
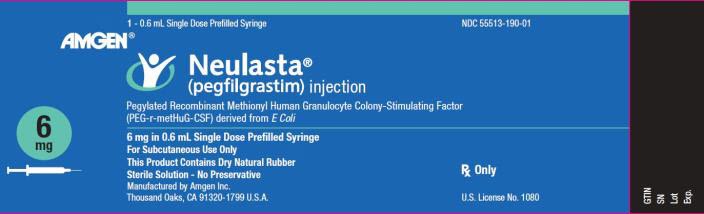
Neulasta pegfilgrastim 6 mg/.6mL Injection, 1 syringe — NDC 55513-190-01 (Billing 55513-0190-01)
This is a package of 1 syringe of Neulasta pegfilgrastim 6 mg/.6mL Injection from Amgen, Inc, marketed since Apr 2002 and currently FDA-listed. It is this product's only package size.
Identity & classification
Regulatory identifiers FDA, NLM and CMS codes for this package
Drug-database identifiers Medi-Span GPI and First Databank GCN / HICL / AHFS classification
- GSN (GCN sequence number): 049872
- GCN: 15666
- HICL (First Databank): 023255
- AHFS class code: 20:16.00.00
- RxCUI (RxNorm): 727539
Where does this data come from?
- FDA openFDA NDC Directory · synced Oct 1, 2026
- FDA label on DailyMed · label index refreshed Oct 1, 2026
- RxNorm (NLM RxNav) · catalog refreshed Oct 1, 2026
- Medi-Span GPI (licensed)
- First Databank (licensed) · refreshed Oct 1, 2026
RxNorm drug class
This medicine belongs to the Leukocyte Growth Factor class.
Where does this data come from?
- RxClass (NLM) · catalog refreshed Oct 1, 2026
Clinical
Pegfilgrastim injection is used to increase white blood cells and decrease the risk of infection in patients receiving chemotherapy for certain types of cancer. Pegfilgrastim injection is also used to increase the chance of survival in people who have been exposed to harmful amounts of radiation, which can cause severe and life-threatening damage to bone marrow. Pegfilgrastim is in a class of medications called colony stimulating factors. It works by helping the body make more neutrophils (a type of blood cell needed to fight infection).
Read the full MedlinePlus article ↗- It helps lower the risk of serious infection with fever in people with non-myeloid cancers getting chemotherapy. Neulasta is also used to improve survival after exposure to dangero...
- For chemotherapy, Neulasta is given once per cycle. It is not given in the 14 days before or the 24 hours after chemotherapy. Your care team will set the exact timing.
- Bone pain and pain in the arms or legs are the most common. Rash, flushing and injection site reactions can also happen.
- Get help for trouble breathing, facial swelling, or hives. Also report pain in the left upper belly or shoulder, fever with breathing problems, or a sickle cell crisis.
Patient education
Supplement & herbal interactions
Where does this data come from?
- MedlinePlus (NLM) · refreshed Oct 1, 2026
- FDA label on DailyMed · label index refreshed Oct 1, 2026
Ask a licensed pharmacist directly — free, answered by our team.
Pricing
A drug doesn't have one price. Each row is a different public payment system, and none is what you'd pay at the counter — that depends on your insurance. The ⓘ on each row explains what it measures.
| Price system | Per each | Per package |
|---|---|---|
| Retail pharmacies payNADAC · weekly | Not in the retail survey — common for institutional, discontinued, or low-volume packs. | |
| Medicaid paysCMS SDUD · 12 mo | $7,413.97 | $7,413.97 / 1 syringe |
| Medicare drug plans payPart D · Q2 2026 | $10,634.70 | $10,634.70 / 1 syringe |
| Medicare Part B allowsASP · J2506 | $143.233 / J2506 unit | — |
Where does this data come from?
- CMS NADAC weekly file
- CMS ASP pricing files · refreshed Sep 20, 2026
- CMS Medicaid State Drug Utilization Data · through Q1 2026
- CMS Part D plan pricing files · refreshed Sep 24, 2026
- VA National Acquisition Center price file
Billing & reimbursement
Where does this data come from?
- CMS ASP NDC-HCPCS crosswalk · refreshed Sep 22, 2026
- DMEPDAC NDC-HCPCS crosswalk
- openFDA NSDE billing units · refreshed Sep 7, 2026
Packaging — all sizes for this product
| Package NDC | Description | Marketing start | Marketing end | Status |
|---|---|---|---|---|
| 55513-0190-01 You're viewing this Main listing | 1 SYRINGE in 1 CARTON / .6 mL in 1 SYRINGE | 2002-04-01 | — | Active |
Therapeutic equivalents
| Product | Labeler | Pack | NADAC/unit | TE | Status | Price vs. this |
|---|---|---|---|---|---|---|
| Neulasta 6 mg/.6mLthis 55513-0190-01 | Amgen, | 1 syringe | — | — | FDA listed | — |
| Fylnetra 6 mg/.6mL 70121-1627-01 | Amneal | 1 syringe | — | — | FDA listed | — |
Where does this data come from?
- FDA openFDA NDC Directory · synced Oct 1, 2026
- FDA Purple Book · refreshed Sep 5, 2026
- CMS NADAC weekly file
Availability & biosimilar status
8 biosimilars are FDA-licensed for this reference biologic — see the list below. (Biologics have no small-molecule generics.)
Why the date isn’t exact: Biosimilar timing can change because patents may be challenged, settled, licensed, added or removed, and litigation can move the real date earlier or later.
🛈 What do these terms mean?
- Biologic patent
- A patent the reference product’s maker has publicly listed. A biosimilar generally can’t launch until these expire — unless they’re invalidated or resolved in a settlement.
- Reference-product exclusivity
- A flat 12 years of FDA market protection from the biologic’s first licensure (the BPCIA). No biosimilar can be licensed before it ends, regardless of patents.
- Interchangeable exclusivity
- The first interchangeable biosimilar can earn a period as the only interchangeable version (pharmacists can substitute it without the prescriber).
- Earliest biosimilar (LOE)
- The latest of all the dates above — the soonest a biosimilar can realistically reach the market. Litigation and settlements can move it earlier.
Biologics have no small-molecule “generics” — competition comes from FDA-licensed biosimilars, tracked in the FDA Purple Book.
| Patent | Type | Use code | Expires |
|---|---|---|---|
| US 9,856,287 ↗ | Biologic patent | — | Jun 21, 2030 |
Is there a biosimilar for NEULASTA 6 MG/0.6 ML SYRINGE?
Why do different websites show different biosimilar dates?
Can a biosimilar launch before the last patent expires?
What does “current Purple Book estimate” mean?
What does “FDA listed” mean?
What does a patent or protection date mean here?
Where does this data come from?
- FDA Purple Book · refreshed Sep 5, 2026
Inactive Ingredients / Excipients
Inactive ingredients, also called excipients, are components of the drug product other than the active ingredient. They may include fillers, dyes, coatings, preservatives, flavors, or other formulation ingredients.
💡 Tap an ingredient (hover on desktop) to see what it is and why it’s used.
-
0.35 mg / 0.6 mL
UNII Q40Q9N063P
Acetic acid is a weak organic acid commonly used in medicines as a buffer and pH adjuster. It helps maintain the proper acidity level to ensure the drug remains stable and effective in its formulation.
-
0.02 mg / 0.6 mL
UNII 7T1F30V5YH
A synthetic emulsifier derived from sorbitol and oleic acid. It helps mix oil and water-based ingredients together and keeps them from separating in liquid formulations.
-
UNII 55X04QC32I
A strong alkaline chemical used to adjust and maintain the pH balance of liquid medicines. It helps keep the medicine stable and ensures it stays effective during storage.
-
30 mg / 0.6 mL
UNII 506T60A25R
Sorbitol is a natural sugar alcohol derived from glucose. It serves as a sweetener, humectant, and bulking agent in medications to improve taste and help maintain moisture in the product.
-
UNII 059QF0KO0R
Water is a liquid solvent that dissolves and mixes ingredients together in liquid medicines, syrups, and injections. It helps distribute the active drug evenly throughout the product.
5 inactive ingredients listed in the exact product block matched to this NDC.
Where does this data come from?
ingredient classCode="IACT" elements from the exact product block matched by this NDC. Label-section narrative from DailyMed / the openFDA label index is shown separately when available.- FDA label on DailyMed · label index refreshed Oct 1, 2026
- FDA openFDA NDC Directory · synced Oct 1, 2026
Inactive ingredient FAQ
Are inactive ingredients the same for every manufacturer?
Why might an inactive ingredient be missing?
Can inactive ingredients matter?
Manufacturer & labeler
Where does this data come from?
- FDA openFDA NDC Directory · synced Oct 1, 2026
- Drugs@FDA
Full prescribing information FDA SPL
🎯 Indications and Usage ▾
1 INDICATIONS AND USAGE Neulasta is a leukocyte growth factor indicated in adult and pediatric patients to Decrease the incidence of infection, as manifested by febrile neutropenia, in patients with non-myeloid malignancies receiving myelosuppressive anti-cancer drugs associated with a clinically significant incidence of febrile neutropenia. ( 1.1 ) Increase survival in patients acutely exposed to myelosuppressive doses of radiation (Hematopoietic Subsyndrome of Acute Radiation Syndrome). ( 1.2 ) Limitations of Use Neulasta is not indicated for the mobilization of peripheral blood progenitor cells for hematopoietic stem cell transplantation.
1.1Patients with Cancer Receiving Myelosuppressive Chemotherapy Neulasta is indicated in adult and pediatric patients to decrease the incidence of infection, as manifested by febrile neutropenia, in patients with non-myeloid malignancies receiving myelosuppressive anti-cancer drugs associated with a clinically significant incidence of febrile neutropenia [see Clinical Studies (14.1) ] . Limitations of Use Neulasta is not indicated for the mobilization of peripheral blood progenitor cells for hematopoietic stem cell transplantation.
1.2Patients with Hematopoietic Subsyndrome of Acute Radiation Syndrome Neulasta is indicated to increase survival in adults and pediatric patients acutely exposed to myelosuppressive doses of radiation [see Dosage and Administration (2.2) and Clinical Studies (14.2) ] .
⏱️ Dosage and Administration ▾
2 DOSAGE AND ADMINISTRATION Patients with cancer receiving myelosuppressive chemotherapy For adult patients of any weight and pediatric patients weighing 45 kg and greater, the recommended dosage is 6 mg subcutaneously once per chemotherapy cycle. ( 2.1 ) For pediatric patients weighing less than 45 kg, use weight-based dosing; refer to Table 1. ( 2.1 ) Do not administer between 14 days before and 24 hours after administration of chemotherapy.
( 2.1 ) Patients acutely exposed to myelosuppressive doses of radiation For adults of any weight and pediatric patients weighing 45 kg and greater, the recommended dosage is two doses, 6 mg each, subcutaneously one week apart. Administer the first dose as soon as possible after suspected or confirmed exposure to myelosuppressive doses of radiation, and a second dose one week after. ( 2.2 ) For pediatric patients weighing less than 45 kg, use weight-based dosing; refer to Table 1.
( 2.1 )
2.1Patients with Cancer Receiving Myelosuppressive Chemotherapy The recommended dosage of Neulasta for adults of any weight and pediatric patients weighing at least 45 kg with cancer receiving myelosuppressive chemotherapy is a single subcutaneous injection of 6 mg administered once per chemotherapy cycle. For dosing in pediatric patients weighing less than 45 kg, refer to Table 1. Do not administer Neulasta between 14 days before and 24 hours after administration of chemotherapy.
Table 1. Dosing of Neulasta for Pediatric Patients Weighing Less Than 45 kg Body Weight Neulasta Dose Volume to Administer Less than 10 kg For pediatric patients weighing less than 10 kg, administer 0.1 mg/kg (0.01 mL/kg) of Neulasta. See below See below 10 kg to 20 kg 1.5 mg 0.15 mL 21 kg to 30 kg 2.5 mg 0.25 mL 31 kg to 44 kg 4 mg 0.4 mL
2.2Patients with Hematopoietic Subsyndrome of Acute Radiation Syndrome The recommended dosage of Neulasta for adults of any weight and pediatric patients weighing at least 45 kg with hematopoietic subsyndrome of acute radiation syndrome is two doses, 6 mg each, administered subcutaneously one week apart. For dosing in pediatric patients weighing less than 45 kg, refer to Table 1 [see Dosage and Administration (2.1) ] . Administer the first dose as soon as possible after suspected or confirmed exposure to radiation levels greater than 2 gray (Gy).
Administer the second dose one week after the first dose. Obtain a baseline complete blood count (CBC). Do not delay administration of Neulasta if a CBC is not readily available.
Estimate a patient's absorbed radiation dose (i.e., level of radiation exposure) based on information from public health authorities, biodosimetry if available, or clinical findings such as time to onset of vomiting or lymphocyte depletion kinetics.
2.3Preparation and Administration Neulasta is supplied in single-dose prefilled syringes for manual use or for use with the on-body injector (OBI) for Neulasta, which is co-packaged with a single-dose prefilled syringe for the OBI, and in single-dose vials [see Dosage Forms and Strengths (3) ] . Before using any of the Neulasta presentations: Remove the carton from the refrigerator and allow the Neulasta prefilled syringe or vial to reach room temperature, 20°C to 25°C (68°F to 77°F), for a minimum of 30 minutes. Do not warm in any other way.
Discard any prefilled syringe or vial left at room temperature for greater than 48 hours. Neulasta is a clear, colorless, preservative-free solution. Parenteral drug products should be inspected visually for particulate matter and discoloration prior to administration, whenever solution and container permit.
Do not administer Neulasta if discoloration or particulates are observed. Prefilled Syringe for Manual Use For adult patients of any weight and pediatric patients weighing 45 kg and greater, the single-dose prefilled syringe for manual use is intended for subcutaneous administration of a single 6 mg/0.6 mL dose of Neulasta. The syringe does not bear graduation marks and therefore does… [Excerpted — this section continues on DailyMed.]
💊 Dosage Forms and Strengths ▾
3 DOSAGE FORMS AND STRENGTHS Neulasta is a clear, colorless, preservative-free solution available as: Injection: 6 mg/0.6 mL in a single-dose prefilled syringe for manual use only. Injection: 6 mg/0.6 mL in a single-dose prefilled syringe co-packaged with the on-body injector (OBI) for Neulasta (Neulasta Onpro kit). Injection: 4 mg/0.4 mL in a single-dose vial.
Injection: 6 mg/0.6 mL solution in a single-dose prefilled syringe for manual use only. ( 3 ) Injection: 6 mg/0.6 mL solution in a single-dose prefilled syringe co-packaged with the on-body injector for Neulasta. ( 3 ) Injection: 4 mg/0.4 mL solution in a single-dose vial.
( 3 )
⛔ Contraindications ▾
4 CONTRAINDICATIONS Neulasta is contraindicated in patients with a history of a serious hypersensitivity reaction to pegfilgrastim or filgrastim. Reactions have included anaphylaxis [see Warnings and Precautions (5.3) ] . In patients with a history of serious hypersensitivity reaction to pegfilgrastim or filgrastim. ( 4 )
⚠️ Warnings and Cautions ▾
5 WARNINGS AND PRECAUTIONS Fatal splenic rupture: Evaluate patients who report left upper abdominal or shoulder pain for an enlarged spleen or splenic rupture. ( 5.1 ) Acute respiratory distress syndrome (ARDS): Evaluate patients who develop fever, lung infiltrates, or respiratory distress. Discontinue Neulasta in patients with ARDS.
( 5.2 ) Serious hypersensitivity reactions, including anaphylaxis: Permanently discontinue Neulasta in patients with serious hypersensitivity reactions. ( 5.3 ) The on-body injector for Neulasta uses acrylic adhesive. For patients who have reactions to acrylic adhesives, use of this product may result in a significant reaction.
( 5.4 ) Fatal sickle cell crises: Discontinue Neulasta if sickle cell crisis occurs. ( 5.5 ) Glomerulonephritis: Evaluate and consider dose-reduction or interruption of Neulasta if causality is likely. ( 5.6 ) Thrombocytopenia: Monitor platelet counts.
( 5.8 ) Myelodysplastic Syndrome (MDS) and Acute Myeloid Leukemia (AML): Monitor patients with breast and lung cancer using Neulasta in conjunction with chemotherapy and/or radiotherapy for signs and symptoms of MDS/AML. ( 5.11 ) Potential device failures: Instruct patients to notify their healthcare provider if they suspect the on-body injector may not have performed as intended. ( 5.12 ) Large and Medium Vessel Arteritis: Discontinue Neulasta if arteritis is suspected.
( 5.13 )
5.1Splenic Rupture Splenic rupture, including fatal cases, can occur following the administration of Neulasta. Evaluate for an enlarged spleen or splenic rupture in patients who report left upper abdominal or shoulder pain after receiving Neulasta.
5.2Acute Respiratory Distress Syndrome Acute respiratory distress syndrome (ARDS) can occur in patients receiving Neulasta. Evaluate patients who develop fever and lung infiltrates or respiratory distress after receiving Neulasta, for ARDS. Discontinue Neulasta in patients with ARDS.
5.3Serious Hypersensitivity Reactions Serious hypersensitivity reactions, including anaphylaxis, can occur in patients receiving Neulasta. The majority of reported events occurred upon initial exposure. Hypersensitivity reactions, including anaphylaxis, can recur within days after the discontinuation of initial therapies to manage the reaction.
Permanently discontinue Neulasta in patients with serious hypersensitivity reactions. Neulasta is contraindicated in patients with a history of a serious hypersensitivity reaction to pegfilgrastim or filgrastim.
5.4Allergies to Acrylics The on-body injector (OBI) for Neulasta uses acrylic adhesive. For patients who have reactions to acrylic adhesives, use of this product may result in a significant reaction.
5.5Use in Patients with Sickle Cell Disorders Severe and sometimes fatal sickle cell crises can occur in patients with sickle cell disorders receiving Neulasta. Discontinue Neulasta if sickle cell crisis occurs.
5.6Glomerulonephritis Glomerulonephritis has occurred in patients receiving Neulasta. The diagnoses were based upon azotemia, hematuria (microscopic and macroscopic), proteinuria, and renal biopsy. Generally, events of glomerulonephritis resolved after dose-reduction or discontinuation of Neulasta. If glomerulonephritis is suspected, evaluate for cause. If causality is likely, consider dose-reduction or interruption of Neulasta.
5.7Leukocytosis White blood cell (WBC) counts of 100 × 10 9 /L or greater have been observed in patients receiving Neulasta. Monitoring of complete blood count (CBC) during Neulasta therapy is recommended.
5.8Thrombocytopenia Neulasta can cause thrombocytopenia. Monitor platelet counts during Neulasta therapy.
5.9Capillary Leak Syndrome Capillary leak syndrome has been reported after G-CSF administration, including Neulasta, and is characterized by hypotension, hypoalbuminemia, edema and hemoconcentration. Episodes vary in frequency, severity and may be life-threatening if treatment is delayed. Patients who develop symptoms of capillary leak… [Excerpted — this section continues on DailyMed.]
🤒 Adverse Reactions ▾
6 ADVERSE REACTIONS The following clinically significant adverse reactions are discussed in greater detail in other sections of the labeling: Splenic Rupture [see Warnings and Precautions (5.1) ] Acute Respiratory Distress Syndrome [see Warnings and Precautions (5.2) ] Serious Hypersensitivity Reactions [see Warnings and Precautions (5.3) ] Allergies to Acrylics [see Warnings and Precautions (5.4) ] Use in Patients with Sickle Cell Disorders [see Warnings and Precautions (5.5) ] Glomerulonephritis [see Warnings and Precautions (5.6) ] Leukocytosis [see Warnings and Precautions (5.7) ] Thrombocytopenia [see Warnings and Precautions (5.8) ] Capillary Leak Syndrome [see Warnings and Precautions (5.9) ] Potential for Tumor Growth Stimulatory Effects on Malignant Cells [see Warnings and Precautions (5.10) ] Myelodysplastic syndrome [see Warnings and Precautions (5.11) ] Acute myeloid leukemia [see Warnings and Precautions (5.11) ] Large and Medium Vessel Arteritis [see Warnings and Precautions (5.13) ] Most common adverse reactions (≥ 5% difference in incidence compared to placebo) are bone pain and pain in extremity.
( 6.1 ) To report SUSPECTED ADVERSE REACTIONS, contact Amgen Inc. at 1-800-77-AMGEN (1-800-772-6436) or FDA at 1-800-FDA-1088 or www.fda.gov/medwatch .
6.1Clinical Trials Experience Because clinical trials are conducted under widely varying conditions, adverse reaction rates observed in the clinical trials of a drug cannot be directly compared with rates in the clinical trials of another drug and may not reflect the rates observed in clinical practice. Neulasta clinical trials safety data are based upon 932 patients receiving Neulasta in seven randomized clinical trials. The population was 21 to 88 years of age and 92% female.
The ethnicity was 75% Caucasian, 18% Hispanic, 5% Black, and 1% Asian. Patients with breast (n = 823), lung and thoracic tumors (n = 53) and lymphoma (n = 56) received Neulasta after nonmyeloablative cytotoxic chemotherapy. Most patients received a single 100 mcg/kg (n = 259) or a single 6 mg (n = 546) dose per chemotherapy cycle over 4 cycles.
The following adverse reaction data in Table 2 are from a randomized, double-blind, placebo-controlled study in patients with metastatic or non-metastatic breast cancer receiving docetaxel 100 mg/m 2 every 21 days (Study 3). A total of 928 patients were randomized to receive either 6 mg Neulasta (n = 467) or placebo (n = 461). The patients were 21 to 88 years of age and 99% female.
The ethnicity was 66% Caucasian, 31% Hispanic, 2% Black, and < 1% Asian, Native American, or other. The most common adverse reactions occurring in ≥ 5% of patients and with a between-group difference of ≥ 5% higher in the pegfilgrastim arm in placebo-controlled clinical trials are bone pain and pain in extremity. Table 2.
Adverse Reactions with ≥ 5% Higher Incidence in Neulasta Patients Compared to Placebo in Study 3 Body System Adverse Reaction Placebo (N = 461) Neulasta 6 mg SC on Day 2 (N = 467) Musculoskeletal and connective tissue disorders Bone pain 26% 31% Pain in extremity 4% 9% Leukocytosis In clinical studies, leukocytosis (WBC counts > 100 × 10 9 /L) was observed in less than 1% of 932 patients with non-myeloid malignancies receiving Neulasta. No complications attributable to leukocytosis were reported in clinical studies.
6.2Postmarketing Experience The following adverse reactions have been identified during post approval use of Neulasta. Because these reactions are reported voluntarily from a population of uncertain size, it is not always possible to reliably estimate their frequency or establish a causal relationship to drug exposure. Splenic rupture and splenomegaly (enlarged spleen) [see Warnings and Precautions (5.1) ] Acute respiratory distress syndrome (ARDS) [see Warnings and Precautions (5.2) ] Allergic reactions/hypersensitivity, including anaphylaxis, skin rash, urticaria, generalized erythema, and flushing [see Warnings and Precautions (5.3) ] Sickle c… [Excerpted — this section continues on DailyMed.]
👥 Use in Specific Populations ▾
8 USE IN SPECIFIC POPULATIONS
8.1Pregnancy Risk Summary Although available data with Neulasta use in pregnant women are insufficient to establish whether there is a drug associated risk of major birth defects, miscarriage, or adverse maternal or fetal outcomes, there are available data from published studies in pregnant women exposed to filgrastim products. These studies have not established an association of filgrastim product use during pregnancy with major birth defects, miscarriage, or adverse maternal or fetal outcomes. In animal studies, no evidence of reproductive/developmental toxicity occurred in the offspring of pregnant rats that received cumulative doses of pegfilgrastim approximately 10 times the recommended human dose (based on body surface area).
In pregnant rabbits, increased embryolethality and spontaneous abortions occurred at 4 times the maximum recommended human dose simultaneously with signs of maternal toxicity (see Data ) . The estimated background risk of major birth defects and miscarriage for the indicated population is unknown. All pregnancies have a background risk of birth defect, loss, or other adverse outcomes.
In the U.S. general population, the estimated background risks of major birth defects and miscarriage in clinically recognized pregnancies is 2-4% and 15-20%, respectively. Data Animal Data Pregnant rabbits were dosed with pegfilgrastim subcutaneously every other day during the period of organogenesis. At cumulative doses ranging from the approximate human dose to approximately 4 times the recommended human dose (based on body surface area), the treated rabbits exhibited decreased maternal food consumption, maternal weight loss, as well as reduced fetal body weights and delayed ossification of the fetal skull; however, no structural anomalies were observed in the offspring from either study.
Increased incidences of post-implantation losses and spontaneous abortions (more than half the pregnancies) were observed at cumulative doses approximately 4 times the recommended human dose, which were not seen when pregnant rabbits were exposed to the recommended human dose. Three studies were conducted in pregnant rats dosed with pegfilgrastim at cumulative doses up to approximately 10 times the recommended human dose at the following stages of gestation: during the period of organogenesis, from mating through the first half of pregnancy, and from the first trimester through delivery and lactation.
No evidence of fetal loss or structural malformations was observed in any study. Cumulative doses equivalent to approximately 3 and 10 times the recommended human dose resulted in transient evidence of wavy ribs in fetuses of treated mothers (detected at the end of gestation but no longer present in pups evaluated at the end of lactation).
8.2Lactation Risk Summary There are no data on the presence of pegfilgrastim in human milk, the effects on the breastfed child, or the effects on milk production. Other filgrastim products are secreted poorly into breast milk, and filgrastim products are not absorbed orally by neonates. The developmental and health benefits of breastfeeding should be considered along with the mother's clinical need for Neulasta and any potential adverse effects on the breastfed child from Neulasta or from the underlying maternal condition.
8.4Pediatric Use The safety and effectiveness of Neulasta for chemotherapy-induced neutropenia and hematopoietic subsyndrome of acute radiation syndrome have been established in pediatric patients. Use of Neulasta in pediatric patients for chemotherapy-induced neutropenia is based on adequate and well-controlled studies in adults with additional pharmacokinetic and safety data in pediatric patients aged 8 months and older with sarcoma [see Clinical Pharmacology (12.3) and Clinical Studies (14.1) ] . The use of Neulasta to increase survival in pediatric patients acutely exposed to myelosuppressive doses of radiation is based on efficacy studies cond… [Excerpted — this section continues on DailyMed.]
🤰 Pregnancy ▾
8.1Pregnancy Risk Summary Although available data with Neulasta use in pregnant women are insufficient to establish whether there is a drug associated risk of major birth defects, miscarriage, or adverse maternal or fetal outcomes, there are available data from published studies in pregnant women exposed to filgrastim products. These studies have not established an association of filgrastim product use during pregnancy with major birth defects, miscarriage, or adverse maternal or fetal outcomes. In animal studies, no evidence of reproductive/developmental toxicity occurred in the offspring of pregnant rats that received cumulative doses of pegfilgrastim approximately 10 times the recommended human dose (based on body surface area).
In pregnant rabbits, increased embryolethality and spontaneous abortions occurred at 4 times the maximum recommended human dose simultaneously with signs of maternal toxicity (see Data ) . The estimated background risk of major birth defects and miscarriage for the indicated population is unknown. All pregnancies have a background risk of birth defect, loss, or other adverse outcomes.
In the U.S. general population, the estimated background risks of major birth defects and miscarriage in clinically recognized pregnancies is 2-4% and 15-20%, respectively. Data Animal Data Pregnant rabbits were dosed with pegfilgrastim subcutaneously every other day during the period of organogenesis. At cumulative doses ranging from the approximate human dose to approximately 4 times the recommended human dose (based on body surface area), the treated rabbits exhibited decreased maternal food consumption, maternal weight loss, as well as reduced fetal body weights and delayed ossification of the fetal skull; however, no structural anomalies were observed in the offspring from either study.
Increased incidences of post-implantation losses and spontaneous abortions (more than half the pregnancies) were observed at cumulative doses approximately 4 times the recommended human dose, which were not seen when pregnant rabbits were exposed to the recommended human dose. Three studies were conducted in pregnant rats dosed with pegfilgrastim at cumulative doses up to approximately 10 times the recommended human dose at the following stages of gestation: during the period of organogenesis, from mating through the first half of pregnancy, and from the first trimester through delivery and lactation.
No evidence of fetal loss or structural malformations was observed in any study. Cumulative doses equivalent to approximately 3 and 10 times the recommended human dose resulted in transient evidence of wavy ribs in fetuses of treated mothers (detected at the end of gestation but no longer present in pups evaluated at the end of lactation).
🧒 Pediatric Use ▾
8.4Pediatric Use The safety and effectiveness of Neulasta for chemotherapy-induced neutropenia and hematopoietic subsyndrome of acute radiation syndrome have been established in pediatric patients. Use of Neulasta in pediatric patients for chemotherapy-induced neutropenia is based on adequate and well-controlled studies in adults with additional pharmacokinetic and safety data in pediatric patients aged 8 months and older with sarcoma [see Clinical Pharmacology (12.3) and Clinical Studies (14.1) ] . The use of Neulasta to increase survival in pediatric patients acutely exposed to myelosuppressive doses of radiation is based on efficacy studies conducted in animals and clinical data supporting the use of Neulasta in patients with cancer receiving myelosuppressive chemotherapy.
Efficacy studies of Neulasta could not be conducted in humans with acute radiation syndrome for ethical and feasibility reasons. Results from population modeling and simulation indicate that two doses of Neulasta (Table 1), administered one week apart provide pediatric patients with exposures comparable to that in adults receiving two 6 mg doses one week apart [see Dosage and Administration (2.1 , 2.2) , Clinical Pharmacology (12.3) and Clinical Studies (14.2) ] .
🧓 Geriatric Use ▾
8.5Geriatric Use Of the 932 patients with cancer who received Neulasta in clinical studies, 139 (15%) were aged 65 and over, and 18 (2%) were aged 75 and over. No overall differences in safety or effectiveness were observed between patients aged 65 and older and younger patients.
🆘 Overdosage ▾
10 OVERDOSAGE Overdosage of Neulasta may result in leukocytosis and bone pain. Events of edema, dyspnea, and pleural effusion have been reported in a single patient who administered Neulasta on 8 consecutive days in error. In the event of overdose, the patient should be monitored for adverse reactions [see Adverse Reactions (6) ] .
Consider contacting the Poison Help line (1-800-222-1222) or a medical toxicologist for additional overdose management recommendations.
🧬 Clinical Pharmacology ▾
12 CLINICAL PHARMACOLOGY
12.1Mechanism of Action Pegfilgrastim is a colony-stimulating factor that acts on hematopoietic cells by binding to specific cell surface receptors, thereby stimulating proliferation, differentiation, commitment, and end cell functional activation.
12.2Pharmacodynamics Animal data and clinical data in humans suggest a correlation between pegfilgrastim exposure and the duration of severe neutropenia as a predictor of efficacy. Selection of the dosing regimen of Neulasta is based on reducing the duration of severe neutropenia.
12.3Pharmacokinetics The pharmacokinetics of pegfilgrastim was studied in 379 patients with cancer. The pharmacokinetics of pegfilgrastim was nonlinear, and clearance decreased with increases in dose. Neutrophil receptor binding is an important component of the clearance of pegfilgrastim, and serum clearance is directly related to the number of neutrophils.
In addition to numbers of neutrophils, body weight appeared to be a factor. Patients with higher body weights experienced higher systemic exposure to pegfilgrastim after receiving a dose normalized for body weight. A large variability in the pharmacokinetics of pegfilgrastim was observed.
The half-life of Neulasta ranged from 15 to 80 hours after subcutaneous injection. In healthy volunteers, the pharmacokinetics of pegfilgrastim were comparable when delivered subcutaneously via a manual prefilled syringe versus via the on-body injector (OBI) for Neulasta. Specific Populations No gender-related differences were observed in the pharmacokinetics of pegfilgrastim, and no differences were observed in the pharmacokinetics of geriatric patients (≥ 65 years of age) compared with younger patients (< 65 years of age) [see Use in Specific Populations (8.5) ] .
Renal Impairment In a study of 30 subjects with varying degrees of renal dysfunction, including end stage renal disease, renal dysfunction had no effect on the pharmacokinetics of pegfilgrastim. Pediatric Patients with Cancer Receiving Myelosuppressive Chemotherapy The pharmacokinetics and safety of pegfilgrastim were studied in 37 pediatric patients with sarcoma in Study 4 [see Clinical Studies (14.1) ] . The mean (± standard deviation [SD]) systemic exposure (AUC 0-inf ) of Neulasta after subcutaneous administration at 100 mcg/kg was 47.9 (± 22.5) mcg ∙ hr/mL in the youngest age group (0 to 5 years, n = 11), 22.0 (± 13.1) mcg ∙ hr/mL in the 6 to 11 years age group (n = 10), and 29.3 (± 23.2) mcg ∙ hr/mL in the 12 to 21 years age group (n = 13).
The terminal elimination half-lives of the corresponding age groups were 30.1 (± 38.2) hours, 20.2 (± 11.3) hours, and 21.2 (± 16.0) hours, respectively. Patients Acutely Exposed to Myelosuppressive Doses of Radiation The pharmacokinetics of pegfilgrastim is not available in patients acutely exposed to myelosuppressive doses of radiation. Based on limited pharmacokinetic data in irradiated non-human primates, the area under the concentration-time curve (AUC), reflecting the exposure to pegfilgrastim in non-human primates following a 300 mcg/kg dose of Neulasta, appears to be greater than in humans receiving a 6 mg dose.
Results from population modeling and simulation indicate that two 6 mg doses of Neulasta administered one week apart in adults result in clinically relevant effects on duration of grade 3 and 4 neutropenia. In addition, weight-based dosing in pediatric patients weighing less than 45 kg [see Dosage and Administration (2.1 , Table 1 and 2.2) ] provides exposures comparable to those in adults receiving two 6 mg doses one week apart.
12.6Immunogenicity The observed incidence of anti-drug antibodies is highly dependent on the sensitivity and specificity of the assay. Differences in assay methods preclude meaningful comparisons of the incidence of anti-drug antibodies in the studies described below with the incidence of anti-drug antibodies in other studies, including those of pegfilgrastim or of other pegfilgrastim products.… [Excerpted — this section continues on DailyMed.]
🧬 Mechanism of Action ▾
12.1Mechanism of Action Pegfilgrastim is a colony-stimulating factor that acts on hematopoietic cells by binding to specific cell surface receptors, thereby stimulating proliferation, differentiation, commitment, and end cell functional activation.
📦 How Supplied / Storage and Handling ▾
16 HOW SUPPLIED/STORAGE AND HANDLING Neulasta single-dose prefilled syringe for manual use Neulasta injection is a clear, colorless solution supplied in a prefilled single-dose syringe for manual use containing 6 mg pegfilgrastim, supplied with a 27-gauge, 1/2-inch needle with an UltraSafe ® Needle Guard. The needle cap of the prefilled syringe contains dry natural rubber (a derivative of latex). Neulasta is provided in a carton containing one sterile 6 mg/0.6 mL prefilled syringe (NDC 55513-190-01).
Neulasta prefilled syringe does not bear graduation marks and is intended only to deliver the entire contents of the syringe (6 mg/0.6 mL) for direct administration. Store refrigerated at 2°C to 8°C (36°F to 46°F) in the original carton to protect from light or physical damage. Discard syringes stored at room temperature for more than 48 hours.
Avoid freezing; if frozen, thaw in the refrigerator before administration. Discard syringe if frozen more than once. Do not shake.
Neulasta Onpro ® kit Neulasta Onpro kit is provided in a carton containing one sterile prefilled syringe and one sterile on-body injector (OBI) for Neulasta (NDC 55513-192-01). The Neulasta injection single-dose prefilled syringe contains 0.64 mL of a clear, colorless solution that delivers 6 mg/0.6 mL of pegfilgrastim when used with the OBI for Neulasta. The prefilled syringe is supplied with a 27-gauge, 1/2-inch needle.
The syringe does not bear graduation marks and is only to be used with the OBI for Neulasta. The needle cap of the prefilled syringe contains dry natural rubber (a derivative of latex). Store Neulasta Onpro kit refrigerated at 2°C to 8°C (36°F to 46°F) in the original carton to protect from light.
Because the OBI for Neulasta is at room temperature during the period of use, Neulasta Onpro kit should not be held at room temperature longer than 12 hours prior to use. Discard Neulasta Onpro kit if stored at room temperature for more than 12 hours. Do not freeze the kit.
Do not shake the prefilled syringe. Do not use the OBI for Neulasta if its packaging has been previously opened. Neulasta single-dose vial Neulasta injection is a clear, colorless, solution, supplied in a vial containing 4 mg pegfilgrastim.
Neulasta is provided in a carton containing one 4 mg/0.4 mL vial (NDC 55513-099-01). Store refrigerated at 2°C to 8°C (36°F to 46°F) in the original carton to protect from light. Do not freeze.
Discard vials stored at room temperature for more than 48 hours. Do not shake.
📦 Storage and Handling ▾
Store refrigerated at 2°C to 8°C (36°F to 46°F) in the original carton to protect from light. Do not freeze. Discard vials stored at room temperature for more than 48 hours. Do not shake.
📋 Description ▾
11 DESCRIPTION Pegfilgrastim is a covalent conjugate of recombinant methionyl human G-CSF (filgrastim) and monomethoxypolyethylene glycol. Filgrastim is a water-soluble 175 amino acid protein with a molecular weight of approximately 19 kilodaltons (kD). Filgrastim is obtained from the bacterial fermentation of a strain of E. coli transformed with a genetically engineered plasmid containing the human G-CSF gene.
To produce pegfilgrastim, a 20 kD monomethoxypolyethylene glycol molecule is covalently bound to the N-terminal methionyl residue of filgrastim. The average molecular weight of pegfilgrastim is approximately 39 kD. Neulasta (pegfilgrastim) is a sterile, clear, colorless, preservative-free solution with a pH of 4.0, supplied in single-dose prefilled syringes for manual use or for use with the on-body injector (OBI) for Neulasta, which is co-packaged with a single-dose prefilled syringe, or in single-dose vials.
Prefilled Syringe for Manual Use or for Use with OBI Neulasta for manual subcutaneous injection is supplied in 0.6 mL prefilled syringes. The prefilled syringe does not bear graduation marks and is designed to deliver the entire contents of the syringe (6 mg/0.6 mL). On-body injector (OBI) for Neulasta is supplied with a prefilled syringe containing 0.64 mL of Neulasta in solution that delivers 0.6 mL of Neulasta in solution when used with the OBI for Neulasta.
The syringe does not bear graduation marks and is only to be used with the OBI for Neulasta. The delivered 0.6 mL dose from either the prefilled syringe for manual subcutaneous injection or the OBI for Neulasta contains pegfilgrastim (6 mg based on protein weight), acetate (0.35 mg), polysorbate 20 (0.02 mg), sodium (0.02 mg), and sorbitol (30 mg) in Water for Injection, USP. Vial Neulasta for subcutaneous injection is supplied in 4 mg/0.4 mL single-dose vials.
Each 0.4 mL vial contains pegfilgrastim (4 mg based on protein weight), acetate (0.24 mg), polysorbate 20 (0.02 mg), sodium (0.01 mg), and sorbitol (20 mg) in Water for Injection, USP.
💬 Information for Patients ▾
17 PATIENT COUNSELING INFORMATION Advise the patient to read the FDA-approved patient labeling (Patient Information and Instructions for Use). Advise patients of the following risks and potential risks with Neulasta: Splenic rupture and splenomegaly Acute Respiratory Distress Syndrome Serious hypersensitivity reactions Sickle cell crisis Glomerulonephritis Increased risk of Myelodysplastic Syndrome and/or Acute Myeloid Leukemia in patients with breast and lung cancer who receive Neulasta in conjunction with chemotherapy and/or radiation therapy Capillary Leak Syndrome Arteritis, including aortitis Advise patients acutely exposed to myelosuppressive doses of radiation (Hematopoietic Subsyndrome of Acute Radiation Syndrome) that efficacy studies of Neulasta for this indication could not be conducted in humans for ethical and feasibility reasons and that, therefore, approval of this use was based on efficacy studies conducted in animals [see Clinical Studies (14.2) ] .
Instruct patients who self-administer Neulasta using the prefilled syringe of the: Importance of following the Instructions for Use. Dangers of reusing syringes. Importance of following local requirements for proper disposal of used syringes.
Advise patients on the use of the on-body injector (OBI) for Neulasta: Review the Patient Information and Patient Instructions for Use with the patient and provide the instructions to the patient. Refer the patient to the dose delivery information written on the Patient Instructions for Use. Tell the patient when their dose delivery of Neulasta will begin and when their dose delivery should be completed.
Advise the patient that serious hypersensitivity reactions can happen with Neulasta. Patients should have a caregiver nearby for the first use. Patients should plan to be in a place where they can appropriately monitor the OBI for Neulasta during the approximately 45 minute Neulasta delivery and for an hour after the delivery.
Advise the patient to avoid traveling, driving, or operating heavy machinery during hours 26 to 29 following application of the OBI for Neulasta. If the OBI for Neulasta is placed on the back of the arm, remind the patient that a caregiver must be available to monitor the OBI for Neulasta. If a patient calls the healthcare provider regarding any OBI for Neulasta problems, the healthcare provider is advised to call Amgen at 1-800-772-6436.
Advise the patient: to call their healthcare provider immediately if the status light on the OBI for Neulasta is flashing red (see the Patient Instructions for Use) . to inform their healthcare provider if the adhesive on the OBI for Neulasta becomes saturated with fluid, or there is dripping, as this may be evidence of significant product leakage, resulting in inadequate or missed dose (see the Patient Instructions for Use) . to keep the OBI for Neulasta dry for approximately the last 3 hours prior to the dose delivery start to better enable potential leak detection. that the OBI for Neulasta should only be exposed to temperatures between 5°C to 40°C (41°F and 104°F). to keep the OBI for Neulasta at least 4 inches away from electrical equipment such as cell phones, cordless telephones, microwaves, and other common appliances.
Failure to keep the OBI for Neulasta at least this recommended distance may interfere with operation and can lead to a missed or incomplete dose of Neulasta. that if the needle is exposed after OBI for Neulasta removal, place the used OBI for Neulasta in a sharps disposal container to avoid accidental needle stick and call their healthcare provider immediately. to remove the OBI for Neulasta after the green light shines continuously and to place the used OBI for Neulasta in a sharps disposal container (see the Patient Instructions for Use) .
Advise the patient: do not reapply the OBI for Neulasta if the OBI for Neulasta comes off before full dose is delivered and instead call their healthcare provider immediately, as they may need a replacement do… [Excerpted — this section continues on DailyMed.]
🧬 Pharmacokinetics ▾
12.3Pharmacokinetics The pharmacokinetics of pegfilgrastim was studied in 379 patients with cancer. The pharmacokinetics of pegfilgrastim was nonlinear, and clearance decreased with increases in dose. Neutrophil receptor binding is an important component of the clearance of pegfilgrastim, and serum clearance is directly related to the number of neutrophils.
In addition to numbers of neutrophils, body weight appeared to be a factor. Patients with higher body weights experienced higher systemic exposure to pegfilgrastim after receiving a dose normalized for body weight. A large variability in the pharmacokinetics of pegfilgrastim was observed.
The half-life of Neulasta ranged from 15 to 80 hours after subcutaneous injection. In healthy volunteers, the pharmacokinetics of pegfilgrastim were comparable when delivered subcutaneously via a manual prefilled syringe versus via the on-body injector (OBI) for Neulasta. Specific Populations No gender-related differences were observed in the pharmacokinetics of pegfilgrastim, and no differences were observed in the pharmacokinetics of geriatric patients (≥ 65 years of age) compared with younger patients (< 65 years of age) [see Use in Specific Populations (8.5) ] .
Renal Impairment In a study of 30 subjects with varying degrees of renal dysfunction, including end stage renal disease, renal dysfunction had no effect on the pharmacokinetics of pegfilgrastim. Pediatric Patients with Cancer Receiving Myelosuppressive Chemotherapy The pharmacokinetics and safety of pegfilgrastim were studied in 37 pediatric patients with sarcoma in Study 4 [see Clinical Studies (14.1) ] . The mean (± standard deviation [SD]) systemic exposure (AUC 0-inf ) of Neulasta after subcutaneous administration at 100 mcg/kg was 47.9 (± 22.5) mcg ∙ hr/mL in the youngest age group (0 to 5 years, n = 11), 22.0 (± 13.1) mcg ∙ hr/mL in the 6 to 11 years age group (n = 10), and 29.3 (± 23.2) mcg ∙ hr/mL in the 12 to 21 years age group (n = 13).
The terminal elimination half-lives of the corresponding age groups were 30.1 (± 38.2) hours, 20.2 (± 11.3) hours, and 21.2 (± 16.0) hours, respectively. Patients Acutely Exposed to Myelosuppressive Doses of Radiation The pharmacokinetics of pegfilgrastim is not available in patients acutely exposed to myelosuppressive doses of radiation. Based on limited pharmacokinetic data in irradiated non-human primates, the area under the concentration-time curve (AUC), reflecting the exposure to pegfilgrastim in non-human primates following a 300 mcg/kg dose of Neulasta, appears to be greater than in humans receiving a 6 mg dose.
Results from population modeling and simulation indicate that two 6 mg doses of Neulasta administered one week apart in adults result in clinically relevant effects on duration of grade 3 and 4 neutropenia. In addition, weight-based dosing in pediatric patients weighing less than 45 kg [see Dosage and Administration (2.1 , Table 1 and 2.2) ] provides exposures comparable to those in adults receiving two 6 mg doses one week apart.
🧬 Pharmacodynamics ▾
12.2Pharmacodynamics Animal data and clinical data in humans suggest a correlation between pegfilgrastim exposure and the duration of severe neutropenia as a predictor of efficacy. Selection of the dosing regimen of Neulasta is based on reducing the duration of severe neutropenia.
🔬 Clinical Studies ▾
14 CLINICAL STUDIES
14.1Patients with Cancer Receiving Myelosuppressive Chemotherapy Neulasta was evaluated in three randomized, double-blind, controlled studies. Studies 1 and 2 were active-controlled studies that employed doxorubicin 60 mg/m 2 and docetaxel 75 mg/m 2 administered every 21 days for up to 4 cycles for the treatment of metastatic breast cancer. Study 1 investigated the utility of a fixed dose of Neulasta.
Study 2 employed a weight-adjusted dose. In the absence of growth factor support, similar chemotherapy regimens have been reported to result in a 100% incidence of severe neutropenia (ANC < 0.5 × 10 9 /L) with a mean duration of 5 to 7 days and a 30% to 40% incidence of febrile neutropenia. Based on the correlation between the duration of severe neutropenia and the incidence of febrile neutropenia found in studies with filgrastim, duration of severe neutropenia was chosen as the primary endpoint in both studies, and the efficacy of Neulasta was demonstrated by establishing comparability to filgrastim-treated patients in the mean days of severe neutropenia.
In Study 1, 157 patients were randomized to receive a single subcutaneous injection of Neulasta (6 mg) on day 2 of each chemotherapy cycle or daily subcutaneous filgrastim (5 mcg/kg/day) beginning on day 2 of each chemotherapy cycle. In Study 2, 310 patients were randomized to receive a single subcutaneous injection of Neulasta (100 mcg/kg) on day 2 or daily subcutaneous filgrastim (5 mcg/kg/day) beginning on day 2 of each chemotherapy cycle. Both studies met the major efficacy outcome measure of demonstrating that the mean days of severe neutropenia of Neulasta-treated patients did not exceed that of filgrastim-treated patients by more than 1 day in cycle 1 of chemotherapy.
The mean days of cycle 1 severe neutropenia in Study 1 were 1.8 days in the Neulasta arm compared to 1.6 days in the filgrastim arm [difference in means 0.2 (95% CI - 0.2, 0.6)] and in Study 2 were 1.7 days in the Neulasta arm compared to 1.6 days in the filgrastim arm [difference in means 0.1 (95% CI - 0.2, 0.4)]. A secondary endpoint in both studies was days of severe neutropenia in cycles 2 through 4 with results similar to those for cycle 1. Study 3 was a randomized, double-blind, placebo-controlled study that employed docetaxel 100 mg/m 2 administered every 21 days for up to 4 cycles for the treatment of metastatic or non-metastatic breast cancer.
In this study, 928 patients were randomized to receive a single subcutaneous injection of Neulasta (6 mg) or placebo on day 2 of each chemotherapy cycle. Study 3 met the major trial outcome measure of demonstrating that the incidence of febrile neutropenia (defined as temperature ≥ 38.2°C and ANC ≤ 0.5 × 10 9 /L) was lower for Neulasta-treated patients as compared to placebo-treated patients (1% versus 17%, respectively, p < 0.001). The incidence of hospitalizations (1% versus 14%) and IV anti-infective use (2% versus 10%) for the treatment of febrile neutropenia was also lower in the Neulasta-treated patients compared to the placebo-treated patients.
Study 4 was a multicenter, randomized, open-label study to evaluate the efficacy, safety, and pharmacokinetics [see Clinical Pharmacology (12.3) ] of Neulasta in pediatric and young adult patients with sarcoma. Patients with sarcoma receiving chemotherapy age 0 to 21 years were eligible. Patients were randomized to receive subcutaneous Neulasta as a single-dose of 100 mcg/kg (n = 37) or subcutaneous filgrastim at a dose 5 mcg/kg/day (n = 6) following myelosuppressive chemotherapy.
Recovery of neutrophil counts was similar in the Neulasta and filgrastim groups. The most common adverse reaction reported was bone pain.
14.2Patients with Hematopoietic Subsyndrome of Acute Radiation Syndrome Efficacy studies of Neulasta could not be conducted in humans with acute radiation syndrome for ethical and feasibility reasons. Approval of this indication was based on efficacy studies conducted in animals… [Excerpted — this section continues on DailyMed.]
🧪 Nonclinical Toxicology ▾
13 NONCLINICAL TOXICOLOGY
13.1Carcinogenesis, Mutagenesis, Impairment of Fertility No carcinogenicity or mutagenesis studies have been performed with pegfilgrastim. Pegfilgrastim did not affect reproductive performance or fertility in male or female rats at cumulative weekly doses approximately 6 to 9 times higher than the recommended human dose (based on body surface area).
📄 Carcinogenesis, Mutagenesis, Impairment of Fertility ▾
13.1Carcinogenesis, Mutagenesis, Impairment of Fertility No carcinogenicity or mutagenesis studies have been performed with pegfilgrastim. Pegfilgrastim did not affect reproductive performance or fertility in male or female rats at cumulative weekly doses approximately 6 to 9 times higher than the recommended human dose (based on body surface area).
📄 Patient Package Insert ▾
Patient Information Neulasta ® (nu-las-tah) (pegfilgrastim) injection Single-Dose Prefilled Syringe and Single-Dose Vial This Patient Information has been approved by the U.S. Food and Drug Administration. Revised: 08/2026 What is Neulasta?
Neulasta is a prescription medicine that is used: in adults and children to reduce the chance of infection due to a low white blood cell count, in people with certain types of cancer (non-myeloid), who receive anti-cancer medicines (chemotherapy) that can cause fever and low blood cell count. to increase survival in adults and children with sudden exposure to high levels of radiation that damage the bone marrow so it cannot make enough blood cells (this is called hematopoietic syndrome of acute radiation syndrome [H-ARS]).
The effectiveness of Neulasta for this use was only studied in animals, because it could not be studied in people. Do not take Neulasta if you have had a serious allergic reaction to pegfilgrastim or filgrastim. Before you receive Neulasta, tell your healthcare provider about all of your medical conditions, including if you: have a sickle cell disorder. have kidney problems. are allergic to latex.
The needle cap on the prefilled syringe contains dry natural rubber (derived from latex). You should not give Neulasta using the prefilled syringe if you have latex allergies. are pregnant or plan to become pregnant. It is not known if Neulasta will harm your unborn baby. are breastfeeding or plan to breastfeed.
It is not known if Neulasta passes into your breast milk. Tell your healthcare provider about all of the medicines you take, including prescription and over-the-counter medicines, vitamins, and herbal supplements. How will I receive Neulasta?
Neulasta is given as an injection under your skin (subcutaneous injection) by a healthcare provider. If your healthcare provider decides that the subcutaneous injections can be given at home by you or your caregiver, follow the detailed "Instructions for Use" that comes with your Neulasta for information on how to prepare and inject a dose of Neulasta using the prefilled syringe. For adults of any weight and children weighing 99 pounds (45 kg) or more: You and your caregiver will be shown how to prepare and inject Neulasta using the prefilled syringe before you use it.
For children weighing less than 99 pounds (45 kg): Do not inject a dose from a Neulasta prefilled syringe. A dose less than 0.6 mL (6 mg) cannot be accurately measured using the Neulasta prefilled syringes. Your healthcare provider will prepare and give the dose of Neulasta using the vials.
If you are receiving Neulasta because you are also receiving chemotherapy, the last dose of Neulasta should be injected at least 14 days before and 24 hours after your dose of chemotherapy. If you miss a dose of Neulasta, talk to your healthcare provider about when you should give your next dose. If you inject too much Neulasta, call your healthcare provider or Poison Help line at 1-800-222-1222 or go to the nearest hospital emergency room right away.
What are possible side effects of Neulasta? Neulasta can cause serious side effects, including: Spleen rupture. Your spleen may become enlarged and can rupture.
A ruptured spleen can cause death. Tell your healthcare provider right away if you develop pain in the left upper stomach area or your left shoulder. A serious lung problem called Acute Respiratory Distress Syndrome (ARDS).
Tell your healthcare provider or get emergency help right away if you develop shortness of breath with or without a fever, trouble breathing, or a fast rate of breathing. Serious allergic reactions. Allergic reactions can cause a rash over your whole body, shortness of breath, wheezing, dizziness, swelling around your mouth or eyes, fast heart rate, and sweating.
If you develop any of these symptoms, stop using Neulasta and tell your healthcare provider or get emergency medical help right away. Sickle cell crises in people with sickle cell disorder. You may… [Excerpted — this section continues on DailyMed.]
📖 Instructions for Use ▾
Neulasta ® (pegfilgrastim) Onpro ® kit Healthcare Provider INSTRUCTIONS FOR USE Guide to Parts Neulasta Prefilled Syringe On-body Injector for Neulasta Important READ THE FOLLOWING INSTRUCTIONS BEFORE USING NEULASTA ONPRO KIT Prescribing Information See Prescribing Information for information on Neulasta. The on-body injector is for adult patients only. The on-body injector is not recommended for patients with Hematopoietic Subsyndrome of Acute Radiation Syndrome.
Neulasta prefilled syringe gray needle cap contains dry natural rubber, which is derived from latex. For patients who have had severe skin reactions to acrylic adhesives, consider the benefit:risk profile before administering pegfilgrastim via the on-body injector for Neulasta. Application Information The on-body injector should be applied to intact, non-irritated skin on the abdomen or back of the arm.
The back of the arm may only be used if there is a caregiver available to monitor the status of the on-body injector. The on-body injector has a self-adhesive backing to attach it to the skin, do not use additional materials to hold it in place as this could dislodge the cannula and lead to a missed or incomplete dose of Neulasta. Environmental Information Do not expose the on-body injector for Neulasta to the following environments as the on-body injector may be damaged and the patient could be injured: Diagnostic imaging (e.g., CT Scan, MRI, Ultrasound, X-ray) Radiation treatment Oxygen rich environments such as hyperbaric chambers Cautions Do not use Neulasta Onpro kit to deliver any other drug product Do not use the on-body injector if its packaging has been previously opened, or the expiration date on the carton or any components has passed.
Do not use if the name Neulasta does not appear on Neulasta Onpro kit carton. Do not modify the on-body injector. Do not attempt to reapply the on-body injector.
Do not use if either the on-body injector or prefilled syringe is dropped. Start again with a new kit Storage Information Store the kit in the refrigerator at 36 ° F to 46 ° F (2 ° C to 8 ° C) until ready for use. If the kit is stored at room temperature (68°F to 77°F [20°C to 25°C] ), for more than 12 hours, do not use.
Start again with a new kit. Keep the prefilled syringe in the kit carton until use to protect from light. Do not freeze the kit.
Do not separate the components of Neulasta Onpro kit until ready for use. For all questions, or if a patient calls you regarding any injector problems, call Amgen at 1-800-772-6436. Step 1: Prepare A Place the syringe tray and the on-body injector tray on a clean, well-lit work surface.
Allow the syringe and on-body injector to come naturally to room temperature for 30 minutes prior to activating. Do not warm the kit components using a heat source. B Choose the patient's injection site.
Ask the patient about their ability to monitor and remove the on-body injector. Use the left or right side of the abdomen, except for a two-inch area right around navel. Use the back of upper arm only if there is a caregiver available to monitor the status of the on-body injector.
Apply the on-body injector to intact, non-irritated skin. Do not apply the on-body injector on surgical sites or areas with scar tissue, moles, or excessive hair. In case of excessive hair, carefully trim hair to get the on-body injector close to the skin.
Do not apply the on-body injector on areas where belts, waistbands, or tight clothing may rub against, disturb, or dislodge the on-body injector. Do not apply the on-body injector on areas where the on-body injector will be affected by folds in the skin. C Clean an area on the injection site larger than the on-body injector adhesive backing.
Thoroughly clean the site with alcohol to enhance on-body adherence to the skin. Only use alcohol to clean the skin. Make sure the skin is oil-free prior to applying the on-body injector.
Allow the skin to completely dry before attaching the on-body injector . Do not touch th… [Excerpted — this section continues on DailyMed.]
📄 Recent Major Changes ▾
Dosage and Administration ( 2 ) 08/2025 Warnings and Precautions, Large and Medium Vessel Arteritis ( 5.13 ) 08/2026
📄 Package Label / Principal Display Panel ▾
PRINCIPAL DISPLAY PANEL - Kit Carton 1 - Single Dose Neulasta Prefilled Syringe 1 - Sterile On-body Injector for Neulasta NDC 55513-192-01 AMGEN Neulasta ® (pegfilgrastim) injection Onpro ® kit Pegylated Recombinant Methionyl Human Granulocyte Colony-Stimulating Factor (PEG-r-metHuG-CSF) derived from E. Coli 6 mg/ 0.6 mL* 6 mg/0.6 mL* * Prefilled syringe contains 0.64 mL to deliver 6 mg/0.6 mL when used with on-body injector For Subcutaneous Use Only Carton Contents (intended to be dispensed as a unit): 1 sterile On-body Injector for Neulasta 1 Neulasta prefilled syringe labeled for use with on-body injector only 1 patient instructions for use 1 healthcare provider instructions for use, 1 Prescribing Information This Product Contains Dry Natural Rubber Store kit refrigerated at 2°C to 8°C (36°F to 46°F) in the original carton to protect from light.
Do Not Freeze or Shake. See full Prescribing Information and Instructions for Use Rx Only LearnOnpro.com PRINCIPAL DISPLAY PANEL - Kit Carton
PRINCIPAL DISPLAY PANEL - 0.6 mL Single Dose Prefilled Syringe NDC 55513-190-01 1 – 0.6 mL Single Dose Prefilled Syringe AMGEN Neulasta ® (pegfilgrastim) injection Pegylated Recombinant Methionyl Human Granulocyte Colony-Stimulating Factor (PEG-r-metHuG-CSF) derived from E Coli 6 mg/ 0.6 mL 6 mg/0.6 mL Single Dose Prefilled Syringe For Subcutaneous Use Only This Product Contains Dry Natural Rubber Sterile Solution - No Preservative Rx Only Manufactured by Amgen Inc. Thousand Oaks, CA 91320-1799 U.S.A. U.S.
License No. 1080 PRINCIPAL DISPLAY PANEL - 0.6 mL Single Dose Prefilled Syringe
PRINCIPAL DISPLAY PANEL - 0.4 mL Vial Carton One 0.4 mL Single-Dose Vial. Discard Unused Portion AMGEN NDC 55513-099-01 Neulasta ® (pegfilgrastim) injection 4 mg/0.4 mL 4 mg/ 0.4 mL For Subcutaneous Use Only Must be administered by a healthcare provider Pegylated Recombinant Methionyl Human Granulocyte Colony-Stimulating Factor (PEG-r-metHuG-CSF) derived from E. Coli Rx Only PRINCIPAL DISPLAY PANEL - 0.4 mL Vial Carton
Medicaid utilization & spend
Medicare Part D spend CMS · PART D · 2026 (Q1)
About this NDC listing & data coverage
What data is (and isn’t) available for this NDC — tap to expand
| NDC identity (package / product / labeler codes) | ✓ Available |
| Labeler | ✓ Available |
| Product & package description | ✓ Available |
| Marketing category & status | ✓ Available |
| Active ingredient / dosage form / route | ✓ Available |
| FDA label (SPL via DailyMed) | ✓ Available |
| Package photos | ✓ Available |
| Inactive ingredients (structured) | ✓ Available |
| NADAC pharmacy acquisition price (CMS) | — Not published for this NDC CMS publishes NADAC only for NDCs reported in its retail-pharmacy survey. |
| Orange Book / therapeutic-equivalence data | ✓ Available |
| HCPCS J-code billing crosswalk | ✓ Available |
| Medicaid utilization (CMS SDUD) | ✓ Available |