
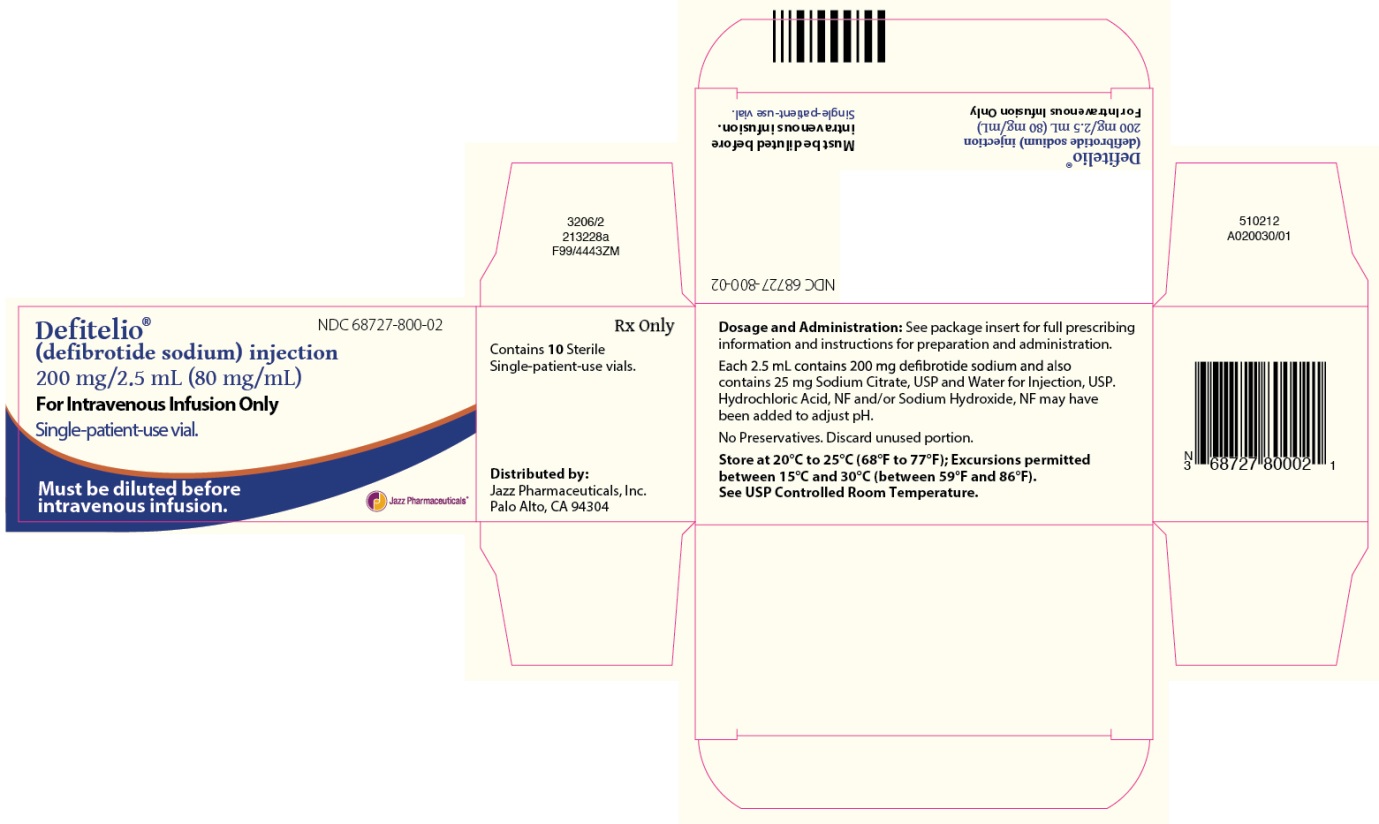
DEFITELIO defibrotide sodium 80 mg/mL Injection, Solution, 25 mL
🆔 Identity & classification
Where does this data come from?
🏷️ RxNorm drug class
This medicine belongs to the Other antithrombotic agents class.
Where does this data come from?
🏭 Manufacturer & labeler
Where does this data come from?
🩺 Clinical
Defibrotide injection is used to treat adults and children with hepatic veno-occlusive disease (VOD; blocked blood vessels inside the liver, also known as sinusoidal obstruction syndrome), who have kidney or lung problems after receiving a hematopoietic stem-cell transplant (HSCT; procedure in which certain blood cells are removed from the body and then returned to the body). Defibrotide injection is in a class of medications called antithrombotic agents. It works by preventing the formation of blood clots.
Read the full MedlinePlus article ↗- Defitelio is used to treat a serious liver complication called hepatic veno-occlusive disease, or VOD — sometimes called sinusoidal obstruction syndrome. It happens when small bloo...
- What exactly is Defitelio used for, and why does my family member need it?
- Defitelio is given through an IV line in a hospital — not at home. Each dose is infused slowly over 2 hours, and it's given four times a day, every 6 hours. The minimum treatment c...
- How is Defitelio given, and how long does treatment last?
Patient education
Supplement & herbal interactions
Some supplements/herbs that may interact with Defibrotide Sodium — tap one for details:
Where does this data come from?
Ask a licensed pharmacist directly — free, answered by our team.
🧪 Inactive Ingredients / Excipients
Inactive ingredients, also called excipients, are components of the drug product other than the active ingredient. They may include fillers, dyes, coatings, preservatives, flavors, or other formulation ingredients.
💡 Tap an ingredient (hover on desktop) to see what it is and why it’s used.
-
UNII QTT17582CB
A strong acid used to adjust and maintain the proper pH level in liquid medicines, ensuring stability and preventing breakdown of active ingredients.
-
UNII 55X04QC32I
A strong alkaline chemical used to adjust and maintain the pH balance of liquid medicines. It helps keep the medicine stable and ensures it stays effective during storage.
-
UNII B22547B95K
A salt derived from citric acid that helps maintain the proper acid-base balance in the medicine. It's used as a buffer to keep the product stable and at the right pH level.
-
UNII 059QF0KO0R
Water is a liquid solvent that dissolves and mixes ingredients together in liquid medicines, syrups, and injections. It helps distribute the active drug evenly throughout the product.
4 inactive ingredients listed in the exact product block matched to this NDC.
Where does this data come from?
ingredient classCode="IACT" elements from the exact product block matched by this NDC. Label-section narrative from DailyMed / the openFDA label index is shown separately when available.Inactive ingredient FAQ
Are inactive ingredients the same for every manufacturer?
Why might an inactive ingredient be missing?
Can inactive ingredients matter?
💲 Pricing
A drug doesn't have one price. Each row is a different public payment system, and none is what you'd pay at the counter — that depends on your insurance. The ⓘ on each row explains what it measures.
| Price system | Per mL | Per package |
|---|---|---|
| Retail pharmacies payNADAC · weekly | Not in the retail survey — common for institutional, discontinued, or low-volume packs. | |
| Medicaid paysCMS SDUD · 12 mo | No recent Medicaid claims on file for this NDC — rare and low-volume NDCs are suppressed in the public data. | |
| Medicare drug plans payPart D · quarterly | No Part D plan price is available for this NDC in our data. | |
Where does this data come from?
🔁 Therapeutic equivalents
| Product | Labeler | Pack | NADAC/unit | TE | Status | Price vs. this |
|---|---|---|---|---|---|---|
| Defitelio 80 mg/mLthis 68727-0800-02 | Jazz | 25 ml | — | — | FDA listed | — |
Where does this data come from?
⏳ Availability & generic status
We did not find an FDA-approved generic match for this exact strength, form and route. Patent/protection dates below may affect future generic timing.
Why the date isn’t exact: Generic timing can change because patents may be challenged, settled, licensed, added, removed, or worked around with a narrower label — and FDA approval does not always mean a pharmacy can get the generic today.
🛈 What do these terms mean?
- Patent
- Legal protection listed in the Orange Book that may delay generic approval or launch. Issued by the U.S. Patent & Trademark Office.
- Substance patent
- Covers the active drug molecule itself — the hardest to design around. A generic generally can’t launch until it expires.
- Formulation (product) patent
- Covers a specific formulation or dosage form. A generic can sometimes work around it with a different formulation.
- Method-of-use patent
- A patent covering one specific approved use of the drug — not necessarily the whole molecule. A generic can sometimes launch with a “skinny label” that carves out the protected use and keeps the others.
- Skinny label
- A generic label that omits a still-patented use when the FDA allows it — letting a generic reach the market for the unprotected uses.
- Exclusivity
- FDA-granted marketing protection, separate from patents — e.g. 5-yr new chemical entity, 7-yr orphan drug, or a +6-month pediatric extension.
- Paragraph IV
- A generic applicant’s formal challenge to a listed patent. It can potentially lead to earlier generic entry, but often involves litigation or a settlement.
- RLD / RS
- Reference Listed Drug — the brand product the FDA uses as the reference for generic applications. Reference Standard — the product the FDA expects generics to compare against in bioequivalence testing.
- TE / AB rating
- FDA therapeutic-equivalence rating. An AB rating generally means the FDA considers a generic therapeutically equivalent to — and substitutable for — the brand.
- LOE (loss of exclusivity)
- The latest patent or exclusivity currently listed — the loss-of-exclusivity / latest-listed-protection date shown on this page. Paragraph-IV challenges and settlements can move the real date earlier; FDA approval and a manufacturer’s decision to market can move it later.
Built from the FDA Orange Book. The bars above are scaled to each protection’s expiry; the red LOE marker is the last one to lapse.
| Patent | Type | Use code | Expires |
|---|---|---|---|
| US 11236328 ↗ | Method of use | U-3301 | Jun 22, 2032 |
| US 11085043 ↗ | Drug product | — | Jun 22, 2032 |
| US 12529052 ↗ | Drug product | — | Jun 22, 2032 |
| US 11746348 ↗ | Drug product | — | Jun 22, 2032 |
| US 12534722 ↗ | Drug product | — | Jun 22, 2032 |
Is there a generic version of DEFITELIO 200 MG/2.5 ML VIAL?
The FDA approved a generic — why can’t I get it at my pharmacy yet?
Why do different websites show different generic release dates?
What does “FDA listed” mean?
What does a patent or protection date mean here?
What does “current Orange Book estimate” mean?
Can a generic come out before the last patent expires?
Can a generic come out after the listed dates?
What is the difference between patents and exclusivity?
Why are there multiple patent dates?
Where does this data come from?
🔬 Reported adverse events (FAERS)
Top reported reactions
Age at onset
Reporter sex
Serious outcomes
Where does this data come from?
📦 Packaging — all sizes for this product
| Package NDC | Description | Marketing start | Status |
|---|---|---|---|
| 68727-0800-01 | 2.5 mL in 1 VIAL (68727-800-01) | 2016-03-30 | Active |
| 68727-0800-02 You're viewing this | 25 mL in 1 CARTON (68727-800-02) | 2016-03-30 | Active |
Pack size FAQ
What quantity is in NDC 68727-0800-02?
What is the difference between NDC 68727-0800-02 and NDC 68727-0800-01?
What NDC number is used to bill for this package of DEFITELIO defibrotide sodium 80 mg/mL Injection, Solution?
🧭 About this NDC listing & data coverage
What data is (and isn’t) available for this NDC — tap to expand
| NDC identity (package / product / labeler codes) | ✓ Available |
| Labeler | ✓ Available |
| Product & package description | ✓ Available |
| Marketing category & status | ✓ Available |
| Active ingredient / dosage form / route | ✓ Available |
| FDA label (SPL via DailyMed) | ✓ Available |
| Package photos | ✓ Available |
| Inactive ingredients (structured) | ✓ Available |
| NADAC pharmacy acquisition price (CMS) | — Not published for this NDC CMS publishes NADAC only for NDCs reported in its retail-pharmacy survey. |
| Orange Book / therapeutic-equivalence data | ✓ Available |
| HCPCS J-code billing crosswalk | — Not published for this NDC Most self-administered / retail products have no J-code — that is normal. |
| Medicaid utilization (CMS SDUD) | — Not published for this NDC CMS reports utilization only for NDCs with Medicaid claims above its privacy threshold. |
Questions about this listing
Why is there no price listed?
Is the NDC printed on the package the same as the 11-digit billing NDC?
What do the three segments of this NDC mean?
Is this package still being marketed?
Does this product come in other package sizes?
Who lists this product with the FDA?
Do I need a prescription for this product?
Where does this data come from?
📄 Full prescribing information FDA SPL
🎯 Indications and Usage ▾
1 INDICATIONS AND USAGE DEFITELIO is indicated for the treatment of adult and pediatric patients with hepatic veno-occlusive disease (VOD), also known as sinusoidal obstruction syndrome (SOS), with renal or pulmonary dysfunction following hematopoietic stem-cell transplantation (HSCT). DEFITELIO is indicated for the treatment of adult and pediatric patients with hepatic veno-occlusive disease (VOD), also known as sinusoidal obstruction syndrome (SOS), with renal or pulmonary dysfunction following hematopoietic stem-cell transplantation (HSCT).
( 1 )
⏱️ Dosage and Administration ▾
2 DOSAGE AND ADMINISTRATION • Administer DEFITELIO 6.25 mg/kg every 6 hours given as a 2-hour intravenous infusion. ( 2.1 ) • Treat for a minimum of 21 days. If after 21 days signs and symptoms of VOD have not resolved, continue treatment until resolution. ( 2.1 )
2.1Recommended Dosage The recommended dosage of DEFITELIO for adult and pediatric patients is 6.25 mg/kg every 6 hours given as a 2‑hour intravenous infusion. The dose should be based on patient’s baseline body weight, defined as the patient’s weight prior to the preparative regimen for HSCT. Administer DEFITELIO for a minimum of 21 days.
If after 21 days signs and symptoms of hepatic VOD have not resolved, continue DEFITELIO until resolution of VOD or up to a maximum of 60 days.
2.2Administration Instructions • DEFITELIO must be diluted prior to infusion [see Dosage and Administration (2.4) ] . • Prior to administration of DEFITELIO, confirm that the patient is not experiencing clinically significant bleeding and is hemodynamically stable on no more than one vasopressor [see Warnings and Precautions (5.1) ] . • Administer DEFITELIO by constant intravenous infusion over a 2-hour period. • Administer the diluted DEFITELIO solution using an infusion set equipped with a 0.2 micron in-line filter.
Flush the intravenous administration line (peripheral or central) with 5% Dextrose Injection, USP or 0.9% Sodium Chloride Injection, USP immediately before and after administration. • Do not co‑administer DEFITELIO and other intravenous drugs concurrently within the same intravenous line.
2.3Treatment Modification Treatment modification, including temporary or permanent discontinuation of DEFITELIO, should follow the recommendations in Table 1. Table 1: Treatment Modifications for Toxicity or Invasive Procedures Event Recommended Action Hypersensitivity Reaction Severe or life-threatening (anaphylaxis) 1. Discontinue DEFITELIO permanently; do not resume treatment.
Bleeding Persistent, severe or potentially life-threatening 1. Withhold DEFITELIO. 2.
Treat the cause of bleeding and give supportive care as clinically indicated. 3. Consider resuming treatment (at the same dose and infusion volume) when bleeding has stopped and the patient is hemodynamically stable.
Recurrent significant bleeding 1. Discontinue DEFITELIO permanently; do not resume treatment. Invasive Procedures 1.
There is no known reversal agent for the profibrinolytic effects of DEFITELIO. Discontinue DEFITELIO infusion at least 2 hours prior to an invasive procedure. 2.
Resume DEFITELIO treatment after the procedure as soon as any procedure-related risk of bleeding is resolved.
2.4Preparation Instructions Dilute DEFITELIO in 5% Dextrose Injection, USP or 0.9% Sodium Chloride Injection, USP to a concentration of 4 mg/mL to 20 mg/mL. Administer the diluted solution over 2 hours. Vials contain no antimicrobial preservatives and are intended for a single-patient-use only.
Partially used vials should be discarded. Use the diluted DEFITELIO solution within 4 hours if stored at room temperature or within 24 hours if stored under refrigeration. Up to four doses of DEFITELIO solution may be prepared at one time, if refrigerated.
Preparation Instructions: • Determine the dose (mg) and number of vials of DEFITELIO based on the individual patient’s baseline weight (weight prior to the preparative regimen for HSCT). • Calculate the volume of DEFITELIO needed, withdraw this amount from the vial(s) and add it to the infusion bag containing 0.9% Sodium Chloride Injection or 5% Dextrose Injection for each dose to make a final concentration of 4 mg/mL to 20 mg/mL. • Gently mix the solution for infusion. • Parenteral drug products should be inspected visually for particulate matter and discoloration prior to administration, whenever solution and container permit.
Only clear solutions without visible particles should be used. Depending on the type and amount of diluent, the color of the diluted solution may vary from colorless t…
💊 Dosage Forms and Strengths ▾
3 DOSAGE FORMS AND STRENGTHS Injection: 200 mg/2.5 mL (80 mg/mL) of defibrotide sodium as a clear, light yellow to brown solution in a single-patient-use glass vial. Injection: 200 mg/2.5 mL (80 mg/mL) in a single-patient-use vial. ( 3 )
⛔ Contraindications ▾
4 CONTRAINDICATIONS The use of DEFITELIO is contraindicated in the following conditions: • Concomitant administration with systemic anticoagulant or fibrinolytic therapy [see Warnings and Precautions (5.1) ] • Known hypersensitivity to DEFITELIO or to any of its excipients [see Warnings and Precautions (5.2) ] • Concomitant administration with systemic anticoagulant or fibrinolytic therapy. ( 4 ) • Known hypersensitivity to DEFITELIO or to any of its excipients. ( 4 )
⚠️ Warnings and Cautions ▾
5 WARNINGS AND PRECAUTIONS • Hemorrhage: Monitor patients for bleeding. Withhold or discontinue DEFITELIO if significant bleeding occurs. ( 2.3 , 5.1 ) • Hypersensitivity Reactions: If severe or life threatening allergic reaction occurs, discontinue DEFITELIO, treat according to standard of care, and monitor until signs and symptoms resolve. ( 2.3 , 5.2 )
5.1Hemorrhage DEFITELIO increased the activity of fibrinolytic enzymes in vitro, and it may increase the risk of bleeding in patients with VOD after hematopoietic stem-cell transplantation (HSCT). Do not initiate DEFITELIO in patients with active bleeding. Monitor patients for signs of bleeding.
If patients on DEFITELIO develop bleeding, discontinue DEFITELIO, treat the underlying cause, and provide supportive care until the bleeding has stopped [see Dosage and Administration (2.3) ] . Concomitant use of DEFITELIO and a systemic anticoagulant or fibrinolytic therapy (not including use for routine maintenance or reopening of central venous lines) may increase the risk of bleeding. Discontinue anticoagulants and fibrinolytic agents prior to DEFITELIO treatment, and consider delaying the start of DEFITELIO administration until the effects of the anticoagulant have abated [see Contraindications (4) ] .
5.2Hypersensitivity Reactions Hypersensitivity reactions have occurred in less than 2% of patients treated with DEFITELIO. These reactions include rash, urticaria and angioedema. One case of an anaphylactic reaction was reported in a patient who had previously received DEFITELIO.
Monitor patients for hypersensitivity reactions, especially if there is a history of previous exposure. If a severe hypersensitivity reaction occurs, discontinue DEFITELIO, treat according to the standard of care, and monitor until symptoms resolve [see Dosage and Administration (2.3) ] .
🤒 Adverse Reactions ▾
6 ADVERSE REACTIONS The following adverse reactions are discussed in greater detail in other sections of the labeling: • Hemorrhage [see Warnings and Precautions (5.1) ] • Hypersensitivity Reactions [see Warnings and Precautions (5.2) ] The most common adverse reactions (incidence ≥10% and independent of causality) with DEFITELIO treatment were hypotension, diarrhea, vomiting, nausea, and epistaxis. ( 6.1 ) To report SUSPECTED ADVERSE REACTIONS, contact Jazz Pharmaceuticals, Inc. at 1-800-520-5568 or FDA at 1-800-FDA-1088 or www.fda.gov/medwatch.
6.1Clinical Trials Experience Because clinical trials are conducted under widely varying conditions, adverse reaction rates observed in the clinical trials of a drug cannot be directly compared to rates in the clinical trials of another drug and may not reflect the rates observed in practice. The safety of DEFITELIO was determined in 176 adult and pediatric patients with hepatic VOD with pulmonary and/or renal dysfunction following HSCT who were treated with DEFITELIO 6.25 mg/kg every 6 hours [see Clinical Studies (14) ] .
Patients were excluded from these trials if at time of study entry they had significant acute bleeding, active grades B-D graft-versus-host disease, or a requirement for multiple vasopressors to provide blood pressure support. For the purposes of adverse event recording in the clinical trials, events were not required to be reported if they were related to the hepatic VOD, or if they were expected to occur after hematopoietic stem-cell transplantation (HSCT), unless they were serious or Grade 4-5. The median age of the safety population was 25 years (range: 1 month to 72 years), and 63% were ≥17 years of age.
A total of 60% of patients were male, 78% were white, 89% had undergone allogeneic HSCT, and the underlying diagnosis was acute leukemia for 43%. At study entry, 13% were dialysis dependent and 18% were ventilator dependent. DEFITELIO was administered for a median of 21 days (range: 1 to 83 days).
Information about adverse reactions resulting in permanent discontinuation of DEFITELIO was available for 102 patients, and 35 (34%) of these patients had an adverse reaction with permanent discontinuation. Adverse reactions leading to permanent discontinuation included pulmonary alveolar hemorrhage in 5 (5%) patients; pulmonary hemorrhage, hypotension, catheter site hemorrhage, and multi-organ failure, each in 3 (3%) patients; and cerebral hemorrhage and sepsis, each in 2 (2%) patients. Information about adverse reactions of any grade was available for all 176 patients.
The most common adverse reactions (incidence ≥10% and independent of causality) were hypotension, diarrhea, vomiting, nausea, and epistaxis. The most common serious adverse reactions (incidence ≥5% and independent of causality) were hypotension (11%) and pulmonary alveolar hemorrhage (7%). Hemorrhage events of any type and any grade were reported for 104 (59%) of the patients, and the events were grade 4-5 in 35 (20%).
Table 2 presents adverse reactions independent of causality ≥10% any grade or Grade 4/5 ≥2% reported in patients treated with DEFITELIO. Table 2: Adverse Reactions a ≥10% or Grade 4-5 Adverse Reactions ≥2% DEFITELIO (n=176) Adverse Reaction a Any grade Grade 4-5 b Hypotension 65 (37%) 12 (7%) Diarrhea 43 (24%) 0 Vomiting 31 (18%) 0 Nausea 28 (16%) 0 Epistaxis 24 (14%) 0 Pulmonary alveolar hemorrhage 15 (9%) 12 (7%) Gastrointestinal hemorrhage 15 (9%) 5 (3%) Sepsis 12 (7%) 9 (5%) Graft versus host disease 11 (6%) 7 (4%) Lung infiltration 10 (6%) 5 (3%) Pneumonia 9 (5%) 5 (3%) Pulmonary hemorrhage 7 (4%) 4 (2%) Infection 6 (3%) 4 (2%) Hemorrhage intracranial 5 (3%) 4 (2%) Hyperuricemia 4 (2%) 4 (2%) Cerebral hemorrhage c 3 (2%) 3 (2%) a Excludes events considered to be due to the underlying disease: multi-organ failure, veno-occlusive disease, respiratory failure, renal failure, and hypoxia b Adverse reactions considered life-threatening or fatal c Cerebral hemorrhage has been included i…
🔄 Drug Interactions ▾
7 DRUG INTERACTIONS Antithrombotic Agents DEFITELIO may enhance the pharmacodynamic activity of antithrombotic/fibrinolytic drugs such as heparin or alteplase. Concomitant use of DEFITELIO with antithrombotic or fibrinolytic drugs is contraindicated because of an increased risk of hemorrhage [see Contraindications (4) ] . • DEFITELIO may enhance the activity of antithrombotic/fibrinolytic drugs. ( 7 )
👥 Use in Specific Populations ▾
8 USE IN SPECIFIC POPULATIONS
8.1Pregnancy Risk Summary There are no available data on DEFITELIO use in pregnant women. When administered to pregnant rabbits during the period of organogenesis at doses that were comparable to the recommended human dose based on body surface area, defibrotide sodium decreased the number of implantations and viable fetuses. Advise pregnant women of the potential risk of miscarriage.
The estimated background risk of major birth defects and miscarriage for the indicated population is unknown. In the U.S. general population, the estimated background risks of major birth defects and miscarriage in clinically recognized pregnancies are 2-4% and 15-20%, respectively. Data Animal Data Embryo-Fetal toxicity assessment was attempted in rats and rabbits, but was not possible because of high maternal mortality, abortion, and fetal resorption at all doses.
Pregnant rats were administered defibrotide sodium from gestational day (GD) 6 to 15 at 0, 240, 1200, and 4800 mg/kg/day by continuous intravenous infusion over 24 hours or at 60, 120, and 240 mg/kg/day by 2-hour infusions 4 times per day. Pregnant rabbits were administered defibrotide sodium at 0, 30, 60, or 120 mg/kg/day from GD 6 to 18 by 2-hour infusions 4 times per day. In another study in pregnant rabbits, 3 separate subgroups of animals were treated with doses of 80 mg/kg/day defibrotide sodium administered by 2-hour infusions 4 times per day for 5 days each in a staggered manner during the organogenesis period.
The dose of 80 mg/kg/day is approximately equivalent to the recommended clinical dose on a mg/m 2 basis. Subgroup 1 was dosed from GD 6 to 10, subgroup 2 was dosed from GD 10 to 14, and subgroup 3 was dosed from GD 14 to 18. An increased incidence of unilateral implantation was observed in defibrotide sodium-treated animals.
Treatment with defibrotide sodium resulted in a decreased number of implantations and viable fetuses.
8.2Lactation Risk Summary There is no information regarding the presence of DEFITELIO in human milk, the effects on the breastfed infant, or the effects on milk production. Because of the potential for serious adverse reactions, including bleeding in a breastfed infant, advise patients that breastfeeding is not recommended during treatment with DEFITELIO.
8.4Pediatric Use The safety and effectiveness of DEFITELIO have been established in pediatric patients. Use of DEFITELIO is supported by evidence from an adequate and well-controlled study and a dose finding study of DEFITELIO in adult and pediatric patients with VOD with evidence of renal or pulmonary dysfunction following HSCT. The clinical trials enrolled 66 pediatric patients in the following age groups: 22 infants (1 month up to less than 2 years), 30 children (2 years up to less than 12 years), and 14 adolescents (12 years to less than 17 years).
The efficacy and safety outcomes were consistent across pediatric and adult patients in the clinical trials [see Adverse Reactions (6) and Clinical Studies (14) ] . Juvenile Animal Toxicity Data A juvenile toxicity study in 21-day-old rats was conducted with intravenous bolus administration of defibrotide sodium at 40, 150, or 320 mg/kg/day for 4 weeks. A delayed mean age of preputial separation was observed at all doses, suggesting a delay in onset of male puberty.
The dose of 40 mg/kg/day is approximately 0.4 times the clinical dose on a mg/m 2 basis for a child. The relevance of this finding for the onset of male puberty in humans is unknown.
8.5Geriatric Use Clinical studies of DEFITELIO did not include sufficient numbers of subjects aged 65 and over to determine whether they respond differently from younger subjects. Other reported clinical experience has not identified differences in responses between the elderly and younger patients.
🤰 Pregnancy ▾
8.1Pregnancy Risk Summary There are no available data on DEFITELIO use in pregnant women. When administered to pregnant rabbits during the period of organogenesis at doses that were comparable to the recommended human dose based on body surface area, defibrotide sodium decreased the number of implantations and viable fetuses. Advise pregnant women of the potential risk of miscarriage.
The estimated background risk of major birth defects and miscarriage for the indicated population is unknown. In the U.S. general population, the estimated background risks of major birth defects and miscarriage in clinically recognized pregnancies are 2-4% and 15-20%, respectively. Data Animal Data Embryo-Fetal toxicity assessment was attempted in rats and rabbits, but was not possible because of high maternal mortality, abortion, and fetal resorption at all doses.
Pregnant rats were administered defibrotide sodium from gestational day (GD) 6 to 15 at 0, 240, 1200, and 4800 mg/kg/day by continuous intravenous infusion over 24 hours or at 60, 120, and 240 mg/kg/day by 2-hour infusions 4 times per day. Pregnant rabbits were administered defibrotide sodium at 0, 30, 60, or 120 mg/kg/day from GD 6 to 18 by 2-hour infusions 4 times per day. In another study in pregnant rabbits, 3 separate subgroups of animals were treated with doses of 80 mg/kg/day defibrotide sodium administered by 2-hour infusions 4 times per day for 5 days each in a staggered manner during the organogenesis period.
The dose of 80 mg/kg/day is approximately equivalent to the recommended clinical dose on a mg/m 2 basis. Subgroup 1 was dosed from GD 6 to 10, subgroup 2 was dosed from GD 10 to 14, and subgroup 3 was dosed from GD 14 to 18. An increased incidence of unilateral implantation was observed in defibrotide sodium-treated animals.
Treatment with defibrotide sodium resulted in a decreased number of implantations and viable fetuses.
🧒 Pediatric Use ▾
8.4Pediatric Use The safety and effectiveness of DEFITELIO have been established in pediatric patients. Use of DEFITELIO is supported by evidence from an adequate and well-controlled study and a dose finding study of DEFITELIO in adult and pediatric patients with VOD with evidence of renal or pulmonary dysfunction following HSCT. The clinical trials enrolled 66 pediatric patients in the following age groups: 22 infants (1 month up to less than 2 years), 30 children (2 years up to less than 12 years), and 14 adolescents (12 years to less than 17 years).
The efficacy and safety outcomes were consistent across pediatric and adult patients in the clinical trials [see Adverse Reactions (6) and Clinical Studies (14) ] . Juvenile Animal Toxicity Data A juvenile toxicity study in 21-day-old rats was conducted with intravenous bolus administration of defibrotide sodium at 40, 150, or 320 mg/kg/day for 4 weeks. A delayed mean age of preputial separation was observed at all doses, suggesting a delay in onset of male puberty.
The dose of 40 mg/kg/day is approximately 0.4 times the clinical dose on a mg/m 2 basis for a child. The relevance of this finding for the onset of male puberty in humans is unknown.
🧓 Geriatric Use ▾
8.5Geriatric Use Clinical studies of DEFITELIO did not include sufficient numbers of subjects aged 65 and over to determine whether they respond differently from younger subjects. Other reported clinical experience has not identified differences in responses between the elderly and younger patients.
🆘 Overdosage ▾
10 OVERDOSAGE There are no known cases of overdose with DEFITELIO. There is no known antidote for DEFITELIO, and DEFITELIO is not dialyzable. If an overdose occurs, institute general supportive measures.
🧬 Clinical Pharmacology ▾
12 CLINICAL PHARMACOLOGY
12.1Mechanism of Action The mechanism of action of defibrotide sodium has not been fully elucidated. In vitro , defibrotide sodium enhances the enzymatic activity of plasmin to hydrolyze fibrin clots. Studies evaluating the pharmacological effects of defibrotide sodium on endothelial cells (ECs) were conducted primarily in the human microvascular endothelial cell line.
In vitro , defibrotide sodium increased tissue plasminogen activator (t-PA) and thrombomodulin expression, and decreased von Willebrand factor (vWF) and plasminogen activator inhibitor‑1 (PAI-1) expression, thereby reducing EC activation and increasing EC‑mediated fibrinolysis. Defibrotide sodium protected ECs from damage caused by chemotherapy, tumor necrosis factor-α (TNF-α), serum starvation, and perfusion.
12.2Pharmacodynamics Cardiac Electrophysiology At a dose 2.4 times the maximum recommended dose, DEFITELIO does not prolong the QTc interval to any clinically relevant extent. PAI-1 Inhibition Plasma concentrations of PAI-1 were assessed on an exploratory basis as a potential pharmacodynamic marker for efficacy in Study 2. PAI-1 is an inhibitor of t-PA and therefore of fibrinolysis.
Mean PAI-1 levels on Days 7 and 14 were lower than those at baseline in patients with complete response (CR) and in those who were alive at Day+100, but this trend did not reach statistical significance. There were no statistically significant differences in mean PAI-1 levels by treatment or outcome.
12.3Pharmacokinetics Absorption After intravenous administration, peak plasma concentrations of defibrotide sodium occur approximately at the end of each infusion. Distribution Defibrotide sodium is highly bound to human plasma proteins (average 93%) and has a volume of distribution of 8.1 to
9.1L. Elimination Metabolism followed by urinary excretion is likely the main route of elimination. The estimated total clearance was 3.4 to
6.1L/h. The elimination half-life of defibrotide sodium is less than 2 hours. Similar plasma concentration profiles were observed in VOD patients after initial and multiple-dose administration of 6.25 mg/kg every 6 hours for 5 days.
Therefore, no accumulation is expected following multiple-dose administration. Metabolism Though the precise pathway of defibrotide sodium degradation in plasma in vivo is largely unknown, it has been suggested that nucleases, nucleotidases, nucleosidases, deaminases, and phosphorylases metabolize polynucleotides progressively to oligonucleotides, nucleotides, nucleosides, and then to the free 2'-deoxyribose sugar, purine and pyrimidine bases. The biotransformation of defibrotide sodium was investigated in vitro by incubation with human hepatocytes from donors of different ages and showed that defibrotide sodium does not undergo appreciable metabolism by human hepatocyte cells.
Excretion After administration of 6.25 mg/kg to 15 mg/kg doses of DEFITELIO as 2-hour infusions, approximately 5-15% of the total dose was excreted in urine as defibrotide sodium, with the majority excreted during the first 4 hours. Specific Populations Age: Pediatric Population Insufficient PK data were collected in pediatric patients to draw conclusions. Renal Impairment The safety, tolerability, and pharmacokinetics of 6.25 mg/kg as 2-hour intravenous infusions of DEFITELIO were evaluated in patients with Hemodialysis-dependent End Stage Renal Disease (ESRD) during hemodialysis and on days off dialysis, and in patients with severe renal disease or ESRD not requiring dialysis.
Defibrotide sodium was not removed by hemodialysis, which had no notable effect on plasma clearance of defibrotide sodium. Terminal half-lives were consistently less than 2 hours, and there was no accumulation of defibrotide sodium following repeated dosing. Defibrotide sodium exposure (AUC) in patients with severe renal impairment or ESRD was 50% to 60% higher than that observed in matched healthy subjects.
Peak concentration (C max ) was 35% to 37…
🧬 Mechanism of Action ▾
12.1Mechanism of Action The mechanism of action of defibrotide sodium has not been fully elucidated. In vitro , defibrotide sodium enhances the enzymatic activity of plasmin to hydrolyze fibrin clots. Studies evaluating the pharmacological effects of defibrotide sodium on endothelial cells (ECs) were conducted primarily in the human microvascular endothelial cell line.
In vitro , defibrotide sodium increased tissue plasminogen activator (t-PA) and thrombomodulin expression, and decreased von Willebrand factor (vWF) and plasminogen activator inhibitor‑1 (PAI-1) expression, thereby reducing EC activation and increasing EC‑mediated fibrinolysis. Defibrotide sodium protected ECs from damage caused by chemotherapy, tumor necrosis factor-α (TNF-α), serum starvation, and perfusion.
📦 How Supplied / Storage and Handling ▾
16 HOW SUPPLIED/STORAGE AND HANDLING DEFITELIO (defibrotide sodium) injection is supplied in a single-patient-use, clear glass vial as a clear, light yellow to brown, sterile, preservative-free solution for intravenous infusion. Each vial (NDC 68727-800-01) contains 200 mg/2.5 mL (at a concentration of 80 mg/mL) of defibrotide sodium. Each carton of DEFITELIO (defibrotide sodium) injection (NDC 68727-800-02) contains 10 vials.
Store DEFITELIO (defibrotide sodium) injection at 20°C-25°C (68°F-77°F); excursions permitted between 15°C to 30°C (59°F to 86°F) (see USP controlled room temperature).
📋 Description ▾
11 DESCRIPTION Defibrotide sodium is an oligonucleotide mixture with profibrinolytic properties. The chemical name of defibrotide sodium is polydeoxyribonucleotide, sodium salt. Defibrotide sodium is a polydisperse mixture of predominantly single-stranded (ss) polydeoxyribonucleotide sodium salts derived from porcine intestinal tissue having a mean weighted molecular weight of 14-19 kDa, and a potency of 27-39 and 28-38 biological units per mg as determined by two separate assays measuring the release of a product formed by contact between defibrotide sodium, plasmin and a plasmin substrate.
The primary structure of defibrotide sodium is shown below. DEFITELIO (defibrotide sodium) injection is a clear, light yellow to brown, sterile, preservative-free solution in a single-patient-use vial for intravenous use. Each milliliter of the injection contains 80 mg of defibrotide sodium and 10 mg of Sodium Citrate, USP, in Water for Injection, USP.
Hydrochloric Acid, NF, and/or Sodium Hydroxide, NF, may have been used to adjust pH to 6.8-7.8. chemical structure
💬 Information for Patients ▾
17 PATIENT COUNSELING INFORMATION • Hemorrhage: Advise patients and caregivers that DEFITELIO may increase the risk of bleeding (hemorrhage). Instruct patients to immediately report any signs or symptoms suggestive of hemorrhage (unusual bleeding, easy bruising, blood in urine or stool, headache, confusion, slurred speech, or altered vision) [see Warnings and Precautions (5.1) ] . • Hypersensitivity Reactions: Ask patients if they have been treated with defibrotide sodium previously. Instruct patients on the risk of allergic reactions, including anaphylaxis.
Describe the symptoms of allergic reactions, including anaphylaxis, and instruct the patient to seek medical attention immediately if they experience such symptoms [see Warnings and Precautions (5.2) ] .