
Veozah fezolinetant 45 mg Tablet, Film Coated — NDC 0469-2760-28 (Billing 00469-2760-28)
This is a package of Veozah fezolinetant 45 mg Tablet, Film Coated from Astellas Pharma US, Inc., marketed since May 2023 and currently FDA-listed. It is this product's only package size.
NDC database record
One package, one record: these facts belong to NDC 0469-2760-28 alone.
- Record
- FDA NDC Directory package listing · Human prescription drug
- Code segments
- 0469 labeler · 2760 product · 28 package
- Package marketed since
- May 12, 2023
- Sample package
- Yes — professional sample, not for sale
- Listing certified through
- Dec 31, 2027
- Barcode (UPC-A, from the NDC)
- 3 0469276028 1
- FDA record last changed
- Jul 24, 2026
Identity & classification
Regulatory identifiers FDA, NLM and CMS codes for this package
Drug-database identifiers Medi-Span GPI and First Databank GCN / HICL / AHFS classification
- GSN (GCN sequence number): 084780
- GCN: 54158
- HICL (First Databank): 048921
- AHFS class code: 28:92.00.00
- RxCUI (RxNorm): 2637141
Where does this data come from?
- FDA openFDA NDC Directory · synced Oct 1, 2026
- FDA label on DailyMed · label index refreshed Oct 3, 2026
- RxNorm (NLM RxNav) · catalog refreshed Oct 1, 2026
- Medi-Span GPI (licensed)
- First Databank (licensed) · refreshed Oct 1, 2026
RxNorm drug class
This medicine belongs to the Neurokinin 3 Receptor Antagonist class.
Where does this data come from?
- RxClass (NLM) · catalog refreshed Oct 1, 2026
Clinical
Patient education
Supplement & herbal interactions
Some supplements/herbs that may interact with Fezolinetant — tap one for details:
Where does this data come from?
- MedlinePlus (NLM) · refreshed Oct 1, 2026
- FDA label on DailyMed · label index refreshed Oct 3, 2026
Ask a licensed pharmacist directly — free, answered by our team.
Pricing
A drug doesn't have one price. Each row is a different public payment system, and none is what you'd pay at the counter — that depends on your insurance. The ⓘ on each row explains what it measures.
| Price system | Per each | Per package |
|---|---|---|
| Retail pharmacies payNADAC · weekly | Not in the retail survey — common for institutional, discontinued, or low-volume packs. | |
| Medicaid paysCMS SDUD · 12 mo | No recent Medicaid claims on file for this NDC — rare and low-volume NDCs are suppressed in the public data. | |
| Medicare drug plans payPart D · Q2 2026 | $19.63 | $549.70 / 28 tablets |
Where does this data come from?
- CMS NADAC weekly file
- CMS ASP pricing files · refreshed Sep 20, 2026
- CMS Medicaid State Drug Utilization Data · refreshed Oct 4, 2026
- CMS Part D plan pricing files · refreshed Sep 24, 2026
- VA National Acquisition Center price file
Packaging — all sizes for this product
| Package NDC | Description | Marketing start | Marketing end | Status |
|---|---|---|---|---|
| 00469-2760-28 You're viewing this Main listing | 4 CARTON in 1 TRAY / 1 BLISTER PACK in 1 CARTON / 7 TABLET, FILM COATED in 1 BLISTER PACK Sample | 2023-05-12 | — | Active |
Therapeutic equivalents
| Product | Labeler | Pack | NADAC/unit | TE | Status | Price vs. this |
|---|---|---|---|---|---|---|
| Veozah 45 mg 00469-2660-30 | Astellas | 30 tablets | $18.582 | — | Availability likely | — |
| Veozah 45 mg 00469-2460-28 | Astellas | 7 tablets | — | — | FDA listed | — |
| Veozah 45 mgthis 00469-2760-28 | Astellas | 7 tablets | — | — | FDA listed | — |
Where does this data come from?
- FDA openFDA NDC Directory · synced Oct 1, 2026
- FDA Orange Book · refreshed Oct 3, 2026
- CMS NADAC weekly file
Availability & generic status
We did not find an FDA-approved generic match for this exact strength, form and route. Patent/protection dates below may affect future generic timing.
Why the date isn’t exact: Generic timing can change because patents may be challenged, settled, licensed, added, removed, or worked around with a narrower label — and FDA approval does not always mean a pharmacy can get the generic today.
🛈 What do these terms mean?
- Patent
- Legal protection listed in the Orange Book that may delay generic approval or launch. Issued by the U.S. Patent & Trademark Office.
- Substance patent
- Covers the active drug molecule itself — the hardest to design around. A generic generally can’t launch until it expires.
- Formulation (product) patent
- Covers a specific formulation or dosage form. A generic can sometimes work around it with a different formulation.
- Method-of-use patent
- A patent covering one specific approved use of the drug — not necessarily the whole molecule. A generic can sometimes launch with a “skinny label” that carves out the protected use and keeps the others.
- Skinny label
- A generic label that omits a still-patented use when the FDA allows it — letting a generic reach the market for the unprotected uses.
- Exclusivity
- FDA-granted marketing protection, separate from patents — e.g. 5-yr new chemical entity, 7-yr orphan drug, or a +6-month pediatric extension.
- Paragraph IV
- A generic applicant’s formal challenge to a listed patent. It can potentially lead to earlier generic entry, but often involves litigation or a settlement.
- RLD / RS
- Reference Listed Drug — the brand product the FDA uses as the reference for generic applications. Reference Standard — the product the FDA expects generics to compare against in bioequivalence testing.
- TE / AB rating
- FDA therapeutic-equivalence rating. An AB rating generally means the FDA considers a generic therapeutically equivalent to — and substitutable for — the brand.
- LOE (loss of exclusivity)
- The latest patent or exclusivity currently listed — the loss-of-exclusivity / latest-listed-protection date shown on this page. Paragraph-IV challenges and settlements can move the real date earlier; FDA approval and a manufacturer’s decision to market can move it later.
Built from the FDA Orange Book. The bars above are scaled to each protection’s expiry; the red LOE marker is the last one to lapse.
| Patent | Type | Use code | Expires |
|---|---|---|---|
| US 10836768 ↗ | Method of use | U-3622 | Mar 28, 2034 |
| US 9987274 ↗ | Method of use | U-3622 | Mar 28, 2034 |
| US 9422299 ↗ | Drug substance | U-3622 | Mar 28, 2034 |
| US 8871761 ↗ | Drug substance | — | Apr 4, 2031 |
| Code | What it grants | Expires |
|---|---|---|
| NCE | New Chemical Entity (5-year) | May 12, 2028 |
Is there a generic version of VEOZAH 45 MG TABLET?
The FDA approved a generic — why can’t I get it at my pharmacy yet?
Why do different websites show different generic release dates?
What does “FDA listed” mean?
What does a patent or protection date mean here?
What does “current Orange Book estimate” mean?
Can a generic come out before the last patent expires?
Can a generic come out after the listed dates?
What is the difference between patents and exclusivity?
Why are there multiple patent dates?
Where does this data come from?
- FDA Orange Book · refreshed Oct 3, 2026
What it looks like
Where does this data come from?
- FDA label on DailyMed · label index refreshed Oct 3, 2026
Inactive Ingredients / Excipients
Inactive ingredients, also called excipients, are components of the drug product other than the active ingredient. They may include fillers, dyes, coatings, preservatives, flavors, or other formulation ingredients.
💡 Tap an ingredient (hover on desktop) to see what it is and why it’s used.
-
UNII 1K09F3G675
Ferric oxide red is an inorganic iron compound used as a colorant in medicines. It gives tablets, capsules, or other dosage forms a red or reddish tint for identification and appearance.
-
UNII 9XZ8H6N6OH
A plant-based cellulose derivative used as a binder to hold tablet ingredients together, a thickener in liquids, and a coating agent to control how fast the medicine dissolves.
-
UNII 0WZ8WG20P6
Hypromellose 2910 is a plant-based cellulose derivative that acts as a thickener, binder, and film-coating agent. It helps control how quickly the medicine dissolves and protects the tablet or capsule from moisture and light.
-
UNII 2165RE0K14
A plant-based thickener made from cellulose that helps control how quickly the medicine dissolves and releases its active ingredient. It also binds ingredients together and improves the tablet's texture and handling during manufacturing.
-
UNII 70097M6I30
Magnesium stearate is a salt made from magnesium and stearic acid, a fatty substance. It's used in tablets and capsules as a lubricant and glidant to help ingredients flow smoothly during manufacturing and prevent sticking.
-
UNII 3OWL53L36A
A natural sugar alcohol derived from seaweed or synthesized in the lab. It's used as a filler to add bulk, a sweetener in sugar-free formulas, and a disintegrant to help tablets break apart in the stomach.
-
UNII OP1R32D61U
Microcrystalline cellulose is a purified form of cellulose, a natural fiber from plant sources. It acts as a binder and filler in tablets and capsules, helping hold ingredients together and give the medicine its shape and size.
-
UNII Q662QK8M3B
Polyethylene glycol 8000 is a synthetic polymer made from ethylene oxide. It acts as a binder and filler in tablets and capsules, helping hold ingredients together and add bulk to the medicine.
-
UNII 7SEV7J4R1U
A powder made from a naturally occurring mineral. In medicines, talc works as a glidant and anti-caking agent, helping tablets and capsules flow smoothly during manufacturing and preventing clumping.
-
UNII 15FIX9V2JP
Titanium dioxide is a bright white mineral powder commonly used as a colorant and opacifying agent. It makes pills and tablets white or lighter in color and helps make coatings non-transparent.
10 inactive ingredients listed in the exact product block matched to this NDC.
Where does this data come from?
ingredient classCode="IACT" elements from the exact product block matched by this NDC. Label-section narrative from DailyMed / the openFDA label index is shown separately when available.- FDA label on DailyMed · label index refreshed Oct 3, 2026
- FDA openFDA NDC Directory · synced Oct 1, 2026
Inactive ingredient FAQ
Are inactive ingredients the same for every manufacturer?
Why might an inactive ingredient be missing?
Can inactive ingredients matter?
Manufacturer & labeler
More NDCs from Astellas Pharma US, Inc. labeler code 00469
- Xtandi enzalutamide 40 mg Tablet NDC 0469-1725-56
- Veozah fezolinetant 45 mg Tablet, Film Coated NDC 0469-2460-28
- Myrbetriq mirabegron 25 mg Tablet, Film Coated, Extended Release NDC 0469-2601-07
- Myrbetriq mirabegron 50 mg Tablet, Film Coated, Extended Release NDC 0469-2602-07
- mirabegron 1 kg/kg Powder NDC 0469-2603-01
- Veozah fezolinetant 45 mg Tablet, Film Coated NDC 0469-2660-30
- Cresemba isavuconazonium sulfate 40 mg Capsule NDC 0469-2860-35
- Prograf Tacrolimus 5 mg/mL Injection, Solution NDC 0469-3016-01
- AmBisome amphotericin B 50 mg/12.5mL Injection, Powder, Lyophilized, For Solution NDC 0469-3051-30
- Mycamine Micafungin sodium 10 mg/mL Injection, Powder, Lyophilized, For Solution NDC 0469-3250-10
- VYLOY ZOLBETUXIMAB 20 mg/mL Injection, Powder, For Suspension NDC 0469-3425-10
- VYLOY ZOLBETUXIMAB 20 mg/mL Injection, Powder, For Suspension NDC 0469-4425-30
Where does this data come from?
- FDA openFDA NDC Directory · synced Oct 1, 2026
- Drugs@FDA
Full prescribing information FDA SPL
🚨 Boxed Warning ▾
WARNING: RISKS OF HEPATOTOXICITY Hepatotoxicity has occurred with the use of VEOZAH in the postmarketing setting ( 5.1 ). • Perform hepatic laboratory tests prior to initiation of treatment to evaluate for hepatic function and injury. Do not start VEOZAH if either aminotransferase is ≥ 2 x the upper limit of normal (ULN) or if the total bilirubin is ≥ 2 x ULN for the evaluating laboratory. • Perform follow-up hepatic laboratory testing monthly for the first 3 months, at 6 months, and 9 months of treatment ( 2.1 , 5.1 ). • Advise patients to discontinue VEOZAH immediately and seek medical attention including hepatic laboratory tests if they experience signs or symptoms that may suggest liver injury (new onset fatigue, decreased appetite, nausea, vomiting, pruritus, jaundice, pale feces, dark urine, or abdominal pain) ( 2.1 , 5.1 ). • Discontinue VEOZAH if transaminase elevations are > 5 x ULN, or if transaminase elevations are > 3 x ULN and the total bilirubin level is > 2 x ULN. • If transaminase elevations > 3 x ULN occur, perform more frequent follow-up hepatic laboratory tests until resolution ( 5.1 ).
WARNING: RISKS OF HEPATOTOXICITY See full prescribing information for complete boxed warning. Hepatotoxicity has occurred with the use of VEOZAH in the postmarketing setting ( 5.1 ). • Perform hepatic laboratory tests prior to initiation of treatment to evaluate for hepatic function and injury. Do not start VEOZAH if either aminotransferase is ≥ 2 x ULN or if the total bilirubin is ≥ 2 x ULN for the evaluating laboratory. • Perform follow-up hepatic laboratory testing monthly for the first 3 months, at 6 months, and 9 months of treatment ( 2.1 , 5.1 ). • Advise patients to discontinue VEOZAH immediately and seek medical attention including hepatic laboratory tests if they experience signs or symptoms that may suggest liver injury (new onset fatigue, decreased appetite, nausea, vomiting, pruritus, jaundice, pale feces, dark urine, or abdominal pain) ( 2.1 , 5.1 ). • Discontinue VEOZAH if transaminase elevations are > 5 x ULN, or if transaminase elevations are > 3 x ULN and the total bilirubin level is > 2 x ULN. • If transaminase elevations > 3 x ULN occur, perform more frequent follow-up hepatic laboratory tests until resolution ( 5.1 ).
🎯 Indications and Usage ▾
1 INDICATIONS AND USAGE VEOZAH ® is indicated for the treatment of moderate to severe vasomotor symptoms due to menopause. VEOZAH is a neurokinin 3 (NK3) receptor antagonist indicated for the treatment of moderate to severe vasomotor symptoms due to menopause. ( 1 )
⏱️ Dosage and Administration ▾
2 DOSAGE AND ADMINISTRATION One 45 mg tablet orally once daily with or without food. Perform baseline hepatic laboratory tests to evaluate for hepatic function and injury before beginning VEOZAH. While using VEOZAH, perform follow-up hepatic laboratory tests monthly for the first 3 months, at 6 months, and 9 months after initiation of therapy or when signs or symptoms suggest liver injury. ( 2.1 )
2.1Recommended Dosage Take a single 45 mg VEOZAH tablet orally once daily with or without food. Take VEOZAH with liquids and swallow whole. Do not cut, crush, or chew tablets.
Administer VEOZAH orally at about the same time each day. If a dose of VEOZAH is missed or not taken at the usual time, administer the missed dose as soon as possible, unless there is less than 12 hours before the next scheduled dose. Return to the regular schedule the following day.
Perform baseline hepatic laboratory tests to evaluate for hepatic function and injury [including serum alanine aminotransferase (ALT), serum aspartate aminotransferase (AST), serum alkaline phosphatase (ALP), and serum bilirubin (total and direct)] before initiating treatment with VEOZAH. Do not start VEOZAH if ALT or AST is ≥ 2 x ULN or if the total bilirubin is ≥ 2 x ULN for the evaluating laboratory. While using VEOZAH, perform follow-up hepatic laboratory tests monthly for the first 3 months, at 6 months, and 9 months after initiation of therapy.
Advise patients to discontinue VEOZAH immediately and seek medical attention including hepatic laboratory tests if they experience signs or symptoms that may suggest liver injury [see Warnings and Precautions ( 5.1 )].
💊 Dosage Forms and Strengths ▾
3 DOSAGE FORMS AND STRENGTHS Tablets: 45 mg, round, light red, film-coated tablets, debossed with the Astellas logo and ‘645’ on the same side. Tablets: 45 mg ( 3 )
⛔ Contraindications ▾
4 CONTRAINDICATIONS VEOZAH is contraindicated in women with any of the following conditions: • Known cirrhosis [see Warnings and Precautions ( 5.1 ), Use in Specific Populations ( 8.7 ), and Clinical Pharmacology ( 12.3 )] . • Severe renal impairment or end-stage renal disease [see Use in Specific Populations ( 8.6 ) and Clinical Pharmacology ( 12.3 )] . • Concomitant use with CYP1A2 inhibitors [see Drug Interactions ( 7.1 ) and Clinical Pharmacology ( 12.3 )] . • Known cirrhosis ( 4 , 5.1 ) • Severe renal impairment or end-stage renal disease ( 4 , 8.6 ) • Concomitant use with CYP1A2 inhibitors ( 4 , 7.1 )
⚠️ Warnings and Cautions ▾
5 WARNINGS AND PRECAUTIONS Hepatotoxicity: Cases of hepatotoxicity and jaundice have been reported in the postmarketing setting. Perform hepatic laboratory tests prior to initiation of VEOZAH to evaluate for hepatic function and injury. Perform follow-up hepatic laboratory tests monthly for the first 3 months, at 6 months, and 9 months after initiation of therapy.
Advise patients to discontinue VEOZAH immediately and seek medical attention including hepatic laboratory tests if they experience signs or symptoms that may suggest liver injury (new onset fatigue, decreased appetite, nausea, vomiting, pruritus, jaundice, pale feces, dark urine, or abdominal pain). ( 5.1 )
5.1Hepatotoxicity In three clinical trials, elevations in serum transaminase [alanine aminotransferase (ALT) and/or aspartate aminotransferase (AST)] levels > 3 x the upper limit of normal (ULN) occurred in 2.3% [exposure adjusted incidence rate (EAIR) of 2.7 per 100 person-years] of women receiving VEOZAH and 0.9% (EAIR of 1.5 per 100 person-years) of women receiving placebo. No elevations in serum total bilirubin (> 2 x ULN) occurred. Women with ALT or AST elevations were generally asymptomatic.
Transaminase levels returned to pretreatment levels (or close to these) without sequelae with dose continuation, and upon dose interruption, or discontinuation. Women with cirrhosis were not studied [see Adverse Reactions ( 6.1 )] . In the postmarketing setting, cases of drug-induced liver injury with elevations of ALT, AST, alkaline phosphatase (ALP), and total bilirubin occurred within 40 days of starting VEOZAH.
Patients reported a general sense of feeling unwell and symptoms of fatigue, nausea, pruritus, jaundice, pale feces, and dark urine. The patients’ signs and symptoms gradually resolved after discontinuation of VEOZAH [see Adverse Reactions ( 6.2 )]. Perform baseline hepatic laboratory tests to evaluate for hepatic function and injury [including serum ALT, serum AST, serum ALP, and serum bilirubin (total and direct)] prior to VEOZAH initiation.
Do not start VEOZAH if ALT or AST is ≥ 2 x ULN or if the total bilirubin is ≥ 2 x ULN for the evaluating laboratory. Perform follow-up hepatic laboratory tests monthly for the first 3 months, at 6 months, and 9 months after initiation of therapy. Advise patients to discontinue VEOZAH immediately and seek medical attention including hepatic laboratory tests if they experience signs or symptoms that may suggest liver injury: • new onset fatigue, decreased appetite, nausea, vomiting, pruritus, jaundice, pale feces, dark urine, or abdominal pain.
Discontinue VEOZAH if: • transaminase elevations are > 5 x ULN. • transaminase elevations are > 3 x ULN and total bilirubin is > 2 x ULN. If transaminase elevations > 3 x ULN occur, perform more frequent follow-up hepatic laboratory tests until resolution. Exclude alternative causes of hepatic laboratory test elevations.
🤒 Adverse Reactions ▾
6 ADVERSE REACTIONS The following serious adverse reactions are discussed elsewhere in the labeling: • Hepatic Transaminase Elevation and Hepatotoxicity [see Warnings and Precautions ( 5.1 )] . The most common adverse reactions with VEOZAH [at least 2% in VEOZAH 45 mg and greater than placebo] are: abdominal pain, diarrhea, insomnia, back pain, hot flush, and hepatic transaminase elevation. ( 6.1 ) To report SUSPECTED ADVERSE REACTIONS, contact Astellas Pharma US, Inc. at 1-800-727-7003 or FDA at 1-800-FDA-1088 or www.fda.gov/medwatch .
6.1Clinical Trials Experience Because clinical trials are conducted under widely varying conditions, adverse reaction rates observed in the clinical trials of a drug cannot be directly compared to rates in the clinical trials of another drug and may not reflect the rates observed in clinical practice. The safety of VEOZAH was evaluated in three 52-week clinical trials [see Clinical Studies ( 14 )] . Across the three clinical trials, a total of 1100 women received VEOZAH.
Trials 1 and 2 were placebo-controlled for the first 12 weeks, followed by re-randomization of women previously receiving placebo to VEOZAH (women on VEOZAH remained on VEOZAH) for an additional 40 weeks of uncontrolled treatment. Trial 3 was a randomized, placebo-controlled, double-blind safety study evaluating the safety of VEOZAH for 52 weeks. The adverse reactions reported in at least 2% in VEOZAH 45 mg and greater than placebo in Trial 3 are presented in Table 1 .
Table 1: Adverse Reactions Reported in at Least 2% in VEOZAH 45 mg and Greater Than Placebo in a Placebo‑Controlled, Double-Blind 52-Week Trial (Trial 3) Adverse Reaction VEOZAH 45 mg (n=609) Total Person-Years=504.2 n (%, EAIR EAIR = Number of individuals experiencing an adverse event divided by exposure time (total person-years) x 100. ) Placebo (n=610) Total Person-Years=475.0 n (%, EAIR ) Abdominal pain Abdominal pain (including Abdominal pain, Abdominal pain lower, Abdominal pain upper). 26 (4.3%, 5.2) 13 (2.1%, 2.7) Diarrhea 24 (3.9%, 4.8) 16 (2.6%, 3.4) Insomnia 24 (3.9%, 4.8) 11 (1.8%, 2.3) Back pain 18 (3.0%, 3.6) 13 (2.1%, 2.7) Hot flush 15 (2.5%, 3.0) 10 (1.6%, 2.1) Hepatic transaminase elevation Hepatic transaminase elevations (including Alanine aminotransferase abnormal, Alanine aminotransferase increased, Aspartate aminotransferase abnormal, Aspartate aminotransferase increased).
14 (2.3%, 2.8) 5 (0.8%, 1.1) In the pooled laboratory data of Trials 1, 2, and 3, elevated hepatic transaminases (> 3 x ULN) occurred in 25 women (2.3%,
2.7EAIR) exposed to VEOZAH 45 mg (n=1100, 912.1 total person-years) as compared to 8 women (0.9%,
1.5EAIR) exposed to placebo (n=952, 549.1 total person-years).
6.2Postmarketing Experience The following adverse reactions have been identified during postapproval use of VEOZAH. Because these reactions are reported voluntarily from a population of uncertain size, it is not always possible to reliably estimate their frequency or establish a causal relationship to drug exposure. Hepatic : Cases of serious drug-induced hepatotoxicity occurred within 40 days of starting VEOZAH.
Patients experienced elevated transaminases (up to 50 x ULN at peak elevation), elevated alkaline phosphatase (up to 4 x ULN at peak elevation), and bilirubin (up to 5 x ULN at peak elevation) coupled with symptoms of fatigue, nausea, pruritus, jaundice, pale feces, and dark urine. After discontinuation of VEOZAH, these abnormalities gradually resolved.
🔄 Drug Interactions ▾
7 DRUG INTERACTIONS
7.1Effect of Other Drugs on VEOZAH CYP1A2 Inhibitors VEOZAH is a substrate of CYP1A2. Concomitant use of VEOZAH with drugs that are weak, moderate, or strong CYP1A2 inhibitors, increase the plasma C max and AUC of VEOZAH [see Clinical Pharmacology ( 12.3 )] . VEOZAH is contraindicated in individuals using CYP1A2 inhibitors.
👥 Use in Specific Populations ▾
8 USE IN SPECIFIC POPULATIONS
8.1Pregnancy Risk Summary There are no data on VEOZAH use in pregnant women to evaluate for a drug-associated risk of major birth defects, miscarriage, or adverse maternal or fetal outcomes. In embryo-fetal toxicity animal studies with fezolinetant, embryo-lethality occurred at high doses above the human therapeutic dose in rats and rabbits, but no teratogenicity was observed. In the pre- and post-natal development animal study, delayed parturition and embryo-lethality occurred at high doses above the human therapeutic dose in rats.
Additionally, in the male offspring delayed male reproductive maturation was observed, characterized by incomplete preputial separation, which affected male fertility at doses above the human therapeutic dose in rats [see Data ] . In the U.S. general population, the estimated background risk of major birth defects or miscarriage in clinically recognized pregnancies are 2-4% and 15-20%, respectively. Data Animal Data In embryo-fetal development toxicity studies in rats and rabbits, embryo-lethality was noted at the highest doses (128- and 174-fold the human AUC 24 at the human therapeutic dose for rats and rabbits, respectively).
The no observed adverse effect level (NOAEL) for embryo-fetal development was 50 mg/kg/day in rats and 45 mg/kg/day in rabbits (62- and 16‑fold the human AUC 24 at the human therapeutic dose for rats and rabbits, respectively). Fezolinetant showed no effects on fertility and early embryonic development in rats [see Nonclinical Toxicology ( 13.1 )] . In the pre- and post-natal development study in rats, the NOAEL for maternal and fetal toxicity was 30 mg/kg/day (36‑fold the human AUC 24 at the human therapeutic dose) based on delayed parturition and embryo-lethality at 100 mg/kg/day.
The NOAEL for F 1 generation development was determined to be 100 mg/kg/day for females (204-fold the human AUC 24 at the human therapeutic dose) and 10 mg/kg/day for males (11-fold the human AUC 24 at the human therapeutic dose). The F 1 male showed delayed male reproductive maturation, characterized as incomplete balanopreputial separation at time of mating, at doses of greater than or equal to 30 mg/kg/day (36-fold the human AUC 24 at the human therapeutic dose), which affected male fertility [see Nonclinical Toxicology ( 13.1 )] .
8.2Lactation Risk Summary There are no data on the presence of fezolinetant in human milk, the effects on the breastfed child, or the effects on milk production. It is not known if fezolinetant is present in human milk. Data Animal Data Following administration of radiolabeled fezolinetant to lactating rats, the radioactivity concentration in milk was higher than that in the plasma at all time points, indicating that fezolinetant-derived components transferred to the tissues in infant rats via breast milk.
8.4Pediatric Use The efficacy and safety of VEOZAH in individuals less than 18 years of age have not been established.
8.5Geriatric Use There have not been sufficient numbers of geriatric women involved in clinical trials utilizing VEOZAH to determine whether those over 65 years of age differ from younger women in their response to VEOZAH.
8.6Renal Impairment VEOZAH is contraindicated in individuals with severe (eGFR 15 to less than 30 mL/min/1.73 m 2 ) renal impairment or end-stage renal disease (eGFR less than 15 mL/min/1.73 m 2 ) [see Clinical Pharmacology ( 12.3 )] . No dose adjustment of VEOZAH is recommended for individuals with mild (eGFR 60 to less than 90 mL/min/1.73 m 2 ) or moderate (eGFR 30 to less than 60 mL/min/1.73 m 2 ) renal impairment.
8.7Hepatic Impairment Child-Pugh Class A or B hepatic impairment increased the exposure of VEOZAH [see Clinical Pharmacology ( 12.3 )] . VEOZAH has not been studied in individuals with Child-Pugh Class C hepatic impairment. VEOZAH is contraindicated in individuals with cirrhosis [see Clinical Pharmacology ( 12.3 )] .
🤰 Pregnancy ▾
8.1Pregnancy Risk Summary There are no data on VEOZAH use in pregnant women to evaluate for a drug-associated risk of major birth defects, miscarriage, or adverse maternal or fetal outcomes. In embryo-fetal toxicity animal studies with fezolinetant, embryo-lethality occurred at high doses above the human therapeutic dose in rats and rabbits, but no teratogenicity was observed. In the pre- and post-natal development animal study, delayed parturition and embryo-lethality occurred at high doses above the human therapeutic dose in rats.
Additionally, in the male offspring delayed male reproductive maturation was observed, characterized by incomplete preputial separation, which affected male fertility at doses above the human therapeutic dose in rats [see Data ] . In the U.S. general population, the estimated background risk of major birth defects or miscarriage in clinically recognized pregnancies are 2-4% and 15-20%, respectively. Data Animal Data In embryo-fetal development toxicity studies in rats and rabbits, embryo-lethality was noted at the highest doses (128- and 174-fold the human AUC 24 at the human therapeutic dose for rats and rabbits, respectively).
The no observed adverse effect level (NOAEL) for embryo-fetal development was 50 mg/kg/day in rats and 45 mg/kg/day in rabbits (62- and 16‑fold the human AUC 24 at the human therapeutic dose for rats and rabbits, respectively). Fezolinetant showed no effects on fertility and early embryonic development in rats [see Nonclinical Toxicology ( 13.1 )] . In the pre- and post-natal development study in rats, the NOAEL for maternal and fetal toxicity was 30 mg/kg/day (36‑fold the human AUC 24 at the human therapeutic dose) based on delayed parturition and embryo-lethality at 100 mg/kg/day.
The NOAEL for F 1 generation development was determined to be 100 mg/kg/day for females (204-fold the human AUC 24 at the human therapeutic dose) and 10 mg/kg/day for males (11-fold the human AUC 24 at the human therapeutic dose). The F 1 male showed delayed male reproductive maturation, characterized as incomplete balanopreputial separation at time of mating, at doses of greater than or equal to 30 mg/kg/day (36-fold the human AUC 24 at the human therapeutic dose), which affected male fertility [see Nonclinical Toxicology ( 13.1 )] .
🧒 Pediatric Use ▾
8.4Pediatric Use The efficacy and safety of VEOZAH in individuals less than 18 years of age have not been established.
🧓 Geriatric Use ▾
8.5Geriatric Use There have not been sufficient numbers of geriatric women involved in clinical trials utilizing VEOZAH to determine whether those over 65 years of age differ from younger women in their response to VEOZAH.
🆘 Overdosage ▾
10 OVERDOSAGE Treatment of overdose consists of discontinuation of VEOZAH therapy with institution of appropriate symptomatic care.
🧬 Clinical Pharmacology ▾
12 CLINICAL PHARMACOLOGY
12.1Mechanism of Action VEOZAH is a neurokinin 3 (NK3) receptor antagonist that blocks neurokinin B (NKB) binding on the kisspeptin/neurokinin B/dynorphin (KNDy) neuron to modulate neuronal activity in the thermoregulatory center. Fezolinetant has high affinity for the NK3 receptor (Ki value of 19.9 to 22.1 nmol/L), which is more than 450-fold higher than binding affinity to NK1 or NK2 receptors.
12.2Pharmacodynamics Treatment with fezolinetant did not show any clear trends in sex hormones measured (follicle-stimulating hormone, testosterone, estrogen, and dehydroepiandrosterone sulfate) in menopausal women. Transient decrease of luteinizing hormone (LH) levels was observed at peak concentrations of fezolinetant. Cardiac Electrophysiology At a dose 20 times the maximum approved recommended dose, fezolinetant does not prolong the QT interval to any clinically relevant extent.
12.3Pharmacokinetics In healthy women, fezolinetant C max and AUC increased proportionally over a dosage range from 20 to 60 mg once daily (0.44 to 1.33 times the approved recommended dosage). Steady-state plasma concentrations of fezolinetant were reached after two once daily doses, with minimal fezolinetant accumulation. Absorption The median (range) time to reach fezolinetant C max is 1.5 (1 to 4) hours in healthy women.
Effect of Food No clinically significant differences in fezolinetant pharmacokinetics were observed following administration with a high‑calorie, high-fat meal containing approximately 1000 calories (500-600 calories from fat, 250 calories from carbohydrates, and 150 calories from protein). Distribution The mean apparent volume of distribution (V z /F) of fezolinetant is 189 L. The plasma protein binding of fezolinetant is 51%.
The blood-to-plasma ratio is 0.9. Elimination The effective half-life (t 1/2 ) of fezolinetant is 9.6 hours in women with vasomotor symptoms. The apparent clearance at steady-state of fezolinetant is
10.8L/h. Metabolism Fezolinetant is primarily metabolized by CYP1A2 and to a lesser extent by CYP2C9 and CYP2C19. A major metabolite of fezolinetant, ES259564, was identified in plasma.
ES259564 is approximately 20-fold less potent than the parent. The metabolite-to-parent ratio ranges from 0.7 to 1.8. Excretion Following oral administration of fezolinetant, 76.9% of the dose was excreted in urine (1.1% unchanged) and 14.7% in feces (0.1% unchanged).
Specific Populations There were no substantive differences in the pharmacokinetics of VEOZAH based on race and body weight (93 to 278 pounds). Women with Renal Impairment Following single-dose administration of 30 mg fezolinetant, there was no effect on VEOZAH exposure (C max and AUC) in women with mild (eGFR 60 to less than 90 mL/min/1.73 m 2 ) to severe (eGFR 15 to less than 30 mL/min/1.73 m 2 ) renal impairment. The AUC of ES259564 (a major metabolite of fezolinetant) in women with moderate (eGFR 30 to less than 60 mL/min/1.73 m 2 ) and severe (eGFR 15 to less than 30 mL/min/1.73 m 2 ) renal impairment increased by approximately 75% and 380%, respectively.
VEOZAH has not been studied in individuals with end-stage renal disease (eGFR less than 15 mL/min/1.73 m 2 ). Women with Hepatic Impairment Following single-dose administration of 30 mg fezolinetant in women with mild Child-Pugh Class A cirrhosis, the mean C max increased by 23% and AUC inf increased by 56%, relative to women with normal hepatic function. In women with moderate Child-Pugh Class B cirrhosis, the mean C max of fezolinetant decreased by 15% and AUC inf increased by 96%.
The C max of ES259564 decreased in both mild and moderate cirrhosis while AUC inf and AUC last increased less than 15%. VEOZAH has not been studied in individuals with severe Child-Pugh Class C cirrhosis. Drug Interaction Studies Clinical Studies and Model-Informed Approaches Strong CYP1A2 Inhibitors: Fezolinetant C max increased by 80% and AUC increased by 840% following concomitant use with fluvoxamine (st… [Excerpted — this section continues on DailyMed.]
🧬 Mechanism of Action ▾
12.1Mechanism of Action VEOZAH is a neurokinin 3 (NK3) receptor antagonist that blocks neurokinin B (NKB) binding on the kisspeptin/neurokinin B/dynorphin (KNDy) neuron to modulate neuronal activity in the thermoregulatory center. Fezolinetant has high affinity for the NK3 receptor (Ki value of 19.9 to 22.1 nmol/L), which is more than 450-fold higher than binding affinity to NK1 or NK2 receptors.
📦 How Supplied / Storage and Handling ▾
16 HOW SUPPLIED/STORAGE AND HANDLING
16.1How Supplied VEOZAH (fezolinetant) 45 mg tablets are supplied as round, light red, film-coated tablets, debossed with the Astellas logo and ‘645’ on the same side. VEOZAH tablets are available in the following package sizes: • Bottles of 30 tablets with child resistant closure, (NDC 0469-2660-30) • Bottles of 90 tablets with child resistant closure, (NDC 0469-2660-90)
16.2Storage and Handling Store at 20°C to 25°C (68°F to 77°F) with excursions permitted from 15°C to 30°C (59°F to 86°F) [see USP Controlled Room Temperature].
📋 Description ▾
11 DESCRIPTION VEOZAH (fezolinetant) is a small-molecule NK3 receptor antagonist. The chemical name of fezolinetant is (4-Fluorophenyl)[(8 R )-8-methyl-3-(3-methyl-1,2,4-thiadiazol-5-yl)-5,6-dihydro[1,2,4]triazolo[4,3- a ]pyrazin-7(8 H )-yl]methanone having a molecular formula of C 16 H 15 FN 6 OS and a molecular weight of 358.39. The structural formula of fezolinetant is: Fezolinetant is a white powder.
It is very slightly soluble in water (0.29 mg/mL). Each VEOZAH (fezolinetant) tablet for oral use contains 45 mg of fezolinetant and the following inactive ingredients: ferric oxide, hydroxypropyl cellulose, hypromellose, low-substituted hydroxypropyl cellulose, magnesium stearate, mannitol, microcrystalline cellulose, polyethylene glycol, talc, and titanium dioxide. structural formula of fezolinetant
💬 Information for Patients ▾
17 PATIENT COUNSELING INFORMATION Advise patients to read the FDA-approved patient labeling (Patient Information). Evaluation of Hepatic Injury During Treatment with VEOZAH Inform patients that they will have to have a blood test to evaluate their liver function before beginning VEOZAH and while using VEOZAH monthly for the first 3 months, at 6 months, and 9 months after initialization of therapy. Advise patients to discontinue VEOZAH immediately and seek medical attention including hepatic laboratory tests if they experience signs or symptoms that may suggest liver abnormalities such as new onset fatigue, decreased appetite, nausea, vomiting, pruritus, jaundice, pale feces, dark urine, or abdominal pain [see Warnings and Precautions ( 5.1 )] .
Serious Adverse Reactions with VEOZAH Inform patients of possible serious adverse reactions of VEOZAH including hepatic transaminase elevation and liver injury [see Warnings and Precautions ( 5.1 )] . Common Adverse Reactions with VEOZAH Inform patients of possible less serious but common adverse reactions of VEOZAH including abdominal pain, diarrhea, insomnia, back pain, and hot flush [see Adverse Reactions ( 6.1 )] . Drug Interactions Advise patients to report their use of any other prescription or nonprescription medications or dietary supplements [see Drug Interactions ( 7.1 )].
Distributed by: Astellas Pharma US, Inc. Northbrook, IL 60062 © 2024 Astellas Pharma US, Inc. or its affiliates 432286-VEO-USA
🧬 Pharmacokinetics ▾
12.3Pharmacokinetics In healthy women, fezolinetant C max and AUC increased proportionally over a dosage range from 20 to 60 mg once daily (0.44 to 1.33 times the approved recommended dosage). Steady-state plasma concentrations of fezolinetant were reached after two once daily doses, with minimal fezolinetant accumulation. Absorption The median (range) time to reach fezolinetant C max is 1.5 (1 to 4) hours in healthy women.
Effect of Food No clinically significant differences in fezolinetant pharmacokinetics were observed following administration with a high‑calorie, high-fat meal containing approximately 1000 calories (500-600 calories from fat, 250 calories from carbohydrates, and 150 calories from protein). Distribution The mean apparent volume of distribution (V z /F) of fezolinetant is 189 L. The plasma protein binding of fezolinetant is 51%.
The blood-to-plasma ratio is 0.9. Elimination The effective half-life (t 1/2 ) of fezolinetant is 9.6 hours in women with vasomotor symptoms. The apparent clearance at steady-state of fezolinetant is
10.8L/h. Metabolism Fezolinetant is primarily metabolized by CYP1A2 and to a lesser extent by CYP2C9 and CYP2C19. A major metabolite of fezolinetant, ES259564, was identified in plasma.
ES259564 is approximately 20-fold less potent than the parent. The metabolite-to-parent ratio ranges from 0.7 to 1.8. Excretion Following oral administration of fezolinetant, 76.9% of the dose was excreted in urine (1.1% unchanged) and 14.7% in feces (0.1% unchanged).
Specific Populations There were no substantive differences in the pharmacokinetics of VEOZAH based on race and body weight (93 to 278 pounds). Women with Renal Impairment Following single-dose administration of 30 mg fezolinetant, there was no effect on VEOZAH exposure (C max and AUC) in women with mild (eGFR 60 to less than 90 mL/min/1.73 m 2 ) to severe (eGFR 15 to less than 30 mL/min/1.73 m 2 ) renal impairment. The AUC of ES259564 (a major metabolite of fezolinetant) in women with moderate (eGFR 30 to less than 60 mL/min/1.73 m 2 ) and severe (eGFR 15 to less than 30 mL/min/1.73 m 2 ) renal impairment increased by approximately 75% and 380%, respectively.
VEOZAH has not been studied in individuals with end-stage renal disease (eGFR less than 15 mL/min/1.73 m 2 ). Women with Hepatic Impairment Following single-dose administration of 30 mg fezolinetant in women with mild Child-Pugh Class A cirrhosis, the mean C max increased by 23% and AUC inf increased by 56%, relative to women with normal hepatic function. In women with moderate Child-Pugh Class B cirrhosis, the mean C max of fezolinetant decreased by 15% and AUC inf increased by 96%.
The C max of ES259564 decreased in both mild and moderate cirrhosis while AUC inf and AUC last increased less than 15%. VEOZAH has not been studied in individuals with severe Child-Pugh Class C cirrhosis. Drug Interaction Studies Clinical Studies and Model-Informed Approaches Strong CYP1A2 Inhibitors: Fezolinetant C max increased by 80% and AUC increased by 840% following concomitant use with fluvoxamine (strong CYP1A2 inhibitor).
Moderate CYP1A2 Inhibitors: Fezolinetant C max increased by 40% and AUC increased by 360% following concomitant use with mexiletine (moderate CYP1A2 inhibitor) 400 mg every 8 hours. Weak CYP1A2 Inhibitors: Fezolinetant C max increased by 30% and AUC increased by 100% following concomitant use with cimetidine (weak CYP1A2 inhibitor) 300 mg every 6 hours. No clinically significant differences in fezolinetant exposure were observed in smokers (moderate CYP1A2 inducer).
In Vitro Studies Cytochrome P450 (CYP) Enzymes: Fezolinetant and ES259564 are not inhibitors of CYP1A2, CYP2B6, CYP2C8, CYP2C9, CYP2C19, CYP2D6, and CYP3A4. Fezolinetant and ES259564 are not inducers of CYP1A2, CYP2B6, and CYP3A4. Transporter Systems Fezolinetant is not a substrate nor an inhibitor of P-glycoprotein (P-gp).
ES259564 is a substrate of P-gp, but not an inhibitor of P-gp. Both fezolinetant and ES259564… [Excerpted — this section continues on DailyMed.]
🧬 Pharmacodynamics ▾
12.2Pharmacodynamics Treatment with fezolinetant did not show any clear trends in sex hormones measured (follicle-stimulating hormone, testosterone, estrogen, and dehydroepiandrosterone sulfate) in menopausal women. Transient decrease of luteinizing hormone (LH) levels was observed at peak concentrations of fezolinetant. Cardiac Electrophysiology At a dose 20 times the maximum approved recommended dose, fezolinetant does not prolong the QT interval to any clinically relevant extent.
🔬 Clinical Studies ▾
14 CLINICAL STUDIES
14.1Effects on Vasomotor Symptoms in Postmenopausal Women The efficacy of VEOZAH for the treatment of moderate to severe vasomotor symptoms due to menopause was evaluated in the first 12-week, randomized, placebo-controlled, double-blind portion of each of two phase 3 clinical trials. In each of these two trials, after the first 12 weeks, women on placebo were then re-randomized to VEOZAH for a 40-week extension to evaluate safety for up to 52 weeks total exposure. In Trials 1 (NCT04003155) and 2 (NCT04003142), 1022 women (522 in Trial 1 and 500 in Trial 2) who had a minimum average of 7 moderate to severe vasomotor symptoms per day were randomized to one of two doses of fezolinetant (including the 45 mg dosage strength) or placebo.
Randomization was stratified by smoking status. The mean age of the postmenopausal women was 54 years. Women self-identified as Caucasian (81%), African American (17%), Asian (1%), and Hispanic/Latina ethnicity (24%).
The study population included menopausal women with one or more of the following: prior hysterectomy (32.1%), prior oophorectomy (21.6%), or prior hormone therapy use (19.9%). Those who were on prior hormone therapy underwent a wash-out period prior to trial participation. The co-primary efficacy endpoints for both trials were the mean change from baseline in moderate to severe vasomotor symptoms frequency and severity to Weeks 4 and 12.
Data from each trial demonstrated statistically significant and clinically meaningful (≥ 2 hot flashes over 24 hours) reduction from baseline in the frequency of moderate to severe vasomotor symptoms for VEOZAH 45 mg compared to placebo at Weeks 4 and 12. Data from each trial also demonstrated a statistically significant reduction from baseline in the severity of moderate to severe vasomotor symptoms (over 24 hours) at Weeks 4 and 12 for VEOZAH 45 mg compared to placebo. Results of the co-primary endpoint for change from baseline to Weeks 4 and 12 in mean frequency of moderate to severe vasomotor symptoms over 24 hours from Trials 1 and 2 are shown in Table 2 .
Table 2: Mean Baseline and Change from Baseline to Weeks 4 and 12 for Mean Frequency of Moderate to Severe Vasomotor Symptoms over 24 Hours in Women Treated with VEOZAH in Trials 1 and 2 Parameter Trial 1 Trial 2 VEOZAH 45 mg (n=174) Placebo (n=175) VEOZAH 45 mg (n=167) Placebo (n=167) Baseline Mean (SD) 10.4 (3.92) 10.5 (3.79) 11.8 (8.26) 11.6 (5.02) Change from Baseline to Week 4 LS Mean (SE) Difference vs Placebo (95% CI) P-value -5.4 (0.30) -2.1 (-2.9, -1.3) < 0.001 Statistically significantly superior compared to placebo at the 0.05 level with multiplicity adjustment.
LS Mean: Least Squares Mean estimated from a mixed model for repeated measures analysis of covariance; SD: Standard Deviation; SE: Standard Error. -3.3 (0.29) -- -- -6.3 (0.33) -2.6 (-3.5, -1.6) < 0.001 -3.7 (0.33) -- -- Change from Baseline to Week 12 LS Mean (SE) Difference vs Placebo (95% CI) P-value -6.4 (0.31) -2.6 (-3.4, -1.7) < 0.001 -3.9 (0.31) -- -- -7.5 (0.39) -2.5 (-3.6, -1.5) < 0.001 -5.0 (0.39) -- -- Results of the co-primary endpoint for change from baseline to Weeks 4 and 12 in mean severity of moderate to severe vasomotor symptoms over 24 hours from Trials 1 and 2 are shown in Table 3 .
Table 3: Mean Baseline and Change from Baseline to Weeks 4 and 12 for Mean Severity of Moderate to Severe Vasomotor Symptoms over 24 Hours in Women Treated with VEOZAH in Trials 1 and 2 Parameter Trial 1 Trial 2 VEOZAH 45 mg (n=174) Placebo (n=175) VEOZAH 45 mg (n=167) Placebo (n=167) Baseline Mean (SD) 2.4 (0.35) 2.4 (0.35) 2.4 (0.34) 2.4 (0.32) Change from Baseline to Week 4 LS Mean (SE) Difference vs Placebo (95% CI) P-value -0.5 (0.04) -0.2 (-0.3, -0.1) 0.002 Statistically significantly superior compared to placebo at the 0.05 level with multiplicity adjustment.
LS Mean: Least Squares Mean estimated from a mixed model for repeated measures analysis of covariance; SD: Standard Deviation; SE: Standard Err… [Excerpted — this section continues on DailyMed.]
🧪 Nonclinical Toxicology ▾
13 NONCLINICAL TOXICOLOGY
13.1Carcinogenesis, Mutagenesis, Impairment of Fertility Carcinogenesis In a 2-year female rat carcinogenicity study and a 26-week carcinogenicity study in rasH2 transgenic mice, there was no evidence of drug-related carcinogenicity at 186-fold and 47-fold the human AUC 24 at the human therapeutic dose of 45 mg, respectively. Mutagenesis Fezolinetant showed no genotoxic potential by the bacterial reverse mutation test, chromosomal aberration test, or in vivo micronucleus test. Impairment of Fertility Fezolinetant had no effect on female fertility or early embryonic development up to 100 mg/kg/day in rats (143-fold the human AUC 24 at the human therapeutic dose).
In the pre- and post-natal development study in rats, the F 1 male showed incomplete balanopreputial separation at doses greater than or equal to 30 mg/kg/day (36-fold the human AUC 24 at the human therapeutic dose), which delayed male reproductive maturation and affected fertility. These effects were not observed following dosing at 10 mg/kg/day (11-fold the human AUC 24 at the human therapeutic dose) [ see Use in Specific Populations ( 8.1 )] .
13.2Animal Toxicology and/or Pharmacology Repeat dose toxicity studies were conducted in intact female rats and cynomolgus monkeys. In female rats, daily administration of fezolinetant for 26 weeks at doses equal to or greater than 30 mg/kg/day (56-fold the human AUC 24 at the human therapeutic dose) showed uterine atrophy and epithelial mucification of the vagina and cervix. In female cynomolgus monkeys, daily administration for 39 weeks at doses equal to or greater than 10 mg/kg/day (19-fold the human AUC 24 at the human therapeutic dose) showed reduced ovarian activity.
📄 Carcinogenesis, Mutagenesis, Impairment of Fertility ▾
13.1Carcinogenesis, Mutagenesis, Impairment of Fertility Carcinogenesis In a 2-year female rat carcinogenicity study and a 26-week carcinogenicity study in rasH2 transgenic mice, there was no evidence of drug-related carcinogenicity at 186-fold and 47-fold the human AUC 24 at the human therapeutic dose of 45 mg, respectively. Mutagenesis Fezolinetant showed no genotoxic potential by the bacterial reverse mutation test, chromosomal aberration test, or in vivo micronucleus test. Impairment of Fertility Fezolinetant had no effect on female fertility or early embryonic development up to 100 mg/kg/day in rats (143-fold the human AUC 24 at the human therapeutic dose).
In the pre- and post-natal development study in rats, the F 1 male showed incomplete balanopreputial separation at doses greater than or equal to 30 mg/kg/day (36-fold the human AUC 24 at the human therapeutic dose), which delayed male reproductive maturation and affected fertility. These effects were not observed following dosing at 10 mg/kg/day (11-fold the human AUC 24 at the human therapeutic dose) [ see Use in Specific Populations ( 8.1 )] .
📄 Patient Package Insert ▾
PATIENT INFORMATION VEOZAH ® (vee-O-zah) (fezolinetant) tablets, for oral use What is the most important information I should know about VEOZAH? VEOZAH can cause serious side effects, including: • Liver Problems. Your healthcare provider will do a blood test to check your liver before you start taking VEOZAH.
Your healthcare provider will also do this blood test monthly for the first 3 months, at month 6, and month 9 after you start taking VEOZAH or if you have signs or symptoms that suggest liver problems. If your liver blood test values are elevated, your healthcare provider may advise you to stop treatment or request additional liver blood tests. Stop VEOZAH right away and call your healthcare provider if you have the following signs or symptoms of liver problems: • feeling more tired than you do usually • decreased appetite • nausea • vomiting • itching • yellowing of the eyes or skin (jaundice) • pale feces • dark urine • pain in the stomach (abdomen) See “ What are the possible side effects of VEOZAH? ” for more information about side effects.
What is VEOZAH? VEOZAH is a prescription medicine used to reduce moderate to severe vasomotor symptoms due to menopause. VEOZAH is not a hormone.
Vasomotor symptoms are the feelings of warmth in the face, neck, and chest, or sudden intense feelings of heat and sweating (“hot flashes” or “hot flushes”). Do not use VEOZAH if you: • have cirrhosis. • have severe kidney problems or kidney failure. • are taking certain medicines called CYP1A2 inhibitors. Ask your healthcare provider if you are not sure.
Before you use VEOZAH, tell your healthcare provider about all of your medical conditions, including if you: • have liver disease or liver problems. • have kidney problems. • have any medical conditions that may become worse while you are using VEOZAH. Tell your healthcare provider about all the medicines you take, including prescription and over-the-counter medicines, vitamins, and herbal supplements. VEOZAH may affect the way other medicines work, and other medicines may affect how VEOZAH works.
Keep a list of your medicines and show it to your healthcare provider and pharmacist when you get a new medicine. How should I take VEOZAH? • Take VEOZAH exactly as your healthcare provider tells you to take it. • Take 1 VEOZAH tablet by mouth with or without food at about the same time each day. • Swallow the VEOZAH tablet whole with liquid. Do not cut, crush, or chew the tablet. • If you miss a dose of VEOZAH, take the missed dose as soon as possible on the same day, with at least 12 hours before the next scheduled dose.
Return to your normal schedule the following day. What are the possible side effects of VEOZAH? VEOZAH can cause serious side effects, including: • See “ What is the most important information I should know about VEOZAH? ” Common side effects of VEOZAH include: • stomach (abdominal) pain • diarrhea • difficulty sleeping (insomnia) • back pain • hot flashes or hot flushes Tell your healthcare provider if you have any side effect that bothers you or does not go away.
These are not all the possible side effects of VEOZAH. Call your doctor for medical advice about side effects. You may report side effects to FDA at 1-800-FDA-1088.
How should I store VEOZAH? • Store VEOZAH at room temperature between 68°F to 77°F (20°C to 25°C). • Dispose of the unused medicine through a take-back option, if available. See www.fda.gov/drugdisposal for more information. • Keep VEOZAH and all medicines out of the reach of children. General information about the safe and effective use of VEOZAH.
Medicines are sometimes prescribed for purposes other than those listed in a Patient Information leaflet. Do not use VEOZAH for a condition for which it was not prescribed. Do not give VEOZAH to other people, even if they have the same symptoms you have.
It may harm them. This Patient Information leaflet summarizes the most important information about VEOZAH. If you would like more information, talk with your… [Excerpted — this section continues on DailyMed.]
📄 Recent Major Changes ▾
Boxed Warning, Hepatotoxicity 12/2024 Dosage and Administration, Recommended Dosage ( 2.1 ) 12/2024 Warnings and Precautions, Hepatotoxicity ( 5.1 ) 12/2024
📄 Package Label / Principal Display Panel ▾
Package/Label Display Panel – VEOZAH Bottle Carton Label NDC 0469-2660-30 Rx Only VEOZAH ® (fezolinetant) tablets 45 mg 30 tablets Package/Label Display Panel- VEOZAH Bottle Carton Label
Package/Label Display Panel – VEOZAH Sample Bottle Carton Label NDC 0469-2460-07 PROFESSIONAL SAMPLE-NOT FOR SALE Rx Only VEOZAH ® (fezolinetant) tablets 45 mg 7 tablets Package/Label Display Panel- VEOZAH Sample Bottle Carton Label
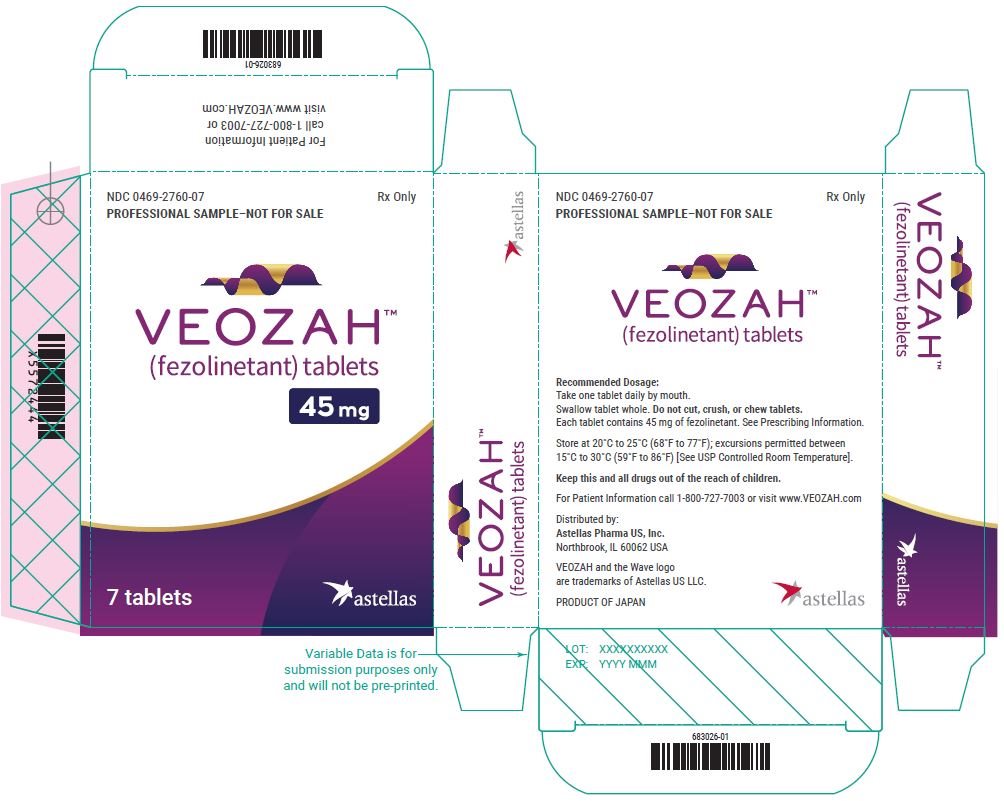
Package/Label Display Panel- VEOZAH Sample Blister Carton Label NDC 0469-2760-07 PROFESSIONAL SAMPLE-NOT FOR SALE Rx Only VEOZAH ® (fezolinetant) tablets 45 mg 7 tablets Package/Label Display Panel- VEOZAH Sample Blister Carton
Medicare Part D spend CMS · PART D · 2026 (Q1)
About this NDC listing & data coverage
What data is (and isn’t) available for this NDC — tap to expand
| NDC identity (package / product / labeler codes) | ✓ Available |
| Labeler | ✓ Available |
| Product & package description | ✓ Available |
| Marketing category & status | ✓ Available |
| Active ingredient / dosage form / route | ✓ Available |
| FDA label (SPL via DailyMed) | ✓ Available |
| Package photos | ✓ Available |
| Inactive ingredients (structured) | ✓ Available |
| NADAC pharmacy acquisition price (CMS) | — Not published for this NDC CMS publishes NADAC only for NDCs reported in its retail-pharmacy survey. |
| Orange Book / therapeutic-equivalence data | ✓ Available |
| HCPCS J-code billing crosswalk | — Not published for this NDC Most self-administered / retail products have no J-code — that is normal. |
| Medicaid utilization (CMS SDUD) | — Not published for this NDC CMS reports utilization only for NDCs with Medicaid claims above its privacy threshold. |