
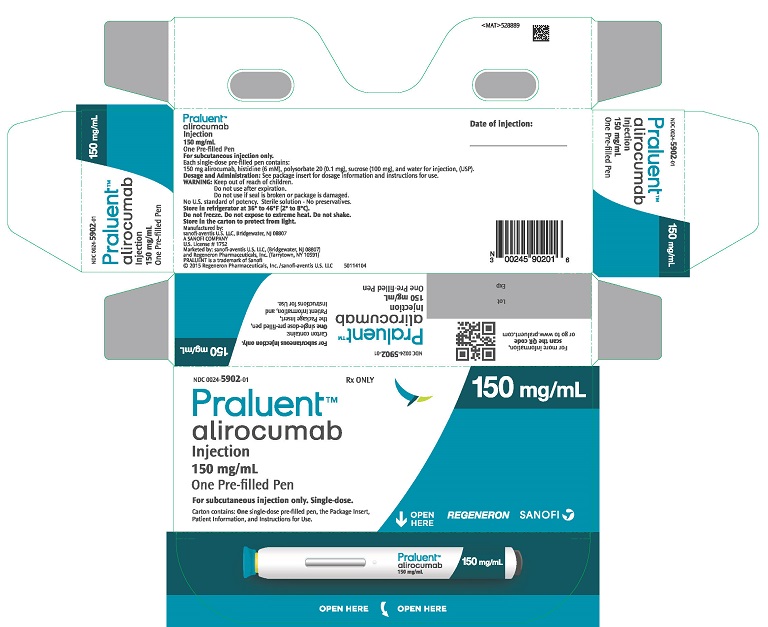
Praluent alirocumab 150 mg/mL Injection, Solution, 1 syringe
🆔 Identity & classification
Where does this data come from?
🏷️ RxNorm drug class
This medicine belongs to the PCSK9 Inhibitor class.
Where does this data come from?
🏭 Manufacturer & labeler
Where does this data come from?
🩺 Clinical
- Praluent is designed to lower your LDL cholesterol — the 'bad' kind that can build up in your arteries. Depending on why it was prescribed, it may be helping you get your cholester...
- What exactly is Praluent supposed to do for me?
- You inject Praluent just under the skin — into your thigh, abdomen, or upper arm — using the pre-filled device. It can take up to 20 seconds to inject. If it has been in the fridge...
- How do I give myself the injection, and does it hurt?
Patient education
Supplement & herbal interactions
Where does this data come from?
Ask a licensed pharmacist directly — free, answered by our team.
🧪 Inactive Ingredients / Excipients
Inactive ingredients, also called excipients, are components of the drug product other than the active ingredient. They may include fillers, dyes, coatings, preservatives, flavors, or other formulation ingredients.
💡 Tap an ingredient (hover on desktop) to see what it is and why it’s used.
-
6 mmol / 1 mL
UNII 4QD397987E
An amino acid used as a buffer and stabilizer in medications. It helps maintain the pH balance and protects the active drug from breaking down during storage and use.
-
0.1 mg / 1 mL
UNII 7T1F30V5YH
A synthetic emulsifier derived from sorbitol and oleic acid. It helps mix oil and water-based ingredients together and keeps them from separating in liquid formulations.
-
100 mg / 1 mL
UNII C151H8M554
A natural sugar derived from sugar cane or sugar beets. It's used as a sweetener, filler, and binder to improve taste, add bulk, and help hold tablet or capsule ingredients together.
-
UNII 059QF0KO0R
Water is a liquid solvent that dissolves and mixes ingredients together in liquid medicines, syrups, and injections. It helps distribute the active drug evenly throughout the product.
4 inactive ingredients listed in the exact product block matched to this NDC.
Where does this data come from?
ingredient classCode="IACT" elements from the exact product block matched by this NDC. Label-section narrative from DailyMed / the openFDA label index is shown separately when available.Inactive ingredient FAQ
Are inactive ingredients the same for every manufacturer?
Why might an inactive ingredient be missing?
Can inactive ingredients matter?
💲 Pricing
A drug doesn't have one price. Each row is a different public payment system, and none is what you'd pay at the counter — that depends on your insurance. The ⓘ on each row explains what it measures.
| Price system | Per mL | Per package |
|---|---|---|
| Retail pharmacies payNADAC · weekly | Not in the retail survey — common for institutional, discontinued, or low-volume packs. | |
| Medicaid paysCMS SDUD · 12 mo | No recent Medicaid claims on file for this NDC — rare and low-volume NDCs are suppressed in the public data. | |
| Medicare drug plans payPart D · Q2 2026 | $250.45 | $250.45 / 1 ml |
Where does this data come from?
🔁 Therapeutic equivalents
| Product | Labeler | Pack | NADAC/unit | TE | Status | Price vs. this |
|---|---|---|---|---|---|---|
| Praluent 150 mg/mLthis 00024-5902-00 | Sanofi-Aventis | 1 syringe | — | — | FDA listed | — |
| Praluent 150 mg/mL 61755-0021-01 | Regeneron | 1 syringe | — | — | FDA listed | — |
Where does this data come from?
⏳ Availability & biosimilar status
Biologics have no small-molecule generics; biosimilar competition is tracked in the FDA Purple Book.
Why the date isn’t exact: Biosimilar timing can change because patents may be challenged, settled, licensed, added or removed, and litigation can move the real date earlier or later.
🛈 What do these terms mean?
- Biologic patent
- A patent the reference product’s maker has publicly listed. A biosimilar generally can’t launch until these expire — unless they’re invalidated or resolved in a settlement.
- Reference-product exclusivity
- A flat 12 years of FDA market protection from the biologic’s first licensure (the BPCIA). No biosimilar can be licensed before it ends, regardless of patents.
- Interchangeable exclusivity
- The first interchangeable biosimilar can earn a period as the only interchangeable version (pharmacists can substitute it without the prescriber).
- Earliest biosimilar (LOE)
- The latest of all the dates above — the soonest a biosimilar can realistically reach the market. Litigation and settlements can move it earlier.
Biologics have no small-molecule “generics” — competition comes from FDA-licensed biosimilars, tracked in the FDA Purple Book.
| Code | What it grants | Expires |
|---|---|---|
| RefProduct | Reference-product exclusivity (12-year, BPCIA) — no biosimilar can be licensed before this date | Jul 24, 2027 |
Is there a biosimilar for PRALUENT 150 MG/ML PEN?
Why do different websites show different biosimilar dates?
Can a biosimilar launch before the last patent expires?
What does “current Purple Book estimate” mean?
What does “FDA listed” mean?
What does a patent or protection date mean here?
Where does this data come from?
📊 Medicare Part D spend CMS · PART D · 2026 (Q1)
🔬 Reported adverse events (FAERS)
Top reported reactions
Reporter sex
Serious outcomes
Where does this data come from?
📦 Packaging — all sizes for this product
| Package NDC | Description | Marketing start | Status |
|---|---|---|---|
| 00024-5902-00 You're viewing this | 1 SYRINGE, GLASS in 1 CARTON (0024-5902-00) / 1 mL in 1 SYRINGE, GLASS | 2015-07-24 | Active |
| 00024-5902-01 | 1 SYRINGE, GLASS in 1 CARTON (0024-5902-01) / 1 mL in 1 SYRINGE, GLASS | 2015-07-24 | Active |
| 00024-5902-02 | 2 SYRINGE, GLASS in 1 CARTON (0024-5902-02) / 1 mL in 1 SYRINGE, GLASS | 2015-07-24 | Discontinued by firm |
Pack size FAQ
What quantity is in NDC 00024-5902-00?
What is the difference between NDC 00024-5902-00 and NDC 00024-5902-01?
What NDC number is used to bill for this package of Praluent alirocumab 150 mg/mL Injection, Solution?
🧭 About this NDC listing & data coverage
What data is (and isn’t) available for this NDC — tap to expand
| NDC identity (package / product / labeler codes) | ✓ Available |
| Labeler | ✓ Available |
| Product & package description | ✓ Available |
| Marketing category & status | ✓ Available |
| Active ingredient / dosage form / route | ✓ Available |
| FDA label (SPL via DailyMed) | ✓ Available |
| Package photos | ✓ Available |
| Inactive ingredients (structured) | ✓ Available |
| NADAC pharmacy acquisition price (CMS) | — Not published for this NDC CMS publishes NADAC only for NDCs reported in its retail-pharmacy survey. |
| Orange Book / therapeutic-equivalence data | ✓ Available |
| HCPCS J-code billing crosswalk | — Not published for this NDC Most self-administered / retail products have no J-code — that is normal. |
| Medicaid utilization (CMS SDUD) | — Not published for this NDC CMS reports utilization only for NDCs with Medicaid claims above its privacy threshold. |
Questions about this listing
Why is there no price listed?
Is the NDC printed on the package the same as the 11-digit billing NDC?
What do the three segments of this NDC mean?
Is this package still being marketed?
Does this product come in other package sizes?
Who lists this product with the FDA?
Do I need a prescription for this product?
Where does this data come from?
📄 Full prescribing information FDA SPL
🎯 Indications and Usage ▾
1 INDICATIONS AND USAGE PRALUENT ® is indicated: To reduce the risk of major adverse cardiovascular (CV) events (coronary heart disease death, myocardial infarction, stroke, or unstable angina requiring hospitalization) in adults at increased risk for these events . As an adjunct to diet and exercise to reduce low- density lipoprotein cholesterol (LDL-C) in: adults with hypercholesterolemia. adults and pediatric patients aged 8 years and older with heterozygous familial hypercholesterolemia (HeFH). adults with homozygous familial hypercholesterolemia (HoFH).
PRALUENT is a proprotein convertase subtilisin kexin type 9 (PCSK9) inhibitor indicated: To reduce the risk of major adverse cardiovascular (CV) events (coronary heart disease death, myocardial infarction, stroke, or unstable angina requiring hospitalization) in adults at increased risk for these events. ( 1 ) As an adjunct to diet and exercise to reduce low-density lipoprotein cholesterol (LDL-C) in : adults with hypercholesterolemia ( 1 ) adults and pediatric patients aged 8 years and older with heterozygous familial hypercholesterolemia (HeFH).
( 1 ) adults with homozygous familial hypercholesterolemia (HoFH). ( 1 )
⏱️ Dosage and Administration ▾
2 DOSAGE AND ADMINISTRATION In adults with hypercholesterolemia, including HeFH ( 2.1 ): The recommended starting dosage of PRALUENT is either 75 mg once every 2 weeks or 300 mg once every 4 weeks administered subcutaneously. For patients receiving PRALUENT 300 mg every 4 weeks, measure LDL-C just prior to the next scheduled dosage, because LDL-C can vary between dosages in some patients. If the LDL-C response is inadequate, the dosage may be adjusted 150 mg subcutaneously every 2 weeks.
In adults with HeFH undergoing LDL apheresis or in adults with HoFH ( 2.1 ): The recommended dosage of PRALUENT is 150 mg once every 2 weeks administered subcutaneously. PRALUENT can be administered without regard to the timing of LDL apheresis. In pediatric patients with HeFH ( 2.2 ): The recommended dosage of PRALUENT for patients with a body weight less than 50 kg is 150 mg once every 4 weeks administered subcutaneously.
The recommended dosage of PRALUENT for patients with a body weight of 50 kg or more is 300 mg once every 4 weeks administered subcutaneously. If the LDL-C response is inadequate, the dosage may be adjusted for patients with a body weight less than 50 kg to 75 mg subcutaneously once every 2 weeks or for patients with a body weight of 50 kg or more to 150 mg subcutaneously once every 2 weeks. Assess LDL-C when clinically appropriate.
The LDL-lowering effect of PRALUENT may be measured as early as 4 weeks after initiation. ( 2.1 ) Administer PRALUENT subcutaneously into areas of the thigh, abdomen, or upper arm that are not tender, bruised, red, or indurated. Rotate injection sites for each administration.
( 2.4 ) To administer the 300 mg dosage, give two 150 mg PRALUENT injections consecutively at two different injection sites. ( 2.4 )
2.1Recommended Dosage in Adults Hypercholesterolemia, including HeFH: The recommended starting dosage of PRALUENT is either 75 mg once every 2 weeks or 300 mg once every 4 weeks administered subcutaneously [see Dosage and Administration (2.4) ] . For patients receiving PRALUENT 300 mg every 4 weeks, measure LDL-C just prior to the next scheduled dose, because LDL-C can vary between doses in some patients [see Clinical Studies (14) ] . If the LDL-C response is inadequate, the dosage may be adjusted to 150 mg subcutaneously every 2 weeks.
HeFH undergoing LDL apheresis or with HoFH: The recommended dosage of PRALUENT is 150 mg once every 2 weeks administered subcutaneously [see Dosage and Administration (2.4) ] . PRALUENT can be administered without regard to the timing of LDL apheresis. Assess LDL-C when clinically appropriate.
The LDL-lowering effect of PRALUENT may be measured as early as 4 weeks after initiation.
2.2Recommended Dosage in Pediatric Patients Aged 8 years and Older With HeFH The recommended dosage of PRALUENT for patients with a body weight less than 50 kg is 150 mg once every 4 weeks administered subcutaneously [see Dosage and Administration (2.4) ] . If the LDL-C lowering response is inadequate, the dosage may be adjusted to 75 mg subcutaneously once every 2 weeks [see Dosage and Administration (2.4) ] . The recommended dosage of PRALUENT for patients with a body weight of 50 kg or more is 300 mg once every 4 weeks administered subcutaneously [see Dosage and Administration (2.4) ] .
If the LDL-C lowering response is inadequate, the dosage may be adjusted to 150 mg subcutaneously once every 2 weeks [see Dosage and Administration (2.4) ] . Assess LDL-C when clinically appropriate. The LDL-lowering effect of PRALUENT may be measured as early as 4 weeks after initiation.
2.3Missed Doses If a dose is missed: Within 7 days from the missed dose, instruct the patient to administer PRALUENT and resume the patient's original schedule. More than 7 days after the missed dose: For every 2-week dosage, instruct the patient to wait until the next dose on the original schedule. For every 4-week dosage, instruct the patient to administer the dose and start a new schedule based on this date.…
💊 Dosage Forms and Strengths ▾
3 DOSAGE FORMS AND STRENGTHS PRALUENT injection is a clear, colorless to pale yellow solution available as follows: 75 mg/mL single-dose pre-filled pen 150 mg/mL single-dose pre-filled pen Injection: 75 mg/mL or 150 mg/mL in a single-dose pre-filled pen. ( 3 )
⛔ Contraindications ▾
4 CONTRAINDICATIONS PRALUENT is contraindicated in patients with a history of a serious hypersensitivity reaction to alirocumab or any of the excipients in PRALUENT. Hypersensitivity vasculitis, angioedema, and hypersensitivity reactions requiring hospitalization have occurred [see Warnings and Precautions (5.1) ]. History of a serious hypersensitivity reaction to alirocumab or any of the excipients in PRALUENT. ( 4 )
⚠️ Warnings and Cautions ▾
5 WARNINGS AND PRECAUTIONS Hypersensitivity reactions: hypersensitivity vasculitis, angioedema, and other hypersensitivity reactions requiring hospitalization, have been reported with PRALUENT treatment. If signs or symptoms of serious hypersensitivity reactions occur, discontinue treatment with PRALUENT, treat according to the standard of care, and monitor until signs and symptoms resolve. ( 5.1 )
5.1Hypersensitivity Reactions Hypersensitivity reactions, including hypersensitivity vasculitis, angioedema, and other hypersensitivity reactions requiring hospitalization, have been reported with PRALUENT treatment. If signs or symptoms of serious hypersensitivity reactions occur, discontinue treatment with PRALUENT, treat according to the standard of care, and monitor until signs and symptoms resolve. PRALUENT is contraindicated in patients with a history of a serious hypersensitivity reaction to alirocumab or any excipient in PRALUENT [see Contraindications (4) ] .
🤒 Adverse Reactions ▾
6 ADVERSE REACTIONS The following adverse reactions are also discussed in the other sections of the labeling: Hypersensitivity Reactions [see Warnings and Precautions (5.1) ] Common (>5% of patients treated with PRALUENT and more frequently than placebo) adverse reactions in adults with: Primary hypercholesterolemia : injection site reactions, and influenza. ( 6 ) Established CV disease: myalgia. ( 6 ) To report SUSPECTED ADVERSE REACTIONS, contact Regeneron at 1-844-734-6643 or FDA at 1-800-FDA-1088 or www.fda.gov/medwatch.
6.1Clinical Trials Experience Because clinical trials are conducted under widely varying conditions, adverse reaction rates observed in the clinical trials of a drug cannot be directly compared to rates in the clinical trials of another drug and may not reflect the rates observed in practice. Adverse Reactions in Adults with Hypercholesterolemia The data in Table 1 are derived from 9 primary hypercholesterolemia placebo-controlled trials that included 2,476 adult patients treated with PRALUENT 75 mg and/or 150 mg every 2 weeks, including 2,135 exposed for 6 months and 1,999 exposed for more than 1 year (median treatment duration of 65 weeks).
The mean age of the population was 59 years, 40% of the population were female, 90% were White, 4% were Black or African American, 3% were Asian, and 3% other races; 6% identified as Hispanic or Latino ethnicity. Adverse reactions reported in at least 2% of PRALUENT-treated patients, and more frequently than in placebo-treated patients, are shown in Table 1. Table 1: Adverse Reactions Occurring in >2% of PRALUENT-Treated Adult Patients and ≥1% More Frequently Than with Placebo Adverse Reactions Placebo (N=1,276) % PRALUENT 75 mg every 2 weeks and 150 mg every 2 weeks combined (N=2,476) % Injection site reactions Includes erythema/redness, itching, swelling, pain/tenderness 5 7 Influenza 5 6 Diarrhea 4 5 Myalgia 3 4 Muscle spasms 2 3 Contusion 1 2 Adverse reactions led to discontinuation of treatment in 5.3% of patients treated with PRALUENT and 5.1% of patients treated with placebo.
The most common adverse reactions leading to treatment discontinuation in patients treated with PRALUENT were allergic reactions (0.6% versus 0.2% for PRALUENT and placebo, respectively) and elevated liver enzymes (0.3% versus <0.1%). In an analysis of ezetimibe-controlled trials in which 864 patients were exposed to PRALUENT for a median of 27 weeks and 618 patients were exposed to ezetimibe for a median of 24 weeks, the types and frequencies of common adverse reactions were similar to those listed above. Adverse Reactions in a Cardiovascular Outcomes Trial in Adults In a CV outcomes trial in which 9,451 patients were exposed to PRALUENT for a median of 31 months and 9,443 patients were exposed to placebo for a median of 32 months, common adverse reactions (greater than 5% of patients treated with PRALUENT and occurring more frequently than placebo) included myalgia (6% PRALUENT, 5% placebo).
Adverse Reactions in Pediatric Patients with HeFH In a 24-week placebo-controlled clinical trial in which 101 pediatric patients aged 8 to 17 years with HeFH were exposed to PRALUENT and 52 pediatric patients with HeFH were exposed to placebo [see Clinical Studies (14.3) ], the safety profile of PRALUENT observed in this population was consistent with the safety profile observed in adults with HeFH. Other Adverse Reactions Local Injection Site Reactions In a pool of placebo-controlled trials evaluating PRALUENT 75 mg and/or 150 mg administered every 2 weeks in adults, local injection site reactions including erythema/redness, itching, swelling, and pain/tenderness were reported more frequently in patients treated with PRALUENT (7.2% versus 5.1% for PRALUENT and placebo, respectively).
Few patients discontinued treatment because of these reactions (0.2% versus 0.4% for PRALUENT and placebo, respectively), but patients receiving PRALUENT had a greater number of injection site reactions, had more repor…
👥 Use in Specific Populations ▾
8 USE IN SPECIFIC POPULATIONS
8.1Pregnancy Risk Summary Available data from clinical trials and postmarketing reports on PRALUENT use in pregnant women are insufficient to evaluate for a drug-associated risk of major birth defects, miscarriage or other adverse maternal or fetal outcomes. In animal reproduction studies, there were no effects on embryo-fetal development when rats were subcutaneously administered alirocumab during organogenesis at dose exposures up to 12-fold the exposure at the maximum recommended human dose of 150 mg every two weeks.
In monkeys, suppression of the humoral immune response was observed in infant monkeys when alirocumab was dosed during organogenesis to parturition at dose exposures 13-fold the exposure at the maximum recommended human dose of 150 mg every two weeks. No additional effects on pregnancy or neonatal/infant development were observed at dose exposures up to 81-fold the maximum recommended human dose of 150 mg every two weeks. Measurable alirocumab serum concentrations were observed in the infant monkeys at birth at comparable levels to maternal serum, indicating that alirocumab, like other IgG antibodies, crosses the placental barrier.
Monoclonal antibodies are transported across the placenta in increasing amounts especially near term; therefore, alirocumab has the potential to be transmitted from the mother to the developing fetus. The estimated background risk of major birth defects and miscarriage for the indicated population(s) is unknown. In the U.S. general population, the estimated background risk of major birth defects and miscarriage in clinically recognized pregnancies is 2%-4% and 15%-20%, respectively.
There is a pregnancy safety study for PRALUENT. If PRALUENT is administered during pregnancy, healthcare providers should report PRALUENT exposure by contacting Regeneron at 1-844-734-6643. Data Animal data In Sprague Dawley rats, no effects on embryo-fetal development were observed when alirocumab was dosed at up to 75 mg/kg/dose by the subcutaneous route on gestation days 6 and 12 at exposures 12-fold the maximum recommended human dose of 150 mg every two weeks, based on serum AUC.
In cynomolgus monkeys, suppression of the humoral immune response to keyhole limpet hemocyanin (KLH) antigen was observed in infant monkeys at 4 to 6 months of age when alirocumab was dosed during organogenesis to parturition at 15 mg/kg/week and 75 mg/kg/week by the subcutaneous route, corresponding to 13-fold and 81-fold the human exposure at the maximum recommended human dose of 150 mg every two weeks, based on serum AUC. The lowest dose tested in the monkey resulted in humoral immune suppression; therefore, it is unknown if this effect would be observed at clinical exposure.
No study designed to challenge the immune system of infant monkeys was conducted. No additional embryo-fetal, prenatal or postnatal effects were observed in infant monkeys, and no maternal effects were observed, when alirocumab was dosed at up to 75 mg/kg/week by the subcutaneous route, corresponding to maternal exposure of 81-fold the exposure at the maximum recommended human dose of 150 mg every two weeks, based on serum AUC.
8.2Lactation Risk Summary There is no information regarding the presence of alirocumab in human milk, the effects on the breastfed infant, or the effects on milk production. The development and health benefits of breastfeeding should be considered along with the mother's clinical need for PRALUENT and any potential adverse effects on the breastfed infant from PRALUENT or from the underlying maternal condition. Human IgG is present in human milk, but published data suggest that breast milk IgG antibodies do not enter the neonatal and infant circulation in substantial amounts.
8.4Pediatric Use The safety and effectiveness of PRALUENT as an adjunct to diet and other LDL-C-lowering therapies for the treatment of HeFH have been established in pediatric patients aged 8 years and older. Use of PRALUENT…
🤰 Pregnancy ▾
8.1Pregnancy Risk Summary Available data from clinical trials and postmarketing reports on PRALUENT use in pregnant women are insufficient to evaluate for a drug-associated risk of major birth defects, miscarriage or other adverse maternal or fetal outcomes. In animal reproduction studies, there were no effects on embryo-fetal development when rats were subcutaneously administered alirocumab during organogenesis at dose exposures up to 12-fold the exposure at the maximum recommended human dose of 150 mg every two weeks.
In monkeys, suppression of the humoral immune response was observed in infant monkeys when alirocumab was dosed during organogenesis to parturition at dose exposures 13-fold the exposure at the maximum recommended human dose of 150 mg every two weeks. No additional effects on pregnancy or neonatal/infant development were observed at dose exposures up to 81-fold the maximum recommended human dose of 150 mg every two weeks. Measurable alirocumab serum concentrations were observed in the infant monkeys at birth at comparable levels to maternal serum, indicating that alirocumab, like other IgG antibodies, crosses the placental barrier.
Monoclonal antibodies are transported across the placenta in increasing amounts especially near term; therefore, alirocumab has the potential to be transmitted from the mother to the developing fetus. The estimated background risk of major birth defects and miscarriage for the indicated population(s) is unknown. In the U.S. general population, the estimated background risk of major birth defects and miscarriage in clinically recognized pregnancies is 2%-4% and 15%-20%, respectively.
There is a pregnancy safety study for PRALUENT. If PRALUENT is administered during pregnancy, healthcare providers should report PRALUENT exposure by contacting Regeneron at 1-844-734-6643. Data Animal data In Sprague Dawley rats, no effects on embryo-fetal development were observed when alirocumab was dosed at up to 75 mg/kg/dose by the subcutaneous route on gestation days 6 and 12 at exposures 12-fold the maximum recommended human dose of 150 mg every two weeks, based on serum AUC.
In cynomolgus monkeys, suppression of the humoral immune response to keyhole limpet hemocyanin (KLH) antigen was observed in infant monkeys at 4 to 6 months of age when alirocumab was dosed during organogenesis to parturition at 15 mg/kg/week and 75 mg/kg/week by the subcutaneous route, corresponding to 13-fold and 81-fold the human exposure at the maximum recommended human dose of 150 mg every two weeks, based on serum AUC. The lowest dose tested in the monkey resulted in humoral immune suppression; therefore, it is unknown if this effect would be observed at clinical exposure.
No study designed to challenge the immune system of infant monkeys was conducted. No additional embryo-fetal, prenatal or postnatal effects were observed in infant monkeys, and no maternal effects were observed, when alirocumab was dosed at up to 75 mg/kg/week by the subcutaneous route, corresponding to maternal exposure of 81-fold the exposure at the maximum recommended human dose of 150 mg every two weeks, based on serum AUC.
🧒 Pediatric Use ▾
8.4Pediatric Use The safety and effectiveness of PRALUENT as an adjunct to diet and other LDL-C-lowering therapies for the treatment of HeFH have been established in pediatric patients aged 8 years and older. Use of PRALUENT for this indication is based on data from a 24-week, randomized, placebo-controlled, double-blind trial in pediatric patients with HeFH. In the trial, 101 patients received PRALUENT and 52 patients received placebo; 26 patients (17%) were 8 to 9 years of age.
This indication is supported by evidence from controlled trials in adults [see Adverse Reactions (6.1) and Clinical Studies (14.3) ]. The safety and effectiveness of PRALUENT have not been established in pediatric patients with HeFH who are younger than 8 years of age or in pediatric patients with other types of hypercholesterolemia.
🧓 Geriatric Use ▾
8.5Geriatric Use In controlled trials, 3663 patients treated with PRALUENT were ≥65 years of age and 734 patients treated with PRALUENT were ≥75 years of age. No overall differences in safety or effectiveness were observed between these patients and younger patients, and other reported clinical experience has not identified differences in responses between the elderly and younger patients, but greater sensitivity of some older individuals cannot be ruled out.
🧬 Clinical Pharmacology ▾
12 CLINICAL PHARMACOLOGY
12.1Mechanism of Action Alirocumab is a human monoclonal antibody that binds to proprotein convertase subtilisin kexin type 9 (PCSK9). PCSK9 binds to the low-density lipoprotein (LDL) receptors (LDLR) on the surface of hepatocytes to promote LDLR degradation within the liver. By inhibiting the binding of PCSK9 to LDLR, alirocumab increases the number of LDLRs available to clear LDL, thereby lowering LDL-C levels.
12.2Pharmacodynamics Alirocumab reduced free PCSK9 in a concentration-dependent manner. Following a single subcutaneous administration of alirocumab 75 or 150 mg, maximal suppression of free PCSK9 occurred within 4 to 8 hours. Free PCSK9 concentrations returned to baseline when alirocumab concentrations decreased below the limit of quantitation.
12.3Pharmacokinetics Absorption After subcutaneous administration of 75 mg to 300 mg alirocumab, median times to maximum serum concentrations (t max ) were 3-7 days. The pharmacokinetics of alirocumab after single subcutaneous administration of 75 mg into the abdomen, upper arm, or thigh were similar. The absolute bioavailability of alirocumab after subcutaneous administration was about 85% as determined by population pharmacokinetics analysis.
A slightly greater than dose proportional increase was observed, with a 2.1-fold to 2.7-fold increase in total alirocumab concentrations for a 2-fold increase in dose from 75 mg every 2 weeks to 150 mg every 2 weeks. Monthly dose normalized exposure with 300 mg every 4 weeks treatment was similar to that of 150 mg every 2 weeks. Steady state was reached after 2 to 3 doses with an accumulation ratio up to a maximum of about 2-fold.
Distribution Following intravenous administration, the volume of distribution was about 0.04 to
0.05L/kg indicating that alirocumab is distributed primarily in the circulatory system. Elimination Specific metabolism studies were not conducted, because alirocumab is a protein. Alirocumab is expected to degrade to small peptides and individual amino acids.
In clinical studies where alirocumab was administered in combination with atorvastatin or rosuvastatin, no relevant changes in statin concentrations were observed in the presence of repeated administration of alirocumab, indicating that cytochrome P450 enzymes (mainly CYP3A4 and CYP2C9) and transporter proteins such as P-gp and OATP were not affected by alirocumab. Two elimination phases were observed for alirocumab. At low concentrations, the elimination is predominately through saturable binding to target (PCSK9), while at higher concentrations the elimination of alirocumab is largely through a non-saturable proteolytic pathway.
Based on a population pharmacokinetic analysis, the median apparent half-life of alirocumab at steady state was 17 to 20 days in patients receiving alirocumab at subcutaneous doses of 75 mg every 2 weeks or 150 mg every 2 weeks. Specific Populations A population pharmacokinetic analysis was conducted on data from 2799 patients. Age, body weight, gender, race, and creatinine clearance were found not to significantly influence alirocumab pharmacokinetics.
Pediatric Patients The pharmacokinetics of alirocumab were evaluated in 140 pediatric patients aged 8 to 17 years with HeFH. Steady-state concentrations were reached at or before week 8 (first PK sampling during repeated dosing) with recommended dosing regimen [see Dosage and Administration (2.2) ] . Renal Impairment Since monoclonal antibodies are not known to be eliminated via renal pathways, renal function is not expected to impact the pharmacokinetics of alirocumab.
No data are available in patients with severe renal impairment. Hepatic Impairment Following administration of a single 75 mg SC dose, alirocumab pharmacokinetic profiles in patients with mild and moderate hepatic impairment were similar to those in patients with normal hepatic function. No data are available in patients with severe hepatic impairment.
Drug-Drug Interactions The median appa…
🧬 Mechanism of Action ▾
12.1Mechanism of Action Alirocumab is a human monoclonal antibody that binds to proprotein convertase subtilisin kexin type 9 (PCSK9). PCSK9 binds to the low-density lipoprotein (LDL) receptors (LDLR) on the surface of hepatocytes to promote LDLR degradation within the liver. By inhibiting the binding of PCSK9 to LDLR, alirocumab increases the number of LDLRs available to clear LDL, thereby lowering LDL-C levels.
📦 How Supplied / Storage and Handling ▾
16 HOW SUPPLIED/STORAGE AND HANDLING PRALUENT injection is a clear, colorless to pale yellow solution, supplied as follows: Pack Size 75 mg/mL Pre-filled Pen 150 mg/mL Pre-filled Pen Pack of 1 pen NDC 0024-5901-01 NDC 0024-5902-01 The needle shield is not made with natural rubber latex. Store in a refrigerator at 36°F to 46°F (2°C to 8°C) in the original carton to protect from light. Do not freeze.
Do not shake. PRALUENT may be kept at room temperature up to 77°F (25°C) in the original carton for 30 days. If not used within the 30 days, discard PRALUENT.
📦 Storage and Handling ▾
Store in a refrigerator at 36°F to 46°F (2°C to 8°C) in the original carton to protect from light. Do not freeze. Do not shake. PRALUENT may be kept at room temperature up to 77°F (25°C) in the original carton for 30 days. If not used within the 30 days, discard PRALUENT.
📋 Description ▾
11 DESCRIPTION Alirocumab is a human monoclonal antibody (IgG1 isotype) that targets proprotein convertase subtilisin kexin type 9 (PCSK9). Alirocumab is a PCSK9 inhibitor produced by recombinant DNA technology in Chinese Hamster Ovary cell suspension culture. Alirocumab consists of two disulfide-linked human heavy chains, each covalently linked through a disulfide bond to a human kappa light chain.
A single N-linked glycosylation site is located in each heavy chain within the CH2 domain of the Fc constant region of the molecule. The variable domains of the heavy and light chains combine to form the PCSK9 binding site within the antibody. Alirocumab has an approximate molecular weight of 146 kDa.
PRALUENT is a sterile, preservative-free, clear, colorless to pale yellow solution for subcutaneous use. PRALUENT 75 mg/mL or 150 mg/mL solution for subcutaneous injection in a single-dose pre-filled pen is supplied in a siliconized 1 mL Type-1 clear glass syringe. Each 75 mg/mL pre-filled pen contains 75 mg alirocumab, histidine (8 mM), polysorbate 20 (0.1 mg), sucrose (100 mg), and Water for Injection USP, to pH 6.0.
Each 150 mg/mL pre-filled pen contains 150 mg alirocumab, histidine (6 mM), polysorbate 20 (0.1 mg), sucrose (100 mg), and Water for Injection USP, to pH 6.0.
💬 Information for Patients ▾
17 PATIENT COUNSELING INFORMATION Advise the patient to read the FDA-approved patient labeling (Patient Information and Instructions for Use). Hypersensitivity Reactions Inform patients that serious hypersensitivity reactions (e.g., angioedema) have been reported in patients treated with PRALUENT. Advise patients on the symptoms of hypersensitivity reactions and instruct them to discontinue PRALUENT and seek medical attention promptly, if such symptoms occur [ see Warnings and Precautions (5.1) ] .
Administration Provide guidance to patients and caregivers on proper subcutaneous injection technique and how to use the pre-filled pen. Inform patients that it may take up to 20 seconds to inject PRALUENT. Inform patients the pre-filled pen should be allowed to warm to room temperature for 30 to 40 minutes prior to use if refrigerated.
Pregnancy Advise women who are exposed to PRALUENT during pregnancy that there is a pregnancy safety study that monitors pregnancy outcomes. Encourage these patients to report their pregnancy to Regeneron at 1 844-734-6643 [see Use in Specific Populations (8.1) ].