
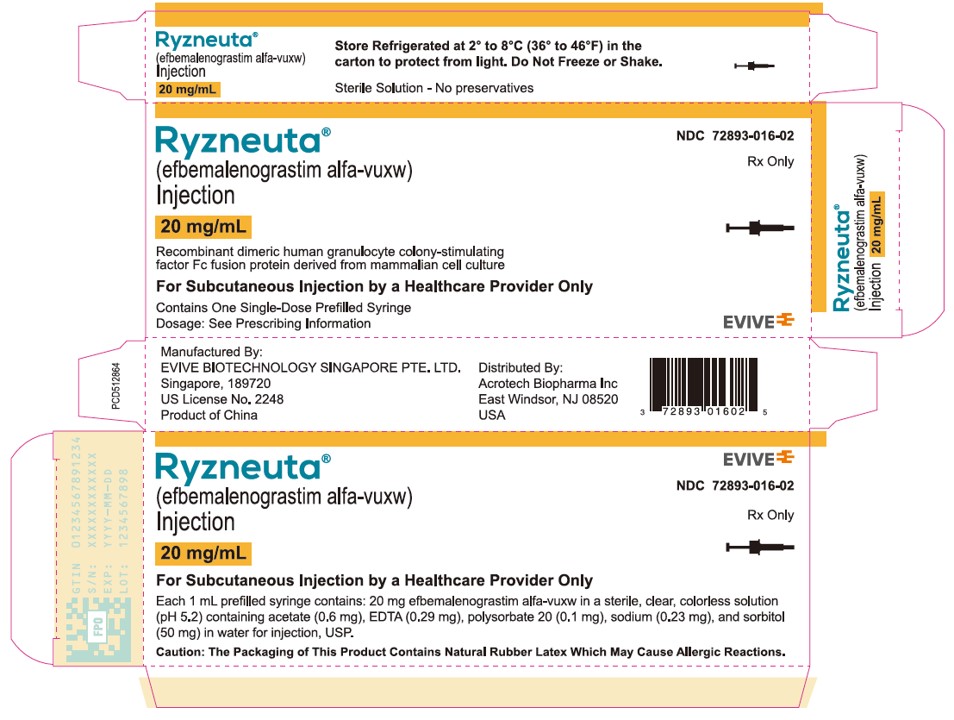
RYZNEUTA efbemalenograstim alfa-vuxw 20 mg/mL Injection, 1 syringe — NDC 72893-016-02 (Billing 72893-0016-02)
This is a package of 1 syringe of RYZNEUTA efbemalenograstim alfa-vuxw 20 mg/mL Injection from Acrotech Biopharma Inc, marketed since Nov 2023 and currently FDA-listed. It is this product's only package size.
NDC database record
One package, one record: these facts belong to NDC 72893-016-02 alone.
- Record
- FDA NDC Directory package listing · Human prescription drug
- Code segments
- 72893 labeler · 016 product · 02 package
- Package marketed since
- Nov 23, 2023
- Sample package
- No — commercial package
- Listing certified through
- Dec 31, 2027
- Barcode (UPC-A, from the NDC)
- 3 7289301602 5
- FDA record last changed
- Jul 24, 2026
Identity & classification
Regulatory identifiers FDA, NLM and CMS codes for this package
Drug-database identifiers Medi-Span GPI and First Databank GCN / HICL / AHFS classification
- GSN (GCN sequence number): 085541
- GCN: 55048
- GPI-14 (Medi-Span): 8240151680E520
- HICL (First Databank): 049318
- AHFS class code: 20:16.00.00
- RxCUI (RxNorm): 2670662
Where does this data come from?
- FDA openFDA NDC Directory · synced Oct 8, 2026
- FDA label on DailyMed · label index refreshed Oct 8, 2026
- RxNorm (NLM RxNav) · catalog refreshed Oct 8, 2026
- Medi-Span GPI (licensed)
- First Databank (licensed) · refreshed Oct 8, 2026
RxNorm drug class
This medicine belongs to the Leukocyte Growth Factor class.
Where does this data come from?
- RxClass (NLM) · catalog refreshed Oct 8, 2026
Clinical
Efbemalenograstim alfa-vuxw injection is used to decrease the chance of infection in people who have nonmyeloid cancer (cancer that does not involve the bone marrow) and are receiving chemotherapy medications that may decrease the number of neutrophils (a type of blood cell needed to fight infection). Efbemalenograstim alfa-vuxw is in a class of medications called colony-stimulating factors. It works by helping the body make more neutrophils.
Read the full MedlinePlus article ↗Patient education
Supplement & herbal interactions
Where does this data come from?
- MedlinePlus (NLM) · refreshed Oct 8, 2026
- FDA label on DailyMed · label index refreshed Oct 8, 2026
Ask a licensed pharmacist directly — free, answered by our team.
Pricing
A drug doesn't have one price. Each row is a different public payment system, and none is what you'd pay at the counter — that depends on your insurance. The ⓘ on each row explains what it measures.
| Price system | Per each | Per package |
|---|---|---|
| Retail pharmacies payNADAC · weekly | Not in the retail survey — common for institutional, discontinued, or low-volume packs. | |
| Medicaid paysCMS SDUD · 12 mo | No recent Medicaid claims on file for this NDC — rare and low-volume NDCs are suppressed in the public data. | |
| Medicare drug plans payPart D · quarterly | No Part D plan price is available for this NDC in our data. | |
Where does this data come from?
- CMS NADAC weekly file
- CMS ASP pricing files · refreshed Sep 20, 2026
- CMS Medicaid State Drug Utilization Data · refreshed Oct 9, 2026
- CMS Part D plan pricing files · refreshed Sep 24, 2026
- VA National Acquisition Center price file
Packaging — all sizes for this product
| Package NDC | Description | Marketing start | Marketing end | Status |
|---|---|---|---|---|
| 72893-0016-02 You're viewing this Main listing | 1 SYRINGE in 1 CARTON / 1 mL in 1 SYRINGE | 2023-11-23 | — | Active |
Therapeutic equivalents
| Product | Labeler | Pack | NADAC/unit | TE | Status | Price vs. this |
|---|---|---|---|---|---|---|
| Ryzneuta 20 mg/mLthis 72893-0016-02 | Acrotech | 1 syringe | — | — | FDA listed | — |
Where does this data come from?
- FDA openFDA NDC Directory · synced Oct 8, 2026
- FDA Purple Book · refreshed Oct 5, 2026
- CMS NADAC weekly file
Availability & biosimilar status
Biologics have no small-molecule generics; biosimilar competition is tracked in the FDA Purple Book.
Why the date isn’t exact: Biosimilar timing can change because patents may be challenged, settled, licensed, added or removed, and litigation can move the real date earlier or later.
🛈 What do these terms mean?
- Biologic patent
- A patent the reference product’s maker has publicly listed. A biosimilar generally can’t launch until these expire — unless they’re invalidated or resolved in a settlement.
- Reference-product exclusivity
- A flat 12 years of FDA market protection from the biologic’s first licensure (the BPCIA). No biosimilar can be licensed before it ends, regardless of patents.
- Interchangeable exclusivity
- The first interchangeable biosimilar can earn a period as the only interchangeable version (pharmacists can substitute it without the prescriber).
- Earliest biosimilar (LOE)
- The latest of all the dates above — the soonest a biosimilar can realistically reach the market. Litigation and settlements can move it earlier.
Biologics have no small-molecule “generics” — competition comes from FDA-licensed biosimilars, tracked in the FDA Purple Book.
| Code | What it grants | Expires |
|---|---|---|
| RefProduct | Reference-product exclusivity (12-year, BPCIA) — no biosimilar can be licensed before this date | Nov 16, 2035 |
Is there a biosimilar for RYZNEUTA 20 MG/ML SYRINGE?
Why do different websites show different biosimilar dates?
Can a biosimilar launch before the last patent expires?
What does “current Purple Book estimate” mean?
What does “FDA listed” mean?
What does a patent or protection date mean here?
Where does this data come from?
- FDA Purple Book · refreshed Oct 5, 2026
Inactive Ingredients / Excipients
Inactive ingredients, also called excipients, are components of the drug product other than the active ingredient. They may include fillers, dyes, coatings, preservatives, flavors, or other formulation ingredients.
💡 Tap an ingredient (hover on desktop) to see what it is and why it’s used.
-
0.6 mg / 1 mL
UNII 569DQM74SC
Acetate ion is a salt form derived from acetic acid. It's commonly used in medicines as a buffer to help maintain the correct pH level and stabilize the product.
-
0.29 mg / 1 mL
UNII 9G34HU7RV0
EDTA is a chemical compound that binds to metals and other substances in medicines. It's used as a preservative and stabilizer to prevent discoloration and extend shelf life.
-
0.1 mg / 1 mL
UNII 7T1F30V5YH
A synthetic emulsifier derived from sorbitol and oleic acid. It helps mix oil and water-based ingredients together and keeps them from separating in liquid formulations.
-
0.23 mg / 1 mL
UNII 9NEZ333N27
Sodium is a mineral element that helps regulate fluid balance and nerve function in the body. In medicines, sodium salts serve as buffers, preservatives, or carriers to stabilize the drug and control its pH.
-
50 mg / 1 mL
UNII 506T60A25R
Sorbitol is a natural sugar alcohol derived from glucose. It serves as a sweetener, humectant, and bulking agent in medications to improve taste and help maintain moisture in the product.
-
UNII 059QF0KO0R
Water is a liquid solvent that dissolves and mixes ingredients together in liquid medicines, syrups, and injections. It helps distribute the active drug evenly throughout the product.
6 inactive ingredients listed in the exact product block matched to this NDC.
Where does this data come from?
ingredient classCode="IACT" elements from the exact product block matched by this NDC. Label-section narrative from DailyMed / the openFDA label index is shown separately when available.- FDA label on DailyMed · label index refreshed Oct 8, 2026
- FDA openFDA NDC Directory · synced Oct 8, 2026
Inactive ingredient FAQ
Are inactive ingredients the same for every manufacturer?
Why might an inactive ingredient be missing?
Can inactive ingredients matter?
Manufacturer & labeler
More NDCs from Acrotech Biopharma Inc labeler code 72893
- EVOMELA Melphalan 50 mg/10mL Injection, Powder, Lyophilized, For Solution NDC 72893-001-01
- Beleodaq Belinostat 500 mg/10mL Injection, Powder, Lyophilized, For Solution NDC 72893-002-01
- Folotyn pralatrexate 20 mg/mL Injection NDC 72893-003-01
- KHAPZORY levoleucovorin 175 mg/3.5mL Injection, Powder, Lyophilized, For Solution NDC 72893-004-01
- Folotyn pralatrexate 40 mg/2mL Injection NDC 72893-005-01
- Zevalin ibritumomab tiuxetan Kit NDC 72893-007-04
- ADQUEY Difamilast 10 mg/g Ointment NDC 72893-017-07
Where does this data come from?
- FDA openFDA NDC Directory · synced Oct 8, 2026
- Drugs@FDA
Full prescribing information FDA SPL
🎯 Indications and Usage ▾
1. INDICATIONS AND USAGE RYZNEUTA is indicated to decrease the incidence of infection, as manifested by febrile neutropenia, in adult patients with non-myeloid malignancies receiving myelosuppressive anti-cancer drugs associated with a clinically significant incidence of febrile neutropenia. Limitations of Use RYZNEUTA is not indicated for the mobilization of peripheral blood progenitor cells for hematopoietic stem cell transplantation.
RYZNEUTA is a leukocyte growth factor indicated to decrease the incidence of infection, as manifested by febrile neutropenia, in adult patients with non-myeloid malignancies receiving myelosuppressive anti-cancer drugs associated with a clinically significant incidence of febrile neutropenia. ( 1 ) Limitations of Use RYZNEUTA is not indicated for the mobilization of peripheral blood progenitor cells for hematopoietic stem cell transplantation. ( 1 )
⏱️ Dosage and Administration ▾
2. DOSAGE AND ADMINISTRATION Recommended Dose: 20 mg administered subcutaneously once per chemotherapy cycle. ( 2.1 ) Administer approximately 24 hours after cytotoxic chemotherapy. Do not administer between 14 days before and 24 hours after administration of cytotoxic chemotherapy. ( 2.1 )
2.1Recommended Dosage The recommended dosage of RYZNEUTA is a single subcutaneous injection of 20 mg administered once per chemotherapy cycle at least 24 hours after cytotoxic chemotherapy. Do not administer RYZNEUTA within 14 days before and <24 hours after administration of cytotoxic chemotherapy.
2.2Administration RYZNEUTA is administered subcutaneously via a single-dose prefilled syringe by a healthcare professional. Prior to use‚ remove the carton from the refrigerator (keeping the prefilled syringe inside the carton) for a minimum of 30 minutes to allow the product to reach room temperature. Discard any product left at room temperature for greater than 48 hours.
Parenteral drug products should be inspected visually for particulate matter and discoloration prior to administration, whenever solution and container permit. Do not administer RYZNEUTA if discoloration or particulates are observed. Caution: This product contains natural rubber latex which may cause allergic reactions.
The needle cap on the prefilled syringe contains natural rubber; people with latex allergies should not administer this product. The RYZNEUTA prefilled syringe does not bear graduation marks and is intended only to deliver the entire contents of the syringe (20 mg/mL) for direct administration to adult patients. Administer injection by pinching the skin and holding.
Inject into the abdomen, the back or side of the upper arms, or the thighs. Rotate injection sites. Do not inject into scar tissue or areas that are reddened, inflamed, or swollen.
If injecting into the abdomen, avoid a 2-inch diameter circle around the navel. Once the entire dose has been injected, the needle safety device will be triggered, pulling the needle automatically from the skin, and into the barrel; the entire needle will be covered by the needle guard.
💊 Dosage Forms and Strengths ▾
3. DOSAGE FORMS AND STRENGTHS Injection: 20 mg/mL clear, colorless, preservative-free solution in a single-dose prefilled syringe. Injection: 20 mg/mL solution in a single-dose prefilled syringe. ( 3 )
⛔ Contraindications ▾
4. CONTRAINDICATIONS RYZNEUTA is contraindicated in patients with a history of serious allergic reactions to granulocyte stimulating factors such as efbemalenograstim alfa-vuxw, pegfilgrastim, or filgrastim products [see Warnings and Precautions ( 5.3 )] . Patients with a history of serious allergic reactions to granulocyte stimulating factors such as efbemalenograstim alfa-vuxw, pegfilgrastim, or filgrastim products. ( 4 )
⚠️ Warnings and Cautions ▾
5. WARNINGS AND PRECAUTIONS Fatal splenic rupture: Evaluate patients who report left upper abdominal or shoulder pain for an enlarged spleen or splenic rupture. ( 5.1 ) Acute respiratory distress syndrome (ARDS): Evaluate patients who develop fever, lung infiltrates, or respiratory distress.
Discontinue RYZNEUTA in patients with ARDS. ( 5.2 ) Serious allergic reactions, including anaphylaxis: Permanently discontinue RYZNEUTA in patients with serious allergic reactions. ( 5.3 ) Sickle cell crises in Patients with Sickle Cell Disorders: Discontinue RYZNEUTA if sickle cell crisis occurs.
( 5.4 ) Glomerulonephritis: Evaluate and consider dose-reduction or interruption of RYZNEUTA if causality is likely. ( 5.5 ) Thrombocytopenia: Monitor platelet counts. ( 5.7 ) Capillary Leak Syndrome: Monitor patients with symptoms; consider intensive care.
( 5.8 ) Myelodysplastic Syndrome (MDS) and Acute Myeloid Leukemia (AML) in Patients with Breast and Lung Cancer: Monitor patients with breast and lung cancer using RYZNEUTA in conjunction with chemotherapy and/or radiotherapy for signs and symptoms of MDS/AML. ( 5.10 )
5.1Splenic Rupture Splenic rupture, including fatal cases, can occur following the administration of recombinant human granulocyte colony-stimulating factor (rhG-CSF) products, such as RYZNEUTA. Evaluate for an enlarged spleen or splenic rupture in patients who report left upper abdominal or shoulder pain after receiving RYZNEUTA.
5.2Acute Respiratory Distress Syndrome Acute respiratory distress syndrome (ARDS) can occur in patients receiving rhG-CSF products, such as RYZNEUTA. Evaluate patients who develop fever and lung infiltrates or respiratory distress after receiving RYZNEUTA for ARDS. Discontinue RYZNEUTA in patients with ARDS.
5.3Serious Allergic Reactions Serious allergic reactions, including anaphylaxis, can occur in patients receiving rhG-CSF products, such as RYZNEUTA. Permanently discontinue RYZNEUTA in patients with serious allergic reactions. RYZNEUTA is contraindicated in patients with a history of serious allergic reactions to RYZNEUTA or other rhG-CSF products such as pegfilgrastim, eflapegrastim or filgrastim products.
5.4Sickle Cell Crisis in Patients with Sickle Cell Disorders Severe and sometimes fatal sickle cell crises can occur in patients with sickle cell disorders receiving rhG-CSF products, such as RYZNEUTA. Discontinue RYZNEUTA if sickle cell crisis occurs.
5.5Glomerulonephritis Glomerulonephritis has occurred in patients receiving rhG-CSF products. The diagnoses were based upon azotemia, hematuria (microscopic and macroscopic), proteinuria, and renal biopsy. Generally, events of glomerulonephritis resolved after dose-reduction or discontinuation of rhG-CSF. If glomerulonephritis is suspected, evaluate for cause. If causality is likely, consider dose-reduction or interruption of RYZNEUTA.
5.6Leukocytosis White blood cell (WBC) counts of 100 × 10 9 /L or greater have been observed in patients receiving rhG-CSF products. Monitor complete blood count (CBC) during RYZNEUTA therapy. Discontinue RYZNEUTA treatment if WBC count of 100 × 10 9 /L or greater occurs.
5.7Thrombocytopenia Thrombocytopenia has been reported in patients receiving rhG-CSF products. Thrombocytopenia occurred in 11% of RYZNEUTA-treated patients. One patient (0.4%) experienced severe thrombocytopenia. Monitor platelet counts.
5.8Capillary Leak Syndrome Capillary leak syndrome has been reported after administration of rhG-CSF products and is characterized by hypotension, hypoalbuminemia, edema and hemoconcentration. Episodes vary in frequency and severity, and may be life-threatening if treatment is delayed. Patients who develop symptoms of capillary leak syndrome should be closely monitored and receive standard symptomatic treatment, which may include a need for intensive care.
5.9Potential for Tumor Growth Stimulatory Effects on Malignant Cells The granulocyte colony-stimulating factor (G-CSF) receptor through which RYZNEUTA… [Excerpted — this section continues on DailyMed.]
⚠️ Warnings ▾
5.1Splenic Rupture Splenic rupture, including fatal cases, can occur following the administration of recombinant human granulocyte colony-stimulating factor (rhG-CSF) products, such as RYZNEUTA. Evaluate for an enlarged spleen or splenic rupture in patients who report left upper abdominal or shoulder pain after receiving RYZNEUTA.
5.2Acute Respiratory Distress Syndrome Acute respiratory distress syndrome (ARDS) can occur in patients receiving rhG-CSF products, such as RYZNEUTA. Evaluate patients who develop fever and lung infiltrates or respiratory distress after receiving RYZNEUTA for ARDS. Discontinue RYZNEUTA in patients with ARDS.
5.3Serious Allergic Reactions Serious allergic reactions, including anaphylaxis, can occur in patients receiving rhG-CSF products, such as RYZNEUTA. Permanently discontinue RYZNEUTA in patients with serious allergic reactions. RYZNEUTA is contraindicated in patients with a history of serious allergic reactions to RYZNEUTA or other rhG-CSF products such as pegfilgrastim, eflapegrastim or filgrastim products.
5.4Sickle Cell Crisis in Patients with Sickle Cell Disorders Severe and sometimes fatal sickle cell crises can occur in patients with sickle cell disorders receiving rhG-CSF products, such as RYZNEUTA. Discontinue RYZNEUTA if sickle cell crisis occurs.
5.5Glomerulonephritis Glomerulonephritis has occurred in patients receiving rhG-CSF products. The diagnoses were based upon azotemia, hematuria (microscopic and macroscopic), proteinuria, and renal biopsy. Generally, events of glomerulonephritis resolved after dose-reduction or discontinuation of rhG-CSF. If glomerulonephritis is suspected, evaluate for cause. If causality is likely, consider dose-reduction or interruption of RYZNEUTA.
5.6Leukocytosis White blood cell (WBC) counts of 100 × 10 9 /L or greater have been observed in patients receiving rhG-CSF products. Monitor complete blood count (CBC) during RYZNEUTA therapy. Discontinue RYZNEUTA treatment if WBC count of 100 × 10 9 /L or greater occurs.
5.7Thrombocytopenia Thrombocytopenia has been reported in patients receiving rhG-CSF products. Thrombocytopenia occurred in 11% of RYZNEUTA-treated patients. One patient (0.4%) experienced severe thrombocytopenia. Monitor platelet counts.
5.8Capillary Leak Syndrome Capillary leak syndrome has been reported after administration of rhG-CSF products and is characterized by hypotension, hypoalbuminemia, edema and hemoconcentration. Episodes vary in frequency and severity, and may be life-threatening if treatment is delayed. Patients who develop symptoms of capillary leak syndrome should be closely monitored and receive standard symptomatic treatment, which may include a need for intensive care.
5.9Potential for Tumor Growth Stimulatory Effects on Malignant Cells The granulocyte colony-stimulating factor (G-CSF) receptor through which RYZNEUTA acts has been found on tumor cell lines. The possibility that RYZNEUTA acts as a growth factor for any tumor type, including myeloid malignancies and myelodysplasia, diseases for which RYZNEUTA is not approved, cannot be excluded.
5.10Myelodysplastic Syndrome (MDS) and Acute Myeloid Leukemia (AML) in Patients with Breast and Lung Cancer MDS and AML have been associated with the use of rhG-CSF products in conjunction with chemotherapy and/or radiotherapy in patients with breast and lung cancer. Monitor patients for signs and symptoms of MDS/AML in these settings.
5.11Aortitis Aortitis has been reported in patients receiving rhG-CSF products. It may occur as early as the first week after start of therapy. Manifestations may include generalized signs and symptoms such as fever, abdominal pain, malaise, back pain, and increased inflammatory markers (e.g., c-reactive protein and white blood cell count).
Consider aortitis in patients who develop these signs and symptoms without known etiology. Discontinue RYZNEUTA if aortitis is suspected.
5.12Nuclear Imaging Increased hematopoietic activity of… [Excerpted — this section continues on DailyMed.]
🤒 Adverse Reactions ▾
6. ADVERSE REACTIONS The following clinically significant adverse reactions are discussed in greater detail in other sections of the labeling: Splenic Rupture [see Warnings and Precautions ( 5.1 )] Acute Respiratory Distress Syndrome [see Warnings and Precautions ( 5.2 )] Serious Allergic Reactions [see Warnings and Precautions ( 5.3 )] Sickle Cell Crisis in Patients with Sickle Cell Disorders [see Warnings and Precautions ( 5.4 )] Glomerulonephritis [see Warnings and Precautions ( 5.5 )] Leukocytosis [see Warnings and Precautions ( 5.6 )] Thrombocytopenia [see Warnings and Precautions ( 5.7 )] Capillary Leak Syndrome [see Warnings and Precautions ( 5.8 )] Potential for Tumor Growth Stimulatory Effects on Malignant Cells [see Warnings and Precautions ( 5.9 )] Myelodysplastic Syndrome (MDS) and Acute Myeloid Leukemia (AML) in Patients with Breast and Lung Cancer [see Warnings and Precautions ( 5.10 )] Aortitis [see Warnings and Precautions ( 5.11 )] Most common adverse reactions (≥10%) were nausea, anemia, and thrombocytopenia.
( 6.1 ) To report SUSPECTED ADVERSE REACTIONS, contact Acrotech Biopharma Inc at 1-888-292-9617 or FDA at 1-800-FDA-1088 or www.fda.gov/medwatch.
6.1Clinical Trials Experience Because clinical trials are conducted under widely varying conditions, adverse reaction rates observed in the clinical trials of a drug cannot be directly compared with rates in the clinical trials of another drug and may not reflect the rates observed in clinical practice. The following adverse reaction data are based on two studies [see Clinical Studies ( 14 )]. The first was a randomized, double-blind, placebo-controlled study in patients with metastatic or non-metastatic breast cancer receiving doxorubicin 60 mg/m 2 and docetaxel 75 mg/m 2 every 21 days (Study GC-627-04).
A total of 122 female patients were randomized to receive either 20 mg RYZNEUTA (n=83) or placebo (n=39) in chemotherapy cycle 1; all patients received RYZNEUTA in cycles 2-4. The second was a randomized, open-label, active-controlled study in patients with stage I to III invasive breast cancer receiving docetaxel 75 mg/m 2 and cyclophosphamide 600 mg/m 2 (Study GC-627-05). A total of 393 patients were randomized to receive either 20 mg RYZNEUTA (n=197) or pegfilgrastim (n=196) in chemotherapy cycles 1 through 4.
In Study GC-627-04, the most common adverse reactions (≥10%) in the RYZNEUTA arm through cycle 1 were nausea, anemia, and thrombocytopenia (see Table 1). Other adverse reactions reported by ≥ 20% of RYZNEUTA-treated Patients with Breast Cancer Receiving Myelosuppressive Chemotherapy in Study GC-627-05 were fatigue and bone pain. Table 1.
Adverse Reactions Adverse reactions that occurred in ≥10% of Ryzneuta-treated patients and ≥5% more than placebo-treated patients. in Study GC-627-04 in RYZNEUTA-treated Patients with Breast Cancer Receiving Myelosuppressive Chemotherapy Through Cycle 1 Adverse Reactions Ryzneuta (n=83) Placebo (n=39) Nausea 42 (51) 15 (39) Anemia 12 (15) 4 (10) Thrombocytopenia 10 (12) 1 (3)
6.1Clinical Trials Experience Because clinical trials are conducted under widely varying conditions, adverse reaction rates observed in the clinical trials of a drug cannot be directly compared with rates in the clinical trials of another drug and may not reflect the rates observed in clinical practice. The following adverse reaction data are based on two studies [see Clinical Studies ( 14 )]. The first was a randomized, double-blind, placebo-controlled study in patients with metastatic or non-metastatic breast cancer receiving doxorubicin 60 mg/m 2 and docetaxel 75 mg/m 2 every 21 days (Study GC-627-04).
A total of 122 female patients were randomized to receive either 20 mg RYZNEUTA (n=83) or placebo (n=39) in chemotherapy cycle 1; all patients received RYZNEUTA in cycles 2-4. The second was a randomized, open-label, active-controlled study in patients with stage I to III invasive breast cancer receiving docetaxel 75 mg/m 2 and cyclophosphami… [Excerpted — this section continues on DailyMed.]
👥 Use in Specific Populations ▾
8. USE IN SPECIFIC POPULATIONS
8.1Pregnancy Risk Summary Although available data with RYZNEUTA use in pregnant women are insufficient to establish whether there is a drug associated risk of major birth defects, miscarriage, or adverse maternal or fetal outcomes, there are available data from published studies in pregnant women exposed to other human G-CSF products. These studies have not established an association of G-CSF product use during pregnancy with major birth defects, miscarriage, or adverse maternal or fetal outcomes. In animal studies, no evidence of reproductive/developmental toxicity occurred in the offspring of pregnant rats and rabbits that received cumulative doses of efbemalenograstim alfa-vuxw approximately 2.6 and 0.7 times, respectively, the recommended human dose (based on body surface area).
The estimated background risk of major birth defects and miscarriage for the indicated population is unknown. All pregnancies have a background risk of birth defect, loss, or other adverse outcomes. In the U.S. general population, the estimated background risks of major birth defects and miscarriage in clinically recognized pregnancies is 2-4% and 15-20%, respectively.
Data Animal Data Three studies were conducted in pregnant rats dosed with efbemalenograstim alfa-vuxw at cumulative doses up to approximately 2.6 times the recommended human dose at the following stages of gestation: during the period of organogenesis, from mating through the first half of pregnancy, and from the first trimester through delivery and lactation. No evidence of fetal loss or structural malformations was observed in any study. Growth and learning were also unaffected.
Pregnant rabbits were dosed with efbemalenograstim alfa-vuxw every other day during organogenesis at cumulative doses up to 0.7 times the recommended human dose showed no signs of fetal loss or structural malformations. Maternal toxicity (decreased weight gain or weight loss and/or death) was observed when higher cumulative doses of efbemalenograstim alfa-vuxw were used in an early dose-ranging study in rabbits.
8.2Lactation Risk Summary There are no data on the presence of efbemalenograstim alfa-vuxw or its metabolite in either human or animal milk, the effects on the breastfed child, or the effects on milk production. The developmental and health benefits of breastfeeding should be considered along with the mother’s clinical need for efbemalenograstim alfa-vuxw and any potential adverse effects on the breastfed child from efbemalenograstim alfa-vuxw or from the underlying maternal condition.
8.4Pediatric Use Safety and effectiveness in pediatric patients have not been established.
8.5Geriatric Use Clinical studies of RYZNEUTA did not include sufficient numbers of subjects aged 65 and over to determine whether they respond differently from younger subjects. Other reported clinical experience with RYZNEUTA has not identified differences in responses between the elderly and younger patients.
🤰 Pregnancy ▾
8.1Pregnancy Risk Summary Although available data with RYZNEUTA use in pregnant women are insufficient to establish whether there is a drug associated risk of major birth defects, miscarriage, or adverse maternal or fetal outcomes, there are available data from published studies in pregnant women exposed to other human G-CSF products. These studies have not established an association of G-CSF product use during pregnancy with major birth defects, miscarriage, or adverse maternal or fetal outcomes. In animal studies, no evidence of reproductive/developmental toxicity occurred in the offspring of pregnant rats and rabbits that received cumulative doses of efbemalenograstim alfa-vuxw approximately 2.6 and 0.7 times, respectively, the recommended human dose (based on body surface area).
The estimated background risk of major birth defects and miscarriage for the indicated population is unknown. All pregnancies have a background risk of birth defect, loss, or other adverse outcomes. In the U.S. general population, the estimated background risks of major birth defects and miscarriage in clinically recognized pregnancies is 2-4% and 15-20%, respectively.
Data Animal Data Three studies were conducted in pregnant rats dosed with efbemalenograstim alfa-vuxw at cumulative doses up to approximately 2.6 times the recommended human dose at the following stages of gestation: during the period of organogenesis, from mating through the first half of pregnancy, and from the first trimester through delivery and lactation. No evidence of fetal loss or structural malformations was observed in any study. Growth and learning were also unaffected.
Pregnant rabbits were dosed with efbemalenograstim alfa-vuxw every other day during organogenesis at cumulative doses up to 0.7 times the recommended human dose showed no signs of fetal loss or structural malformations. Maternal toxicity (decreased weight gain or weight loss and/or death) was observed when higher cumulative doses of efbemalenograstim alfa-vuxw were used in an early dose-ranging study in rabbits.
🧒 Pediatric Use ▾
8.4Pediatric Use Safety and effectiveness in pediatric patients have not been established.
🧓 Geriatric Use ▾
8.5Geriatric Use Clinical studies of RYZNEUTA did not include sufficient numbers of subjects aged 65 and over to determine whether they respond differently from younger subjects. Other reported clinical experience with RYZNEUTA has not identified differences in responses between the elderly and younger patients.
🆘 Overdosage ▾
10. OVERDOSAGE Overdosage of RYZNEUTA may result in leukocytosis and bone pain. In the event of overdose, general supportive measures should be instituted, as necessary. Monitor the patient for adverse reactions [see Adverse Reactions ( 6 )] .
🧬 Clinical Pharmacology ▾
12. CLINICAL PHARMACOLOGY
12.1Mechanism of Action Efbemalenograstim alfa-vuxw is a colony-stimulating factor that acts on hematopoietic cells by binding to specific cell surface receptors, thereby stimulating proliferation, differentiation, commitment, and end cell functional activation.
12.2Pharmacodynamics Over the tested dose range of 30 to 360 μg/kg in healthy adult males, neutrophils generally increased in a dose-dependent manner; however, the effect on neutrophils plateaued at the top two doses of 240 and 360 μg/kg. In patients with breast cancer given EC chemotherapy, in cycle 1, median ANC peaked on Day 4 (about 24 hours after efbemalenograstim alfa-vuxw administration) for the 240 and 320 μg/kg doses, declined to nadir on Days 8–9, and then recovered to a count of 2.0 × 10 9 /L and higher by Day 11.
The ANC levels in cycles 2, 3, and 4 tended to be higher than those observed in cycle 1. In patients with breast cancer given TAC chemotherapy, in cycle 1, median ANC peaked on Day 3 (about 24 hours after efbemalenograstim alfa-vuxw administration) in cycle 1 and 3, reached nadir on Day 8, and recovered to 2.0 x 10 9 /L on Days 9 and 10 for the 320 μg/kg and 240 μg/kg doses, respectively. Mean ANC nadir was higher with 320 μg/kg compared to 240 μg/kg in both cycle 1 (0.75 x 10 9 /L and 0.29 x 10 9 /L, respectively), and cycle 3 (1.19 x 10 9 /L and 0.57 x 10 9 /L, respectively).
12.3Pharmacokinetics The pharmacokinetics of efbemalenograstim alfa-vuxw was studied in female patients with breast cancer and healthy male subjects. Efbemalenograstim alfa-vuxw exhibited nonlinear and time-dependent pharmacokinetics over the dose range of 30 to 360 μg/kg. At the approved recommended dose in healthy subjects, the mean (%CV) C max was 1202 ng/mL (56%), and the AUC 0-inf was 76357 h*ng/mL (65%).
At the approved recommended dose in patients with breast cancer, the geometric mean (CV%) Cmax was 1085 ng/mL (92%) in Cycle 1 and 525 ng/mL (163%) in Cycle 3; the geometric mean (CV%) AUC 0-inf was 54858 h*ng/mL (110%) in Cycle 1 and 26217 h*ng/mL (167%) in Cycle 3. Absorption The median t max of efbemalenograstim alfa-vuxw administered as 80 to 320 μg/kg in female patients with breast cancer receiving EC chemotherapy ranged from 24 hours to 48 hours in Cycle 1 and 9 to 30 hours in Cycle 3. The median t max of efbemalenograstim alfa-vuxw administered as 240 to 320 μg/kg in female participants with breast cancer receiving TAC chemotherapy was 36 hours in Cycle 1 and ranged from 24 to 30 hours in Cycle 3.
Distribution The geometric mean (CV%) apparent volume of distribution of efbemalenograstim alfa-vuxw was
18.8 L (257%) in Cycle 1 and
40.7L (387%) in Cycle 3 in female patients with breast cancer. Elimination The geometric mean (CV%) apparent clearance of efbemalenograstim alfa-vuxw was
0.36 L/h (110%) in Cycle 1 and
0.76L/h (167%) in Cycle 3. The geometric mean (CV%) elimination half-life of efbemalenograstim alfa-vuxw was 35.6 h (108%) in Cycle 1 and 36.9 h (120%) in Cycle 3. Neutrophil receptor binding is an important component of the clearance of efbemalenograstim alfa-vuxw, and serum clearance is directly related to the number of neutrophils.
Metabolism Efbemalenograstim alfa-vuxw is expected to be metabolized into small peptides by catabolic pathways. Specific Populations No clinically significant differences were observed based on age (20 to 83 years) or body weight (40 to 137 kg) [see Use in Specific Populations ( 8.5 )]. The impact of sex, race/ethnicity, renal impairment, hepatic impairment, and pregnancy on the pharmacokinetics of efbemalenograstim alfa-vuxw are unknown.
12.6Immunogenicity The observed incidence of anti-drug antibodies is highly dependent on the sensitivity and specificity of the assay. Differences in assay methods preclude meaningful comparisons of the incidence of anti-drug antibodies in the studies described below with the incidence of anti-drug antibodies in other studies, including those of efbemalenogras… [Excerpted — this section continues on DailyMed.]
🧬 Mechanism of Action ▾
12.1Mechanism of Action Efbemalenograstim alfa-vuxw is a colony-stimulating factor that acts on hematopoietic cells by binding to specific cell surface receptors, thereby stimulating proliferation, differentiation, commitment, and end cell functional activation.
📦 How Supplied / Storage and Handling ▾
16. HOW SUPPLIED/STORAGE AND HANDLING RYZNEUTA (efbemalenograstim alfa-vuxw) injection is a clear, colorless, preservative-free solution supplied in a prefilled single-dose syringe with a 27-gauge, 1/2-inch needle and an UltraSafe Passive™ Needle Guard, containing 20 mg of efbemalenograstim alfa-vuxw. Caution: This product contains natural rubber latex which may cause allergic reactions.
The needle cap of the prefilled syringe contains natural rubber; persons with latex allergies should not administer this product. RYZNEUTA is provided in a dispensing pack containing one 20 mg/mL prefilled syringe (NDC 72893-016-02). RYZNEUTA prefilled syringe does not bear graduation marks and is intended only to deliver the entire contents of the syringe (20 mg/mL) for direct administration to adult patients.
Store refrigerated at 2°C to 8°C (36°F to 46°F) in the carton to protect from light. Do not shake. Discard syringes stored at room temperature for more than 48 hours.
Do not freeze. Discard syringe if frozen.
📋 Description ▾
11. DESCRIPTION Efbemalenograstim alfa-vuxw, a leukocyte growth factor, is a 413 amino acid recombinant fusion protein consisting of human G-CSF, a 16 amino-acid linker, and the Fc portion of human IgG2. In solution, efbemalenograstim alfa-vuxw forms covalently-linked dimers (disulfide bonds between Fc moieties), resulting in an immunoglobulin-like structure.
The dimer is a water-soluble, glycosylated protein with a molecular weight of approximately 93.4 kilodaltons (kDa), of which 89.5 kDa is attributed to amino acids (protein sequence) and the remainder is from glycosylation. Efbemalenograstim alfa-vuxw is obtained from genetically-engineered strain of Chinese hamster ovary (CHO) cells grown in a serum-free medium. RYZNEUTA (efbemalenograstim alfa-vuxw) injection is supplied in 1 mL prefilled single-dose syringes for manual subcutaneous injection.
The prefilled syringe does not bear graduation marks and is designed to deliver the entire contents of the syringe (20 mg/mL). Each syringe contains 20 mg efbemalenograstim alfa-vuxw in a sterile, clear, colorless, preservative-free solution (pH 5.2) containing acetate (0.6 mg), EDTA (0.29 mg), polysorbate 20 (0.1 mg), sodium (0.23 mg), and sorbitol (50 mg) in Water for Injection, USP.
🧬 Pharmacokinetics ▾
12.3Pharmacokinetics The pharmacokinetics of efbemalenograstim alfa-vuxw was studied in female patients with breast cancer and healthy male subjects. Efbemalenograstim alfa-vuxw exhibited nonlinear and time-dependent pharmacokinetics over the dose range of 30 to 360 μg/kg. At the approved recommended dose in healthy subjects, the mean (%CV) C max was 1202 ng/mL (56%), and the AUC 0-inf was 76357 h*ng/mL (65%).
At the approved recommended dose in patients with breast cancer, the geometric mean (CV%) Cmax was 1085 ng/mL (92%) in Cycle 1 and 525 ng/mL (163%) in Cycle 3; the geometric mean (CV%) AUC 0-inf was 54858 h*ng/mL (110%) in Cycle 1 and 26217 h*ng/mL (167%) in Cycle 3. Absorption The median t max of efbemalenograstim alfa-vuxw administered as 80 to 320 μg/kg in female patients with breast cancer receiving EC chemotherapy ranged from 24 hours to 48 hours in Cycle 1 and 9 to 30 hours in Cycle 3. The median t max of efbemalenograstim alfa-vuxw administered as 240 to 320 μg/kg in female participants with breast cancer receiving TAC chemotherapy was 36 hours in Cycle 1 and ranged from 24 to 30 hours in Cycle 3.
Distribution The geometric mean (CV%) apparent volume of distribution of efbemalenograstim alfa-vuxw was
18.8 L (257%) in Cycle 1 and
40.7L (387%) in Cycle 3 in female patients with breast cancer. Elimination The geometric mean (CV%) apparent clearance of efbemalenograstim alfa-vuxw was
0.36 L/h (110%) in Cycle 1 and
0.76L/h (167%) in Cycle 3. The geometric mean (CV%) elimination half-life of efbemalenograstim alfa-vuxw was 35.6 h (108%) in Cycle 1 and 36.9 h (120%) in Cycle 3. Neutrophil receptor binding is an important component of the clearance of efbemalenograstim alfa-vuxw, and serum clearance is directly related to the number of neutrophils.
Metabolism Efbemalenograstim alfa-vuxw is expected to be metabolized into small peptides by catabolic pathways. Specific Populations No clinically significant differences were observed based on age (20 to 83 years) or body weight (40 to 137 kg) [see Use in Specific Populations ( 8.5 )]. The impact of sex, race/ethnicity, renal impairment, hepatic impairment, and pregnancy on the pharmacokinetics of efbemalenograstim alfa-vuxw are unknown.
🧬 Pharmacodynamics ▾
12.2Pharmacodynamics Over the tested dose range of 30 to 360 μg/kg in healthy adult males, neutrophils generally increased in a dose-dependent manner; however, the effect on neutrophils plateaued at the top two doses of 240 and 360 μg/kg. In patients with breast cancer given EC chemotherapy, in cycle 1, median ANC peaked on Day 4 (about 24 hours after efbemalenograstim alfa-vuxw administration) for the 240 and 320 μg/kg doses, declined to nadir on Days 8–9, and then recovered to a count of 2.0 × 10 9 /L and higher by Day 11.
The ANC levels in cycles 2, 3, and 4 tended to be higher than those observed in cycle 1. In patients with breast cancer given TAC chemotherapy, in cycle 1, median ANC peaked on Day 3 (about 24 hours after efbemalenograstim alfa-vuxw administration) in cycle 1 and 3, reached nadir on Day 8, and recovered to 2.0 x 10 9 /L on Days 9 and 10 for the 320 μg/kg and 240 μg/kg doses, respectively. Mean ANC nadir was higher with 320 μg/kg compared to 240 μg/kg in both cycle 1 (0.75 x 10 9 /L and 0.29 x 10 9 /L, respectively), and cycle 3 (1.19 x 10 9 /L and 0.57 x 10 9 /L, respectively).
🔬 Clinical Studies ▾
14. CLINICAL STUDIES The efficacy of RYZNEUTA to decrease the incidence of infection, as manifested by febrile neutropenia, in patients with nonmyeloid malignancies receiving myelosuppressive anti-cancer drugs was evaluated in two randomized, controlled studies. Study GC-627-04 [NCT02872103] was a randomized, double-blind, placebo-controlled study that employed doxorubicin 60 mg/m 2 and docetaxel 75 mg/m 2 administered every 21 days for up to 4 cycles for the treatment of metastatic or non-metastatic breast cancer.
In this study, 122 patients were randomized to receive a single subcutaneous injection of RYZNEUTA (20 mg) or placebo on day 2 of chemotherapy cycle 1. All patients received RYZNEUTA (20 mg) on day 2 of chemotherapy cycles 2 – 4. The patients were 30 to 69 years of age and all female.
The race was 99% Caucasian, and 1% Black. The ethnicity was 2% Hispanic or Latino. Efficacy was based upon the mean duration of severe (Grade 4) neutropenia in cycle 1 which was lower for RYZNEUTA-treated patients as compared to placebo-treated patients (LS mean: 1.4 days versus 4.3 days, respectively, p<0.001 [95% CI: 2.4, 3.5]).
The incidence of febrile neutropenia (defined as temperature ≥38.3℃, or >38.0℃ sustained for at least 1 hour, and ANC < 0.5 × 10 9 /L) was also lower for RYZNEUTA-treated patients compared to placebo-treated patients in cycle 1 (4.8% versus 25.6%, respectively, p=0.0016; 20.8% difference [95% CI: 1.8%, 38.8%]. Study GC-627-05 [NCT03252431] was a randomized, active-controlled study that compared RYZNEUTA to pegfilgrastim. Study GC-627-05 employed docetaxel 75 mg/m 2 and cyclophosphamide 600 mg/m 2 administered every 21 days for up to 4 cycles for the treatment of non-metastatic breast cancer.
In this study, 393 patients were randomized to receive a single subcutaneous injection of RYZNEUTA (20 mg) or pegfilgrastim (6 mg) on day 2 of each chemotherapy cycle. The patients were 26 to 83 years of age and all female and Caucasian. The ethnicity was 0.3% Hispanic or Latino.
The study demonstrated that the mean days of severe (Grade 4) neutropenia of RYZNEUTA-treated patients did not exceed that of pegfilgrastim-treated patients by more than 0.6 days in cycle 1 of chemotherapy. The mean days of severe neutropenia in cycle 1 were 0.2 days in both the RYZNEUTA and pegfilgrastim arms (difference in means 0.0 days [95% CI -0.1, 0.1]). Subgroup analyses of treatment effect were not useful due to small subgroup sizes.
🧪 Nonclinical Toxicology ▾
13. NONCLINICAL TOXICOLOGY
13.1Carcinogenesis, Mutagenesis, Impairment of Fertility No carcinogenicity or mutagenesis studies have been conducted with efbemalenograstim alfa-vuxw. Efbemalenograstim alfa did not affect reproductive performance or fertility in male or female rats at cumulative weekly doses approximately 2.2 times higher than the recommended human dose (based on body surface area).
📄 Carcinogenesis, Mutagenesis, Impairment of Fertility ▾
13.1Carcinogenesis, Mutagenesis, Impairment of Fertility No carcinogenicity or mutagenesis studies have been conducted with efbemalenograstim alfa-vuxw. Efbemalenograstim alfa did not affect reproductive performance or fertility in male or female rats at cumulative weekly doses approximately 2.2 times higher than the recommended human dose (based on body surface area).
📄 Patient Package Insert ▾
Patient Information RYZNEUTA® (rīz-new-ta) (efbemalenograstim alfa-vuxw) injection Single-Dose Prefilled Syringe What is RYZNEUTA? RYZNEUTA is a is a man-made form of granulocyte colony-stimulating factor (G-CSF). G-CSF is a substance produced by the body.
It stimulates the growth of neutrophils, a type of white blood cell important in the body’s fight against infection. RYZNEUTA is used in adults. It is not known if RYZNEUTA is safe and effective in children.
Do not take RYZNEUTA if you have had a serious allergic reaction to granulocyte stimulating factors such as efbemalenograstim alfa-vuxw, pegfilgrastim products or filgrastim products. Before you receive RYZNEUTA, tell your healthcare provider about all of your medical conditions, including if you: have a sickle cell disorder. have kidney problems. are allergic to latex. The needle cap on the prefilled syringe contains dry natural rubber (derived from latex). are pregnant or plan to become pregnant.
It is not known if RYZNEUTA will harm your unborn baby. are breastfeeding or plan to breastfeed. It is not known if RYZNEUTA passes into your breast milk. Tell your healthcare provider about all of the medicines you take, including prescription and over-the-counter medicines, vitamins, and herbal supplements.
How will I receive RYZNEUTA? RYZNEUTA is given as an injection under your skin (subcutaneous injection) by a healthcare provider. RYZNEUTA should be injected at least 24 hours after your dose of chemotherapy and at least 14 days before your next dose of chemotherapy.
If you miss a dose of RYZNEUTA, talk to your healthcare provider about when you should receive your next dose. What are possible side effects of RYZNEUTA? RYZNEUTA may cause serious side effects, including: Spleen rupture.
Your spleen may become enlarged and can rupture. A ruptured spleen can cause death. Call your healthcare provider right away if you have pain in the left upper stomach area or your left shoulder.
A serious lung problem called Acute Respiratory Distress Syndrome (ARDS). Call your healthcare provider or get emergency care right away if you have shortness of breath with or without a fever, trouble breathing, or a fast rate of breathing. Serious allergic reactions.RYZNEUTA can cause serious allergic reactions.
These reactions can cause a rash over your whole body, swelling around your mouth or eyes, wheezing, shortness of breath, dizziness, and fast heart rate. If you have any of these symptoms, stop using RYZNEUTA and call your healthcare provider or get emergency medical help right away. Risk of Sickle cell crises.
You may have a serious sickle cell crisis, which could lead to death, if you have a sickle cell disorder and receive RYZNEUTA. Call your healthcare provider right away if you have symptoms of sickle cell crisis such as pain or difficulty breathing. Kidney injury (glomerulonephritis).
Products like RYZNEUTA can cause kidney injury. Call your healthcare provider right away if you develop any of the following symptoms: swelling of your face or ankles dark colored urine or blood in your urine you urinate less than usual Increased white blood cell count (leukocytosis). Your healthcare provider will check your blood during treatment with RYZNEUTA.
Decreased platelet count (thrombocytopenia). Your healthcare provider will check your blood during treatment with RYZNEUTA. Capillary Leak Syndrome.
RYZNEUTA can cause fluid to leak from blood vessels into your body’s tissues. This condition is called “Capillary Leak Syndrome” (CLS). CLS can quickly cause you to have symptoms that may become life-threatening.
Get emergency medical help right away if you develop any of the following symptoms: swelling or puffiness and urinating less than usual trouble breathing swelling of your stomach area (abdomen) and feeling of fullness dizziness or feeling faint a general feeling of tiredness Myelodysplastic Syndrome (MDS) and Acute Myeloid Leukemia (AML). If you have breast cancer or lung cancer, when RYZN… [Excerpted — this section continues on DailyMed.]
📄 Package Label / Principal Display Panel ▾
Syringe Label RYZNEUTA (efbemalenograstim alfa-vuxw) Injection 20 mg/mL For Subcutaneous Use Only Single-dose Prefilled Syringe Mfr By: EVIVE BIOTECHNOLOGY SINGAPORE PTE. LTD Dist.By: Acrotech Biopharma Inc, East Windsor, NJ 08520 USA Syringe Label
Carton Label RYZNEUTA (efbemalenograstim alfa-vuxw) Injection 20 mg/mL Store refrigerated at 2° to 8°C (36° to 46°F) in the carton to protect from light. Do Not Freeze or Shake. Sterile Solution - No preservatives RYZNEUTA NDC 72893-016-02 (efbemalenograstim alfa-vuxw) Rx Only Injection 20 mg/mL Recombinant dimeric human granulocyte colony-stimulating factor Fc fusion protein derived from mammalian cell culture For Subcutaneous Injection by a Healthcare Provider Only Contains One Single-Dose Prefilled Syringe Dosage: See Prescribing Information Manufactured By: EVIVE BIOTECHNOLOGY SINGAPORE PTE.
LTD. Singapore, 189720 Distributed by: Acrotech Biopharma Inc, East Windsor, NJ 08520 USA US License No. 2248 Product of China RYZNEUTA NDC 72893-016-02 (efbemalenograstim alfa-vuxw) Injection Rx Only 20 mg/mL For Subcutaneous Injection by a Healthcare Provider Only Each 1 mL prefilled syringe contains: 20 mg efbemalenograstim alfa-vuxw in a sterile, clear, colorless solution (pH 5.2) containing acetate (0.6 mg), EDTA (0.29 mg), polysorbate 20 (0.1 mg), sodium (0.23 mg), and sorbitol (50 mg) in water for injection, USP.
Caution: The Packaging of This Product Contains Natural Rubber Latex Which May Cause Allergic Reactions. Carton Label
Tray Lid Label RYZNEUTA (efbemalenograstim alfa-vuxw) Injection 20 mg/mL For Subcutaneous Use Only Recombinant dimeric human granulocyte colony-stimulating factor Fc fusion protein derived from mammalian cell culture One Single-Dose Prefilled Syringe Rx Only NDC 72893-016-02 Each 1 mL prefilled syringe contains: 20 mg efbemalenograstim alfa-vuxw in a sterile, clear, colorless solution (pH 5.2) containing acetate (0.6 mg), EDTA (0.29 mg), polysorbate 20 (0.1 mg), sodium (0.23 mg), and sorbitol (50 mg) in water for injection, USP Dosage: See Prescribing Information Store refrigerated at 2° to 8°C (36° to 46°F) in the carton to protect from light.
Do Not Freeze or Shake. Sterile Solution – No Preservatives Caution: The Packaging of This Product Contains Natural Rubber Latex Which May Cause Allergic Reactions. Mfr.
By: EVIVE BIOTECHNOLOGY SINGAPORE PTE. LTD. Dist.
By: Acrotech Biopharma Inc, East Windsor, NJ 08520 USA US LIC No. 2248 Tray Lid Label
Reported adverse events (FAERS)
Top reported reactions
Reporter sex
Where does this data come from?
About this NDC listing & data coverage
What data is (and isn’t) available for this NDC — tap to expand
| NDC identity (package / product / labeler codes) | ✓ Available |
| Labeler | ✓ Available |
| Product & package description | ✓ Available |
| Marketing category & status | ✓ Available |
| Active ingredient / dosage form / route | ✓ Available |
| FDA label (SPL via DailyMed) | ✓ Available |
| Package photos | ✓ Available |
| Inactive ingredients (structured) | ✓ Available |
| NADAC pharmacy acquisition price (CMS) | — Not published for this NDC CMS publishes NADAC only for NDCs reported in its retail-pharmacy survey. |
| Orange Book / therapeutic-equivalence data | ✓ Available |
| HCPCS J-code billing crosswalk | — Not published for this NDC Most self-administered / retail products have no J-code — that is normal. |
| Medicaid utilization (CMS SDUD) | — Not published for this NDC CMS reports utilization only for NDCs with Medicaid claims above its privacy threshold. |