
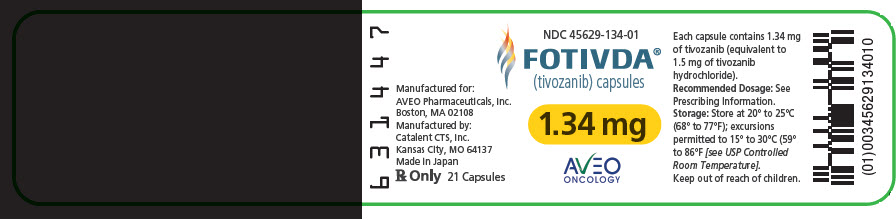
FOTIVDA Tivozanib 1.34 mg Capsule, 21-count — NDC 45629-134-02 (Billing 45629-0134-02)
This is a package of 21 capsules of FOTIVDA Tivozanib 1.34 mg Capsule from AVEO Pharmaceuticals, Inc., marketed since Mar 2021 and currently FDA-listed.
Identity & classification
Regulatory identifiers FDA, NLM and CMS codes for this package
Drug-database identifiers Medi-Span GPI and First Databank GCN / HICL / AHFS classification
- GSN (GCN sequence number): 079735
- GCN: 46287
- HICL (First Databank): 045740
- AHFS class code: 10:00.00.00
Where does this data come from?
- FDA openFDA NDC Directory · synced Oct 1, 2026
- FDA label on DailyMed · label index refreshed Oct 1, 2026
- RxNorm (NLM RxNav) · catalog refreshed Oct 1, 2026
- Medi-Span GPI (licensed)
- First Databank (licensed) · refreshed Oct 1, 2026
Clinical
- Fotivda (tivozanib) treats adults with advanced kidney cancer, called renal cell carcinoma, that has come back or stopped responding. It is used after at least two earlier treatmen...
- Take one capsule by mouth once daily, with or without food. You take it for 21 days and then stop for 7 days, repeating in 28-day cycles. Swallow it whole with water and never open...
- Common ones are tiredness, high blood pressure, diarrhea, less appetite, nausea, a hoarse voice, an underactive thyroid, cough and mouth sores. Tell us if they bother you. Your doc...
- Call for chest pain, signs of a stroke, severe headache, seizures, vision changes, confusion, unusual bleeding, severe belly pain, or swelling and shortness of breath. These can be...
Patient education
Supplement & herbal interactions
Some supplements/herbs that may interact with Tivozanib Hydrochloride — tap one for details:
Where does this data come from?
- MedlinePlus (NLM) · refreshed Oct 1, 2026
- FDA label on DailyMed · label index refreshed Oct 1, 2026
Ask a licensed pharmacist directly — free, answered by our team.
Pricing
A drug doesn't have one price. Each row is a different public payment system, and none is what you'd pay at the counter — that depends on your insurance. The ⓘ on each row explains what it measures.
| Price system | Per each | Per package |
|---|---|---|
| Retail pharmacies payNADAC · weekly | Not in the retail survey — common for institutional, discontinued, or low-volume packs. | |
| Medicaid paysCMS SDUD · 12 mo | $1,514.54 | $31,805.39 / 21 capsules |
| Medicare drug plans payPart D · Q2 2026 | $1,506.30 | $31,632.39 / 21 capsules |
Where does this data come from?
- CMS NADAC weekly file
- CMS ASP pricing files · refreshed Sep 20, 2026
- CMS Medicaid State Drug Utilization Data · through Q1 2026
- CMS Part D plan pricing files · refreshed Sep 24, 2026
- VA National Acquisition Center price file
Packaging — all sizes for this product
| Package NDC | Description | Marketing start | Marketing end | Status |
|---|---|---|---|---|
| 45629-0134-01 45629-134-01 Main listing | 24 BOTTLE, PLASTIC in 1 CARTON / 21 CAPSULE in 1 BOTTLE, PLASTIC | 2021-03-10 | — | Discontinued by firm |
| 45629-0134-02 You're viewing this | 1 BOTTLE, PLASTIC in 1 CARTON / 21 CAPSULE in 1 BOTTLE, PLASTIC | 2025-01-30 | — | Active |
Pack size FAQ
What quantity is in this package?
What NDC number is used to bill for this package of FOTIVDA Tivozanib 1.34 mg Capsule?
Therapeutic equivalents
| Product | Labeler | Pack | NADAC/unit | TE | Status | Price vs. this |
|---|---|---|---|---|---|---|
| Fotivda 1.34 mgthis 45629-0134-02 | AVEO | 21 capsules | — | — | FDA listed | — |
Where does this data come from?
- FDA openFDA NDC Directory · synced Oct 1, 2026
- FDA Orange Book · refreshed Oct 3, 2026
- CMS NADAC weekly file
Availability & generic status
We did not find an FDA-approved generic match for this exact strength, form and route. Patent/protection dates below may affect future generic timing.
Why the date isn’t exact: Generic timing can change because patents may be challenged, settled, licensed, added, removed, or worked around with a narrower label — and FDA approval does not always mean a pharmacy can get the generic today.
🛈 What do these terms mean?
- Patent
- Legal protection listed in the Orange Book that may delay generic approval or launch. Issued by the U.S. Patent & Trademark Office.
- Substance patent
- Covers the active drug molecule itself — the hardest to design around. A generic generally can’t launch until it expires.
- Formulation (product) patent
- Covers a specific formulation or dosage form. A generic can sometimes work around it with a different formulation.
- Method-of-use patent
- A patent covering one specific approved use of the drug — not necessarily the whole molecule. A generic can sometimes launch with a “skinny label” that carves out the protected use and keeps the others.
- Skinny label
- A generic label that omits a still-patented use when the FDA allows it — letting a generic reach the market for the unprotected uses.
- Exclusivity
- FDA-granted marketing protection, separate from patents — e.g. 5-yr new chemical entity, 7-yr orphan drug, or a +6-month pediatric extension.
- Paragraph IV
- A generic applicant’s formal challenge to a listed patent. It can potentially lead to earlier generic entry, but often involves litigation or a settlement.
- RLD / RS
- Reference Listed Drug — the brand product the FDA uses as the reference for generic applications. Reference Standard — the product the FDA expects generics to compare against in bioequivalence testing.
- TE / AB rating
- FDA therapeutic-equivalence rating. An AB rating generally means the FDA considers a generic therapeutically equivalent to — and substitutable for — the brand.
- LOE (loss of exclusivity)
- The latest patent or exclusivity currently listed — the loss-of-exclusivity / latest-listed-protection date shown on this page. Paragraph-IV challenges and settlements can move the real date earlier; FDA approval and a manufacturer’s decision to market can move it later.
Built from the FDA Orange Book. The bars above are scaled to each protection’s expiry; the red LOE marker is the last one to lapse.
| Patent | Type | Use code | Expires |
|---|---|---|---|
| US 11504365 ↗ | Method of use | U-3476 | Nov 5, 2039 |
| US 11504365 ↗ | Method of use | U-3476 | Nov 5, 2039 |
| US 7166722 ↗ | Drug substance | — | Nov 16, 2028 |
| US 7166722 ↗ | Drug substance | — | Nov 16, 2028 |
| Code | What it grants | Expires |
|---|---|---|
| NCE | New Chemical Entity (5-year) | Mar 10, 2026 |
| NCE | New Chemical Entity (5-year) | Mar 10, 2026 |
Is there a generic version of FOTIVDA 1.34 MG CAPSULE?
The FDA approved a generic — why can’t I get it at my pharmacy yet?
Why do different websites show different generic release dates?
What does “FDA listed” mean?
What does a patent or protection date mean here?
What does “current Orange Book estimate” mean?
Can a generic come out before the last patent expires?
Can a generic come out after the listed dates?
What is the difference between patents and exclusivity?
Why are there multiple patent dates?
Where does this data come from?
- FDA Orange Book · refreshed Oct 3, 2026
What it looks like
Where does this data come from?
- FDA label on DailyMed · label index refreshed Oct 1, 2026
Inactive Ingredients / Excipients
Inactive ingredients, also called excipients, are components of the drug product other than the active ingredient. They may include fillers, dyes, coatings, preservatives, flavors, or other formulation ingredients.
💡 Tap an ingredient (hover on desktop) to see what it is and why it’s used.
-
UNII 70097M6I30
Magnesium stearate is a salt made from magnesium and stearic acid, a fatty substance. It's used in tablets and capsules as a lubricant and glidant to help ingredients flow smoothly during manufacturing and prevent sticking.
-
UNII 3OWL53L36A
A natural sugar alcohol derived from seaweed or synthesized in the lab. It's used as a filler to add bulk, a sweetener in sugar-free formulas, and a disintegrant to help tablets break apart in the stomach.
2 inactive ingredients listed in the exact product block matched to this NDC.
Where does this data come from?
ingredient classCode="IACT" elements from the exact product block matched by this NDC. Label-section narrative from DailyMed / the openFDA label index is shown separately when available.- FDA label on DailyMed · label index refreshed Oct 1, 2026
- FDA openFDA NDC Directory · synced Oct 1, 2026
Inactive ingredient FAQ
Are inactive ingredients the same for every manufacturer?
Why might an inactive ingredient be missing?
Can inactive ingredients matter?
Manufacturer & labeler
Where does this data come from?
- FDA openFDA NDC Directory · synced Oct 1, 2026
- Drugs@FDA
Full prescribing information FDA SPL
🎯 Indications and Usage ▾
1 INDICATIONS AND USAGE FOTIVDA is indicated for the treatment of adult patients with relapsed or refractory advanced renal cell carcinoma (RCC) following two or more prior systemic therapies. FOTIVDA is a kinase inhibitor indicated for the treatment of adult patients with relapsed or refractory advanced renal cell carcinoma (RCC) following two or more prior systemic therapies. ( 1 )
⏱️ Dosage and Administration ▾
2 DOSAGE AND ADMINISTRATION Recommended Dose: 1.34 mg once daily with or without food for 21 days on treatment followed by 7 days off treatment (28-day cycle) until disease progression or unacceptable toxicity. ( 2.1 ) Dose interruptions and/or dose reduction may be needed to manage adverse reactions. ( 2.2 ) For patients with moderate hepatic impairment, reduce the dose to 0.89 mg for 21 days on treatment followed by 7 days off treatment (28-day cycle).
( 2.3 )
2.1Recommended Dosing The recommended dosage of FOTIVDA is 1.34 mg taken orally once daily for 21 days on treatment followed by 7 days off treatment for a 28-day cycle. Continue treatment until disease progression or until unacceptable toxicity occurs. Take FOTIVDA with or without food.
Swallow the FOTIVDA capsule whole with a glass of water. Do not open the capsule. If a dose is missed, the next dose should be taken at the next scheduled time.
Do not take two doses at the same time.
2.2Dose Modifications for Adverse Reactions Initiate medical management for diarrhea, nausea, or vomiting prior to dose interruption or reduction. If dose modifications are required for adverse reactions, reduce the dosage of FOTIVDA to 0.89 mg for 21 days on treatment followed by 7 days off treatment for a 28-day cycle. Recommendations for dosage modifications are provided in Table 1 .
Table 1. Dosage Modifications for Adverse Reactions Adverse Reaction Severity Grades are based on the National Cancer Institute Common Terminology Criteria for Adverse Events (CTCAE). Dosage Modifications for FOTIVDA Hypertension [see Warnings and Precautions (5.1) ] Grade 3 Withhold for Grade 3 that persists despite optimal anti-hypertensive therapy.
Resume at reduced dose when hypertension is controlled at less than or equal to Grade 2. Grade 4 Permanently discontinue. Cardiac Failure [see Warnings and Precautions (5.2) ] Grade 3 Withhold until improves to Grade 0 to 1 or baseline.
Resume at a reduced dose or discontinue depending on the severity and persistence of adverse reaction. Grade 4 Permanently discontinue. Arterial Thromboembolic Events [see Warnings and Precautions (5.3) ] Any Grade Permanently discontinue.
Hemorrhagic Events [see Warnings and Precautions (5.5) ] Grade 3 or 4 Permanently discontinue. Proteinuria [see Warnings and Precautions (5.6) ] 2 grams or greater proteinuria in 24 hours Withhold until less than or equal to 2 grams of proteinuria per 24 hours. Resume at a reduced dose.
Permanently discontinue for nephrotic syndrome. Reversible Posterior Leukoencephalopathy Syndrome [see Warnings and Precautions (5.10) ] Any Grade Permanently discontinue. Other Adverse Reactions Persistent or intolerable Grade 2 or 3 adverse reaction Grade 4 laboratory abnormality Withhold until improves to Grade 0 to 1 or baseline.
Resume at reduced dose. Grade 4 adverse reaction Permanently discontinue.
2.3Dosage Modifications for Moderate Hepatic Impairment Reduce the recommended dosage of FOTIVDA to 0.89 mg capsule taken orally once daily for 21 days on treatment followed by 7 days off treatment for a 28-day cycle for patients with moderate hepatic impairment [see USE IN SPECIFIC POPULATIONS (8.7) ].
💊 Dosage Forms and Strengths ▾
3 DOSAGE FORMS AND STRENGTHS Capsules: 1.34 mg: bright yellow opaque cap imprinted with "TIVZ" in dark blue ink and a bright yellow opaque body imprinted with "SD" in dark blue ink. 0.89 mg: dark blue opaque cap imprinted with "TIVZ" in yellow ink and a bright yellow opaque body imprinted with "LD" in dark blue ink. Capsules: 1.34 mg and 0.89 mg ( 3 )
⛔ Contraindications ▾
4 CONTRAINDICATIONS None. None. ( 4 )
⚠️ Warnings and Cautions ▾
5 WARNINGS AND PRECAUTIONS Hypertension and Hypertensive Crisis: Control blood pressure prior to initiating FOTIVDA. Monitor for hypertension and treat as needed. For persistent hypertension despite use of anti-hypertensive medications, reduce the FOTIVDA dose.
( 5.1 ) Cardiac Failure: Monitor for signs or symptoms of cardiac failure throughout treatment with FOTIVDA. ( 5.2 ) Cardiac Ischemia and Arterial Thromboembolic Events: Closely monitor patients who are at increased risk for these events. Permanently discontinue FOTIVDA for severe arterial thromboembolic events, such as myocardial infarction and stroke.
( 5.3 ) Venous Thromboembolic Events: Closely monitor patients who are at increased risk for these events. Permanently discontinue FOTIVDA for severe venous thromboembolic events. ( 5.4 ) Hemorrhagic Events: Closely monitor patients who are at risk for or who have a history of bleeding.
( 5.5 ) Proteinuria: Monitor throughout treatment with FOTIVDA. For moderate to severe proteinuria, reduce the dose or temporarily interrupt treatment with FOTIVDA. ( 5.6 ) Perforations and Fistulas: Monitor for symptoms.
Discontinue FOTIVDA for severe or life-threatening gastrointestinal perforation. ( 5.7 ) Thyroid Dysfunction: Monitor before initiation and throughout treatment with FOTIVDA. ( 5.8 ) Risk of Impaired Wound Healing: Withhold FOTIVDA for at least 24 days before elective surgery.
Do not administer for at least 2 weeks following major surgery and adequate wound healing. The safety of resumption of FOTIVDA after resolution of wound healing complications has not been established. ( 5.9 ) Reversible Posterior Leukoencephalopathy Syndrome (RPLS): Discontinue FOTIVDA if signs or symptoms of RPLS occur.
( 5.10 ) Embryo-Fetal Toxicity: Can cause fetal harm. Advise patients of the potential risk to a fetus and to use effective contraception. ( 5.11 , 8.1 , 8.3 ) Allergic Reactions to Tartrazine: The 0.89 mg capsule of FOTIVDA contains FD&C Yellow No.5 (tartrazine) which may cause allergic-type reactions (including bronchial asthma) in certain susceptible patients.
( 5.12 )
5.1Hypertension and Hypertensive Crisis FOTIVDA can cause severe hypertension and hypertensive crisis. Hypertension occurred in 45% of patients treated with FOTIVDA, with 22% of the events ≥ Grade 3. Median time to onset of hypertension was 2 weeks (range: 0 – 192 weeks).
Hypertensive crisis occurred in 0.8% of patients . One patient (0.1%) died due to hypertensive emergency after FOTIVDA overdose [see OVERDOSAGE (10) ] . FOTIVDA has not been studied in patients with systolic blood pressure > 150 mmHg or diastolic blood pressure > 100 mmHg.
Control blood pressure prior to treatment with FOTIVDA. Monitor blood pressure after 2 weeks and at least monthly thereafter during treatment with FOTIVDA. Treat patients with anti- hypertensive therapy when hypertension occurs during treatment with FOTIVDA.
Withhold FOTIVDA for severe hypertension despite optimal anti-hypertensive therapy. For persistent hypertension despite use of anti-hypertensive medications, reduce the FOTIVDA dose [see DOSAGE AND ADMINISTRATION (2.2) ] . Discontinue FOTIVDA if hypertension is severe and persistent despite anti-hypertensive therapy and dose reduction of FOTIVDA, or in patients who experience hypertensive crisis.
If FOTIVDA is interrupted, monitor patients receiving anti-hypertensive medications for hypotension.
5.2Cardiac Failure FOTIVDA can cause serious, sometimes fatal, cardiac failure. Cardiac failure in FOTIVDA- treated patients occurred in 1.6%, with 1% of events ≥ Grade 3, and 0.6% events were fatal. FOTIVDA has not been studied in patients with symptomatic cardiac failure within the preceding 6 months before FOTIVDA treatment initiation.
Periodically monitor patients for symptoms of cardiac failure throughout treatment with FOTIVDA. Management of cardiac failure events may require interruption, dose reduction, or permanent discontinuation of FOTIVDA therapy [see DOSAGE AND ADMINI… [Excerpted — this section continues on DailyMed.]
🤒 Adverse Reactions ▾
6 ADVERSE REACTIONS The following clinically significant adverse reactions are also described elsewhere in the labeling: Hypertension and Hypertensive Crisis [see WARNINGS AND PRECAUTIONS (5.1) ] Cardiac Failure [see WARNINGS AND PRECAUTIONS (5.2) ] Cardiac Ischemia and Arterial Thromboembolic Events [see WARNINGS AND PRECAUTIONS (5.3) ] Venous Thromboembolic Events [see WARNINGS AND PRECAUTIONS (5.4) ] Hemorrhagic Events [see WARNINGS AND PRECAUTIONS (5.5) ] Proteinuria [see WARNINGS AND PRECAUTIONS (5.6) ] Gastrointestinal Perforation and Fistula Formation [see WARNINGS AND PRECAUTIONS (5.7) ] Thyroid Dysfunction [see WARNINGS AND PRECAUTIONS (5.8) ] Risk of Impaired Wound Healing [see WARNINGS AND PRECAUTIONS (5.9) ] Reversible Posterior Leukoencephalopathy Syndrome (RPLS) [see WARNINGS AND PRECAUTIONS (5.10) ] The most common (≥20%) adverse reactions were fatigue, hypertension, diarrhea, decreased appetite, nausea, dysphonia, hypothyroidism, cough, and stomatitis, and the most common Grade 3 or 4 laboratory abnormalities (≥5%) were sodium decreased, lipase increased, and phosphate decreased.
( 6.1 ) To report SUSPECTED ADVERSE REACTIONS, contact AVEO Pharmaceuticals, Inc. at 1-833-FOTIVDA (1-833-368-4832) or FDA at 1-800-FDA-1088 or www.fda.gov/medwatch .
6.1Clinical Trial Experience Because clinical studies are conducted under widely varying conditions, adverse reaction rates observed in clinical studies of a drug cannot be directly compared to rates in the clinical studies of another drug and may not reflect the rates observed in practice. The pooled safety population described in WARNINGS AND PRECAUTIONS reflects exposure to FOTIVDA administered at 1.34 mg orally once daily with or without food for 21 days on treatment followed by 7 days off treatment for a 28-day cycle in 1008 patients with advanced RCC in TIVO-3 and five other monotherapy studies.
Among 1008 patients who received FOTIVDA, 52% were exposed for 6 months or longer and 34% were exposed for greater than one year. Relapsed or Refractory Advanced RCC Following Two or More Prior Systemic Therapies The safety of FOTIVDA was evaluated in TIVO-3, a randomized, open-label trial in 350 patients with relapsed or refractory advanced RCC who received 2 or 3 prior systemic treatments [see CLINICAL STUDIES (14) ] . Patients were randomized (1:1) to receive FOTIVDA 1.34 mg orally once daily for 21 days on treatment followed by 7 days off treatment for a 28-day cycle, or to receive sorafenib 400 mg orally twice a day continuously until disease progression or unacceptable toxicity.
Among patients who received FOTIVDA, 53% were exposed for 6 months or longer and 31% were exposed for greater than one year. Serious adverse reactions occurred in 45% of patients who received FOTIVDA. Serious adverse reactions in > 2% of patients included bleeding (3.5%), venous thromboembolism (3.5%), arterial thromboembolism (2.9%), acute kidney injury (2.3%), and hepatobiliary disorders (2.3%).
Fatal adverse reactions occurred in 8% of patients who received FOTIVDA, including pneumonia (1.7%), hepatobiliary disorders (1.2%), respiratory failure (1.2%), myocardial infarction (0.6%), cerebrovascular accident (0.6%), and subdural hematoma (0.6%). Permanent discontinuation of FOTIVDA due to an adverse reaction occurred in 21% of patients. Adverse reactions which resulted in permanent discontinuation of FOTIVDA in > 2 patients included hepatobiliary disorders, fatigue, and pneumonia.
Dosage interruptions of FOTIVDA due to an adverse reaction occurred in 48% of patients. Adverse reactions which required dosage interruption in > 5% of patients included fatigue, hypertension, decreased appetite, and nausea. Dose reductions of FOTIVDA due to an adverse reaction occurred in 24% of patients.
Adverse reactions which required dose reductions in > 3% of patients included fatigue, diarrhea, and decreased appetite. The most common (≥ 20%) adverse reactions were fatigue, hypertension, diarrhea, decreased ap… [Excerpted — this section continues on DailyMed.]
🔄 Drug Interactions ▾
7 DRUG INTERACTIONS CYP3A Inducers: Avoid concomitant use of strong CYP3A inducers. ( 7.1 )
7.1Effect of Other Drugs on FOTIVDA Strong CYP3A Inducers Concomitant use of FOTIVDA with a strong CYP3A inducer decreases tivozanib exposure [see CLINICAL PHARMACOLOGY (12.3) ], which may reduce FOTIVDA anti-tumor activity. Avoid concomitant use of strong CYP3A inducers with FOTIVDA.
👥 Use in Specific Populations ▾
8 USE IN SPECIFIC POPULATIONS Lactation: Advise not to breastfeed. ( 8.2 ) Females and Males of Reproductive Potential: Can impair fertility. ( 8.3 ) Hepatic Impairment: Adjust dosage in patients with moderate hepatic impairment. Avoid use in patients with severe hepatic impairment. ( 2.3 , 8.7 )
8.1Pregnancy Risk Summary Based on findings in animal studies and its mechanism of action, FOTIVDA can cause fetal harm when administered to a pregnant woman [see CLINICAL PHARMACOLOGY (12.1) ] . There are no available data on FOTIVDA use in pregnant woman to inform the drug-associated risk. In embryo-fetal developmental studies, oral administration of tivozanib to pregnant animals during the period of organogenesis caused maternal toxicity, fetal malformations and embryo- fetal death at doses below the maximum recommended clinical dose on a mg/m 2 basis [see DATA ] .
Advise pregnant woman of the potential risk to a fetus. The estimated background risk of major birth defects and miscarriage for the indicated population is unknown. All pregnancies have a background risk of birth defect, loss, or other adverse outcomes.
In the U.S. general population, the estimated background risk of major birth defects and miscarriage in clinically-recognized pregnancies is 2% to 4% and 15% to 20% respectively. Data Animal Data In an embryo-fetal developmental study in pregnant rats, daily oral administration of tivozanib at doses ≥ 0.03 mg/kg/day (0.2 times the maximum recommended clinical dose on a mg/m 2 basis) during the period of organogenesis resulted in maternal toxicity, increases in early and late resorptions, and an increase in fetal external malformations (body edema, short/kinked tail), and skeletal developmental delays.
In an embryo-fetal developmental study in pregnant rabbits, daily oral administration of tivozanib at 1 mg/kg/day (14.5 times the maximum recommended clinical dose on a mg/m 2 basis) during the period of organogenesis resulted in fetal malformations including ventricular septal defects and major vessel anomalies. No maternal toxicity was reported at doses up to 1 mg/kg/day.
8.2Lactation Risk Summary There are no data on the presence of tivozanib in human milk, or the effects of tivozanib on the breastfed child, or on milk production. Because of the potential for serious adverse reactions in a breastfed child, advise a lactating woman not to breastfeed during treatment with FOTIVDA and for one month after the last dose.
8.3Females and Males of Reproductive Potential FOTIVDA can cause fetal harm when administered to a pregnant woman [see USE IN SPECIFIC POPULATIONS (8.1) ] . Pregnancy Testing Verify pregnancy status of females of reproductive potential prior to starting treatment with FOTIVDA. Contraception Females Advise females of reproductive potential to use effective contraception during treatment with FOTIVDA and for one month after the last dose [see USE IN SPECIFIC POPULATIONS (8.1) ] .
Males Advise males with female partners of reproductive potential to use effective contraception during treatment with FOTIVDA and for one month after the last dose [see NONCLINICAL TOXICOLOGY (13.1) ] . Infertility Females and Males Based on findings in animal studies, FOTIVDA can impair fertility in females and males of reproductive potential [see NONCLINICAL TOXICOLOGY (13.1) ] .
8.4Pediatric Use The safety and effectiveness of FOTIVDA in pediatric patients have not been established. Animal Data Juvenile animal studies have not been conducted with tivozanib. In a 13-week repeat-dose study, oral administration of tivozanib to young and growing cynomolgus monkeys resulted in growth plate hypertrophy, absence of active corpora lutea, and no maturing follicles at doses ≥ 0.3 mg/kg/day (4.4 times the maximum recommended clinical dose on a mg/m 2 basis).
In a 13-week repeat-dose study in rats, teeth abnormalities (thin, brittle teeth, tooth loss, malocclusions) and growth plate hypertrophy were observed following oral administration o… [Excerpted — this section continues on DailyMed.]
🤰 Pregnancy ▾
8.1Pregnancy Risk Summary Based on findings in animal studies and its mechanism of action, FOTIVDA can cause fetal harm when administered to a pregnant woman [see CLINICAL PHARMACOLOGY (12.1) ] . There are no available data on FOTIVDA use in pregnant woman to inform the drug-associated risk. In embryo-fetal developmental studies, oral administration of tivozanib to pregnant animals during the period of organogenesis caused maternal toxicity, fetal malformations and embryo- fetal death at doses below the maximum recommended clinical dose on a mg/m 2 basis [see DATA ] .
Advise pregnant woman of the potential risk to a fetus. The estimated background risk of major birth defects and miscarriage for the indicated population is unknown. All pregnancies have a background risk of birth defect, loss, or other adverse outcomes.
In the U.S. general population, the estimated background risk of major birth defects and miscarriage in clinically-recognized pregnancies is 2% to 4% and 15% to 20% respectively. Data Animal Data In an embryo-fetal developmental study in pregnant rats, daily oral administration of tivozanib at doses ≥ 0.03 mg/kg/day (0.2 times the maximum recommended clinical dose on a mg/m 2 basis) during the period of organogenesis resulted in maternal toxicity, increases in early and late resorptions, and an increase in fetal external malformations (body edema, short/kinked tail), and skeletal developmental delays.
In an embryo-fetal developmental study in pregnant rabbits, daily oral administration of tivozanib at 1 mg/kg/day (14.5 times the maximum recommended clinical dose on a mg/m 2 basis) during the period of organogenesis resulted in fetal malformations including ventricular septal defects and major vessel anomalies. No maternal toxicity was reported at doses up to 1 mg/kg/day.
🧒 Pediatric Use ▾
8.4Pediatric Use The safety and effectiveness of FOTIVDA in pediatric patients have not been established. Animal Data Juvenile animal studies have not been conducted with tivozanib. In a 13-week repeat-dose study, oral administration of tivozanib to young and growing cynomolgus monkeys resulted in growth plate hypertrophy, absence of active corpora lutea, and no maturing follicles at doses ≥ 0.3 mg/kg/day (4.4 times the maximum recommended clinical dose on a mg/m 2 basis).
In a 13-week repeat-dose study in rats, teeth abnormalities (thin, brittle teeth, tooth loss, malocclusions) and growth plate hypertrophy were observed following oral administration of tivozanib at doses ≥ 0.1 mg/kg/day (0.7 times the maximum recommended clinical dose on a mg/m 2 basis).
🧓 Geriatric Use ▾
8.5Geriatric Use Of the 1008 patients with advanced RCC treated with FOTIVDA, 29% were ≥ 65 years of age and 4% were ≥ 75 of age. No overall differences in safety were observed between patients ≥ 65 versus < 65 years of age. Of the 175 patients with advanced RCC following two or more prior systemic therapies randomized to FOTIVDA, 44% were ≥ 65 years of age and 9% were ≥ 75 of age.
No overall differences in effectiveness were observed between patients ≥ 65 versus < 65 years of age.
🆘 Overdosage ▾
10 OVERDOSAGE Overdosage with FOTIVDA can cause severe hypertension and hypertensive crisis that may result in death [see WARNINGS AND PRECAUTIONS (5.1) ] . During clinical studies, three patients inadvertently received doses ≥ 2.68 mg (≥ 2 times the recommended dose) of FOTIVDA. One patient who received two daily doses of 8.9 mg of FOTIVDA experienced hypertensive crisis with severe hypertensive retinopathy; a second patient who received three doses of 1.34 mg in one day experienced fatal uncontrolled hypertension; and a third patient who received two doses of 1.34 mg FOTIVDA in one day experienced persistent hypertension lasting over 5 days.
There is no specific treatment or antidote for FOTIVDA overdose. In cases of suspected overdose, withhold FOTIVDA, closely monitor patients for hypertension and hypertensive crisis and other potential adverse reactions. Immediately manage signs or symptoms of hypertension and provide other supportive care as clinically indicated.
🧬 Clinical Pharmacology ▾
12 CLINICAL PHARMACOLOGY
12.1Mechanism of Action Tivozanib is a tyrosine kinase inhibitor. In vitro cellular kinase assays demonstrated that tivozanib inhibits phosphorylation of vascular endothelial growth factor receptor (VEGFR)-1, VEGFR-2 and VEGFR-3 and inhibits other kinases including c-kit and PDGFR β at clinically relevant concentrations. In tumor xenograft models in mice and rats, tivozanib inhibited angiogenesis, vascular permeability, and tumor growth of various tumor cell types including human renal cell carcinoma.
12.2Pharmacodynamics Exposure-Response Relationship Tivozanib exposure-response relationships and the time course of pharmacodynamic response have not been fully characterized. Cardiac Electrophysiology At the recommended dose of FOTIVDA, no large mean increases (i.e., 20 msec) in QTc interval were observed.
12.3Pharmacokinetics The pharmacokinetics of tivozanib were evaluated in patients with solid tumors administered 1.34 mg once daily unless otherwise specified. Steady-state tivozanib AUC and C max increased in a dose-proportional manner over the dose range of 0.89 to 1.78 mg once daily (0.67 to 1.3 times the recommended dose). Steady-state was reached by 14 days and the accumulation ratio after administration of 1.34 mg once daily was approximately 6- to 7- fold.
Mean steady-state tivozanib [coefficient of variation (CV%)] C max was 86.9 (44.7%) ng/mL and AUC 0-24h was 1510 (46.1%) ng*h/mL. Absorption The median T max of tivozanib is 10 hours with a range of 3 to 24 hours. Effect of Food No clinically significant differences in tivozanib AUC or C max were observed following administration of a high fat meal (approximately 500-600 fat calories, 250 carbohydrate calories and 150 protein calories) in healthy subjects.
Distribution The apparent volume of distribution (V/F) of tivozanib is 123 L. Protein binding of tivozanib is ≥ 99%, primarily to albumin in vitro and is independent of concentration. The mean blood-to-plasma concentration ratios ranged from 0.495 to 0.615 in healthy subjects.
Elimination The apparent clearance (CL/F) of tivozanib is
0.75L/h and the half-life is 111 hours. Metabolism Tivozanib is metabolized predominantly by CYP3A4. Following oral administration of a single radiolabeled 1.34 mg dose of tivozanib to healthy subjects, unchanged tivozanib constituted 90% of the radioactive drug components in serum.
Excretion Following oral administration of a single radiolabeled 1.34 mg dose of tivozanib to healthy subjects, 79% of the administered dose was recovered in feces (26% unchanged) and 12% in urine (unchanged tivozanib not detected). Specific Populations No clinically significant differences in the pharmacokinetics of tivozanib were observed based on age (18 years to 88 years), sex, race (93% Caucasian, 3% African American, 2% Asian, 2% others), body weight (39 kg to 158 kg), mild to severe renal impairment (CLcr 15-89 mL/min as estimated by Cockcroft-Gault) or mild hepatic impairment (total bilirubin less than or equal to ULN with AST greater than ULN or total bilirubin greater than 1 to 1.5 times ULN with any AST).
The effect of end-stage renal disease or severe hepatic impairment on tivozanib pharmacokinetics is unknown [see USE IN SPECIFIC POPULATIONS (8.6) and (8.7) ] . Patients with Hepatic Impairment Compared to subjects with normal hepatic function, tivozanib AUC tau increased by 1% in patients with mild (total bilirubin less than or equal to ULN with AST greater than ULN or total bilirubin greater than 1 to 1.5 times ULN with any AST) hepatic impairment. Compared to subjects with normal hepatic function, tivozanib AUC tau increased by 62% in patients with moderate (total bilirubin greater than 1.5 to 3 times ULN with any AST) hepatic impairment.
The effect of severe (total bilirubin greater than 3 to 10 times ULN with any AST) hepatic impairment on tivozanib pharmacokinetics has not been studied [see DOSAGE AND ADMINISTRATION (2.3) and USE IN SPECIFIC POPLATIONS (8.7) ].… [Excerpted — this section continues on DailyMed.]
🧬 Mechanism of Action ▾
12.1Mechanism of Action Tivozanib is a tyrosine kinase inhibitor. In vitro cellular kinase assays demonstrated that tivozanib inhibits phosphorylation of vascular endothelial growth factor receptor (VEGFR)-1, VEGFR-2 and VEGFR-3 and inhibits other kinases including c-kit and PDGFR β at clinically relevant concentrations. In tumor xenograft models in mice and rats, tivozanib inhibited angiogenesis, vascular permeability, and tumor growth of various tumor cell types including human renal cell carcinoma.
📦 How Supplied / Storage and Handling ▾
16 HOW SUPPLIED/STORAGE AND HANDLING How Supplied FOTIVDA (tivozanib) capsules, for oral use are supplied as follows: Capsule Strength Opaque Capsule Color Capsule Markings Pack Size NDC Code Tivozanib 1.34 mg (equivalent to 1.5 mg tivozanib hydrochloride) Bright yellow cap and body "TIVZ" imprinted with dark blue ink on cap; "SD" imprinted with dark blue ink on body Bottle of 21 NDC 45629-134-01 Tivozanib 0.89 mg (equivalent to 1.0 mg tivozanib hydrochloride) Dark blue cap and bright yellow body "TIVZ" imprinted with yellow ink on cap; "LD" imprinted with dark blue ink on body Bottle of 21 NDC 45629-089-01 Tivozanib 1.34 mg (equivalent to 1.5 mg tivozanib hydrochloride) Bright yellow cap and body "TIVZ" imprinted with dark blue ink on cap; "SD" imprinted with dark blue ink on body Bottle of 21 in a Carton NDC 45629-134-02 Tivozanib 0.89 mg (equivalent to 1.0 mg tivozanib hydrochloride) Dark blue cap and bright yellow body "TIVZ" imprinted with yellow ink on cap; "LD" imprinted with dark blue ink on body Bottle of 21 in a Carton NDC 45629-089-02 Storage and Handling Store at 20 ° C to 25 ° C (68 ° F to 77 ° F); excursions permitted between 15°C to 30 ° C (59°F to 86 ° F) [see USP CONTROLLED ROOM TEMPERATURE] .
Keep out of reach of children.
📦 Storage and Handling ▾
Storage and Handling Store at 20 ° C to 25 ° C (68 ° F to 77 ° F); excursions permitted between 15°C to 30 ° C (59°F to 86 ° F) [see USP CONTROLLED ROOM TEMPERATURE] . Keep out of reach of children.
📋 Description ▾
11 DESCRIPTION Tivozanib is a kinase inhibitor. Tivozanib hydrochloride, the active ingredient, has the chemical name 1-{2-chloro-4-[(6,7-dimethoxyquinolin-4-yl)oxy]phenyl}-3-(5-methylisoxazol-3-yl)urea hydrochloride hydrate. The molecular formula is C 22 H 19 ClN 4 O 5 ∙ HCl ∙ H 2 O and the molecular weight is 509.34 Daltons.
The chemical structure is: Tivozanib hydrochloride is a white to light brown crystalline powder that is practically insoluble in water (0.09 mg/mL). FOTIVDA 1.34 mg capsule contains 1.5 mg of tivozanib hydrochloride (equivalent to 1.34 mg tivozanib) with inactive ingredients: mannitol and magnesium stearate. Capsule composition: gelatin, titanium dioxide, FDA yellow iron oxide, and Blue SB-6018 (ink).
FOTIVDA 0.89 mg capsule contains 1.0 mg of tivozanib hydrochloride (equivalent to 0.89 mg tivozanib) with inactive ingredients: mannitol and magnesium stearate. Capsule composition: gelatin, titanium dioxide, FDA yellow iron oxide, FD&C Blue #2, Blue SB-6018 (ink) and Yellow SB-3017 (ink). The Yellow SB-3017 ink contains FD&C Yellow No.5 (tartrazine).
Chemical Structure
💬 Information for Patients ▾
17 PATIENT COUNSELING INFORMATION Advise the patient to read the FDA-approved patient labeling [see PATIENT INFORMATION ] . Hypertension and Hypertensive Crisis Inform patients that hypertension or hypertensive crisis may occur during FOTIVDA treatment. Advise patients to undergo routine blood pressure monitoring and to contact their healthcare provider if blood pressure is elevated.
Advise patients that if they experience signs or symptoms of hypertension to immediately contact their healthcare provider [see WARNINGS AND PRECAUTIONS (5.1) ]. Cardiac Failure Advise patients to immediately contact their healthcare provider if they develop symptoms of cardiac failure [see WARNINGS AND PRECAUTIONS (5.2) ]. Cardiac Ischemia and Arterial Thromboembolic Events Inform patients that arterial thromboembolic events (including fatal outcomes) may occur during FOTIVDA treatment.
Advise patients to immediately contact their healthcare provider if new onset of chest discomfort, sudden weakness, or other events suggestive of a thrombotic event occurs [see WARNINGS AND PRECAUTIONS (5.3) ]. Venous Thromboembolic Events Advise patients to immediately contact their healthcare provider if they develop symptoms of dyspnea or localized limb edema [see WARNINGS AND PRECAUTIONS (5.4) ]. Hemorrhagic Events Instruct patients to contact their healthcare provider to seek immediate medical attention for signs or symptoms of unusual bleeding or hemorrhage [see WARNINGS AND PRECAUTIONS (5.5) ] .
Perforations and Fistulas Inform patients that gastrointestinal perforations or fistulas may develop during FOTIVDA treatment. Advise patients to immediately contact their healthcare provider if they experience persistent or severe abdominal pain [see WARNINGS AND PRECAUTIONS (5.7) ] . Risk of Impaired Wound Healing Inform patients that FOTIVDA may impair wound healing.
Advise patients that temporary interruption of FOTIVDA is recommended prior to elective surgery. Advise patients to contact their healthcare provider before any planned surgeries, including dental surgery [see DOSAGE AND ADMINISTRATION (2.1) and WARNINGS AND PRECAUTIONS (5.9) ] . Reversible Posterior Leukoencephalopathy Syndrome Inform patients that RPLS may occur during FOTIVDA treatment.
Advise patients to immediately contact their healthcare provider in the event of seizures, headaches, visual disturbances, confusion or difficulty thinking [see WARNINGS AND PRECAUTIONS (5.10) ]. Overdosage Instruct patients to contact their healthcare provider immediately if they inadvertently take too much FOTIVDA [see OVERDOSAGE (10) ] . Embryo-Fetal Toxicity Advise females of reproductive potential of the potential risk to a fetus.
Advise patients to inform their healthcare provider of a known or suspected pregnancy [see WARNINGS AND PRECAUTIONS (5.10) and USE IN SPECIFIC POPULATIONS (8.1) ]. Advise females of reproductive potential to use effective contraception during treatment with FOTIVDA and for one month after the last dose. Advise male patients with female partners of reproductive potential to use effective contraception during treatment with FOTIVDA and for one month after the last dose [see USE IN SPECIFIC POPULATIONS (8.1) and (8.3) and NONCLINICAL TOXICOLOGY (13.1) ].
Lactation Advise women not to breastfeed during treatment with FOTIVDA and for one month after the last dose [see USE IN SPECIFIC POPULATIONS (8.2) ]. Infertility Advise males and females of reproductive potential that FOTIVDA can impair fertility [see USE IN SPECIFIC POPULATIONS (8.3) ] . Allergic Reactions to Tartrazine (FD&C Yellow No.5) FOTIVDA 0.89 mg capsule contains FD&C Yellow No.5 (tartrazine) as an imprint ink which may cause allergic-type reactions (including bronchial asthma) in certain susceptible patients.
Although the overall incidence of FD&C Yellow No.5 (tartrazine) sensitivity in the general population is low, it is frequently seen in patients who also have aspirin hypersensitivity [see WARNINGS AND PRECAUTIONS (5.12)… [Excerpted — this section continues on DailyMed.]
🧬 Pharmacokinetics ▾
12.3Pharmacokinetics The pharmacokinetics of tivozanib were evaluated in patients with solid tumors administered 1.34 mg once daily unless otherwise specified. Steady-state tivozanib AUC and C max increased in a dose-proportional manner over the dose range of 0.89 to 1.78 mg once daily (0.67 to 1.3 times the recommended dose). Steady-state was reached by 14 days and the accumulation ratio after administration of 1.34 mg once daily was approximately 6- to 7- fold.
Mean steady-state tivozanib [coefficient of variation (CV%)] C max was 86.9 (44.7%) ng/mL and AUC 0-24h was 1510 (46.1%) ng*h/mL. Absorption The median T max of tivozanib is 10 hours with a range of 3 to 24 hours. Effect of Food No clinically significant differences in tivozanib AUC or C max were observed following administration of a high fat meal (approximately 500-600 fat calories, 250 carbohydrate calories and 150 protein calories) in healthy subjects.
Distribution The apparent volume of distribution (V/F) of tivozanib is 123 L. Protein binding of tivozanib is ≥ 99%, primarily to albumin in vitro and is independent of concentration. The mean blood-to-plasma concentration ratios ranged from 0.495 to 0.615 in healthy subjects.
Elimination The apparent clearance (CL/F) of tivozanib is
0.75L/h and the half-life is 111 hours. Metabolism Tivozanib is metabolized predominantly by CYP3A4. Following oral administration of a single radiolabeled 1.34 mg dose of tivozanib to healthy subjects, unchanged tivozanib constituted 90% of the radioactive drug components in serum.
Excretion Following oral administration of a single radiolabeled 1.34 mg dose of tivozanib to healthy subjects, 79% of the administered dose was recovered in feces (26% unchanged) and 12% in urine (unchanged tivozanib not detected). Specific Populations No clinically significant differences in the pharmacokinetics of tivozanib were observed based on age (18 years to 88 years), sex, race (93% Caucasian, 3% African American, 2% Asian, 2% others), body weight (39 kg to 158 kg), mild to severe renal impairment (CLcr 15-89 mL/min as estimated by Cockcroft-Gault) or mild hepatic impairment (total bilirubin less than or equal to ULN with AST greater than ULN or total bilirubin greater than 1 to 1.5 times ULN with any AST).
The effect of end-stage renal disease or severe hepatic impairment on tivozanib pharmacokinetics is unknown [see USE IN SPECIFIC POPULATIONS (8.6) and (8.7) ] . Patients with Hepatic Impairment Compared to subjects with normal hepatic function, tivozanib AUC tau increased by 1% in patients with mild (total bilirubin less than or equal to ULN with AST greater than ULN or total bilirubin greater than 1 to 1.5 times ULN with any AST) hepatic impairment. Compared to subjects with normal hepatic function, tivozanib AUC tau increased by 62% in patients with moderate (total bilirubin greater than 1.5 to 3 times ULN with any AST) hepatic impairment.
The effect of severe (total bilirubin greater than 3 to 10 times ULN with any AST) hepatic impairment on tivozanib pharmacokinetics has not been studied [see DOSAGE AND ADMINISTRATION (2.3) and USE IN SPECIFIC POPLATIONS (8.7) ]. Drug Interaction Studies Clinical Studies Strong CYP3A Inducers : Concomitant use of multiple doses of rifampin (strong CYP3A inducer) did not change tivozanib C max but decreased tivozanib AUC 0-INF by 52%. Strong CYP3A Inhibitors : No clinically significant differences in the pharmacokinetics of tivozanib were observed when multiple doses of ketoconazole (strong CYP3A inhibitor) was co-administered with tivozanib.
In Vitro Studies Cytochrome P450 (CYP) Enzymes : Tivozanib does not inhibit CYP1A2, CYP2A6, CYP2B6, CYP2C8, CYP2C9, CYP2C19, CYP2D6 or CYP3A4 at clinically relevant concentrations. Tivozanib does not induce CYP1A, CYP2B6, CYP2C9, CYP2C19, or CYP3A at clinically relevant concentrations. Uridine Diphosphate (UDP)-glucuronosyl Transferase (UGT) Enzymes : Tivozanib does not inhibit UGT at clinically relevant concentrations.… [Excerpted — this section continues on DailyMed.]
🧬 Pharmacodynamics ▾
12.2Pharmacodynamics Exposure-Response Relationship Tivozanib exposure-response relationships and the time course of pharmacodynamic response have not been fully characterized. Cardiac Electrophysiology At the recommended dose of FOTIVDA, no large mean increases (i.e., 20 msec) in QTc interval were observed.
🔬 Clinical Studies ▾
14 CLINICAL STUDIES The efficacy of FOTIVDA was evaluated in TIVO-3 (NCT02627963), a randomized (1:1), open- label, multicenter trial of FOTIVDA versus sorafenib in patients with relapsed or refractory advanced RCC who received 2 or 3 prior systemic treatments including at least one VEGFR kinase inhibitor other than sorafenib or tivozanib. Patients were randomized to receive FOTIVDA 1.34 mg orally once daily for 21 days on treatment followed by 7 days off treatment for a 28-day cycle, or to receive sorafenib 400 mg orally twice a day continuously, until disease progression or unacceptable toxicity.
Randomization was stratified by prior therapy [two kinase inhibitors (KIs), a KI plus an immune checkpoint inhibitor, or a KI plus other systemic agents] and by International Metastatic Renal Cell Carcinoma Database Consortium (IMDC) prognostic score. Patients were excluded if they had more than 3 prior treatments or Central Nervous System metastases. The main efficacy outcome measure was progression-free survival (PFS) assessed by a blinded independent radiology review committee.
Other efficacy endpoints were objective response rate (ORR) and overall survival (OS). The median age was 63 years (range: 30 to 90 years), 73% were male, 95% were Caucasian, ECOG performance status was 0 in 48% and 1 in 49% of patients (respectively), and 98% of patients had clear cell or clear cell component histology. Prior therapy included two KIs (45%), a KI plus an immune checkpoint inhibitor (26%), and a KI plus another systemic agent (29%).
At the time of study entry, 20% of patients had favorable, 61% intermediate, and 19% poor IMDC prognoses. Efficacy results are summarized in Table 4 and Figure 1 . Table 4.
Efficacy Results in TIVO-3 (ITT) Endpoint FOTIVDA N= 175 Sorafenib N= 175 CI: Confidence interval; HR: Hazard ratio (FOTIVDA/sorafenib); NE: not estimable. Progression Free Survival (PFS) Assessed by blinded independent radiology review committee according to RECIST v1.1. Events, n (%) 123 (70) 123 (70) Progressive Disease 103 (59) 109 (62) Death 20 (11) 14 (8) Median (95% CI), months 5.6 (4.8, 7.3) 3.9 (3.7, 5.6) HR (95% CI) Based on the Cox proportional hazards model stratified by IMDC prognostic score and prior therapy.
0.73(0.56, 0.95) P-value Based on the log-rank test stratified by IMDC prognostic score and prior therapy. 0.016 Overall Survival Deaths, n (%) 125 (71) 126 (72) Median (95% CI), months 16.4 (13.4, 21.9) 19.2 (14.9, 24.2) HR (95% CI) 0.97 (0.75, 1.24) Objective Response Rate % (95% CI) 18 (12, 24) 8 (4, 13) Median duration of response in months (95% CI) NE (9.8, NE) 5.7 (5.6, NE) Figure 1. Kaplan-Meier Plot of PFS in TIVO-3 Figure 1
🧪 Nonclinical Toxicology ▾
13 NONCLINICAL TOXICOLOGY
13.1Carcinogenesis, Mutagenesis, Impairment of Fertility Carcinogenicity studies have not been conducted with tivozanib. Tivozanib was not mutagenic in a bacterial reverse mutation (Ames) assay and was not clastogenic in an in vitro cytogenetic assay in Chinese hamster ovary cells or in an in vivo mouse bone marrow micronucleus assay. In animal studies assessing mating and fertility parameters, oral doses ≥ 0.03 mg/kg/day (0.2 times the maximum recommended clinical dose on a mg/m 2 basis) in rats were associated with increased epididymis and testis weights, and doses ≥ 0.3 mg/kg/day (2 times the maximum recommended clinical dose on a mg/m 2 basis) reduced mating and produced infertility.
An increase in embryo lethality was noted at doses ≥ 0.1 mg/kg/day (0.7 times the maximum recommended clinical dose on a mg/m 2 basis).
📄 Carcinogenesis, Mutagenesis, Impairment of Fertility ▾
13.1Carcinogenesis, Mutagenesis, Impairment of Fertility Carcinogenicity studies have not been conducted with tivozanib. Tivozanib was not mutagenic in a bacterial reverse mutation (Ames) assay and was not clastogenic in an in vitro cytogenetic assay in Chinese hamster ovary cells or in an in vivo mouse bone marrow micronucleus assay. In animal studies assessing mating and fertility parameters, oral doses ≥ 0.03 mg/kg/day (0.2 times the maximum recommended clinical dose on a mg/m 2 basis) in rats were associated with increased epididymis and testis weights, and doses ≥ 0.3 mg/kg/day (2 times the maximum recommended clinical dose on a mg/m 2 basis) reduced mating and produced infertility.
An increase in embryo lethality was noted at doses ≥ 0.1 mg/kg/day (0.7 times the maximum recommended clinical dose on a mg/m 2 basis).
📄 Patient Package Insert ▾
This Patient Information has been approved by the U.S. Food and Drug Administration. Issued: 08/2024 PATIENT INFORMATION FOTIVDA ® (fo-TIV-dah) (tivozanib) capsules What is FOTIVDA?
FOTIVDA is a prescription medicine used to treat adults with advanced kidney cancer (advanced renal cell carcinoma or RCC) that has been treated with 2 or more prior medicines and has come back or did not respond to treatment. It is not known if FOTIVDA is safe and effective in children. Before taking FOTIVDA, tell your healthcare provider about all your medical conditions, including if you: have high blood pressure. have a history of heart failure. have a history of blood clots in your veins or arteries (types of blood vessels), including stroke, heart attack, or change in vision. have bleeding problems. have thyroid problems. have liver problems. have an unhealed wound. plan to have surgery or have had recent surgery, including dental surgery.
You should stop taking FOTIVDA at least 24 days before planned surgery. See " What are the possible side effects of FOTIVDA? " are allergic to FD&C Yellow No.5 (tartrazine) or aspirin. are pregnant or plan to become pregnant. FOTIVDA can harm your unborn baby.
Females who are able to become pregnant: Your healthcare provider should do a pregnancy test before you start treatment with FOTIVDA. Use effective birth control (contraception) during treatment and for 1 month after your last dose of FOTIVDA. Talk to your healthcare provider about birth control methods that may be right for you during this time.
Tell your healthcare provider right away if you become pregnant or think you might be pregnant during treatment with FOTIVDA. Males with female partners who are able to become pregnant: Use effective birth control (contraception) during treatment and for 1 month after your last dose of FOTIVDA. If your female partner becomes pregnant during your treatment with FOTIVDA, tell your healthcare provider right away. are breastfeeding or plan to breastfeed.
It is not known if FOTIVDA passes into your breast milk. Do not breastfeed during treatment and for 1 month after your last dose of FOTIVDA. Tell your healthcare provider about all the medicines you take , including prescription and over-the-counter medicines, vitamins, and herbal supplements.
Taking FOTIVDA with certain other medicines may affect how FOTIVDA works. Know the medicines you take. Keep a list of them to show your healthcare provider and pharmacist when you get a new medicine.
How should I take FOTIVDA? Take FOTIVDA exactly as your healthcare provider tells you to take it. Take FOTIVDA 1 time each day for 21 days on treatment, followed by 7 days off treatment.
This is 1 cycle of treatment. You will repeat this cycle for as long as your healthcare provider tells you to. FOTIVDA can be taken with or without food.
Swallow the FOTIVDA capsule whole with a glass of water. Do not open the capsule. If you miss a dose of FOTIVDA, take your next dose at your next scheduled time.
Do not take 2 doses in the same day. If you take too much FOTIVDA, call your healthcare provider or go to the nearest hospital emergency room right away. What are the possible side effects of FOTIVDA?
FOTIVDA may cause serious side effects, including: High blood pressure (hypertension). High blood pressure is common with FOTIVDA and may sometimes be severe. FOTIVDA may also cause a sudden, severe increase in your blood pressure (hypertensive crisis) that can lead to death.
Your healthcare provider should check your blood pressure after 2 weeks and at least monthly during treatment with FOTIVDA. Your healthcare provider may prescribe medicine to treat your high blood pressure if you develop blood pressure problems. You should check your blood pressure regularly during treatment with FOTIVDA and tell your healthcare provider if you have increased blood pressure.
Tell your healthcare provider right away if you get any of the following signs or symptoms: confusion headaches dizziness ches… [Excerpted — this section continues on DailyMed.]
📄 Recent Major Changes ▾
Warnings and Precautions ( 5.7 ) 8/2024
📄 Package Label / Principal Display Panel ▾
PRINCIPAL DISPLAY PANEL - 0.89 mg Capsule Bottle Label NDC 45629-089-02 Rx Only 21 Capsules FOTIVDA ® (tivozanib) capsules 0.89 mg Actual Size PRINCIPAL DISPLAY PANEL - 0.89 mg Capsule Bottle Label
PRINCIPAL DISPLAY PANEL - 1.34 mg Capsule Bottle Label NDC 45629-134-02 Rx Only 21 Capsules FOTIVDA ® (tivozanib) capsules 1.34 mg Actual Size PRINCIPAL DISPLAY PANEL - 1.34 mg Capsule Bottle Label
Medicaid utilization & spend
Medicaid utilization by pack size
Medicare Part D spend CMS · PART D · 2026 (Q1)
About this NDC listing & data coverage
What data is (and isn’t) available for this NDC — tap to expand
| NDC identity (package / product / labeler codes) | ✓ Available |
| Labeler | ✓ Available |
| Product & package description | ✓ Available |
| Marketing category & status | ✓ Available |
| Active ingredient / dosage form / route | ✓ Available |
| FDA label (SPL via DailyMed) | ✓ Available |
| Package photos | ✓ Available |
| Inactive ingredients (structured) | ✓ Available |
| NADAC pharmacy acquisition price (CMS) | — Not published for this NDC CMS publishes NADAC only for NDCs reported in its retail-pharmacy survey. |
| Orange Book / therapeutic-equivalence data | ✓ Available |
| HCPCS J-code billing crosswalk | — Not published for this NDC Most self-administered / retail products have no J-code — that is normal. |
| Medicaid utilization (CMS SDUD) | ✓ Available |